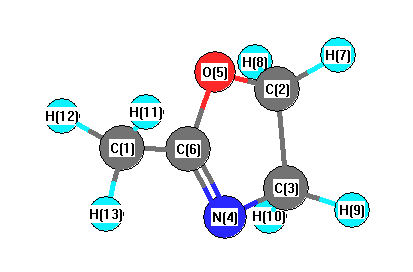
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3118 |
3100 |
2.85 |
|
|
|
| 2 |
A |
2990 |
2972 |
11.37 |
|
|
|
| 3 |
A |
2978 |
2960 |
51.25 |
|
|
|
| 4 |
A |
2962 |
2945 |
46.01 |
|
|
|
| 5 |
A |
1696 |
1686 |
120.16 |
|
|
|
| 6 |
A |
1447 |
1438 |
3.74 |
|
|
|
| 7 |
A |
1419 |
1411 |
0.32 |
|
|
|
| 8 |
A |
1399 |
1391 |
7.79 |
|
|
|
| 9 |
A |
1355 |
1347 |
60.38 |
|
|
|
| 10 |
A |
1314 |
1306 |
8.20 |
|
|
|
| 11 |
A |
1275 |
1268 |
10.91 |
|
|
|
| 12 |
A |
1206 |
1199 |
68.75 |
|
|
|
| 13 |
A |
1045 |
1039 |
6.19 |
|
|
|
| 14 |
A |
981 |
976 |
83.47 |
|
|
|
| 15 |
A |
955 |
950 |
0.74 |
|
|
|
| 16 |
A |
923 |
918 |
17.75 |
|
|
|
| 17 |
A |
883 |
878 |
20.95 |
|
|
|
| 18 |
A |
737 |
733 |
4.89 |
|
|
|
| 19 |
A |
653 |
650 |
2.13 |
|
|
|
| 20 |
A |
336 |
334 |
5.00 |
|
|
|
| 21 |
A |
3059 |
3041 |
5.84 |
|
|
|
| 22 |
A |
3028 |
3010 |
42.24 |
|
|
|
| 23 |
A |
2993 |
2976 |
6.46 |
|
|
|
| 24 |
A |
1394 |
1386 |
8.98 |
|
|
|
| 25 |
A |
1191 |
1184 |
0.10 |
|
|
|
| 26 |
A |
1160 |
1153 |
1.29 |
|
|
|
| 27 |
A |
1069 |
1063 |
0.00 |
|
|
|
| 28 |
A |
1008 |
1002 |
3.16 |
|
|
|
| 29 |
A |
795 |
790 |
0.00 |
|
|
|
| 30 |
A |
572 |
568 |
2.79 |
|
|
|
| 31 |
A |
229 |
227 |
9.11 |
|
|
|
| 32 |
A |
137 |
136 |
1.22 |
|
|
|
| 33 |
A |
59 |
59 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23182.4 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 23047.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.010 |
|
|
|
| 2 |
C |
0.081 |
|
|
|
| 3 |
C |
0.011 |
|
|
|
| 4 |
N |
-0.245 |
|
|
|
| 5 |
O |
-0.248 |
|
|
|
| 6 |
C |
0.065 |
|
|
|
| 7 |
H |
0.039 |
|
|
|
| 8 |
H |
0.039 |
|
|
|
| 9 |
H |
0.037 |
|
|
|
| 10 |
H |
0.037 |
|
|
|
| 11 |
H |
0.059 |
|
|
|
| 12 |
H |
0.059 |
|
|
|
| 13 |
H |
0.056 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.945 |
-0.740 |
0.000 |
1.200 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.368 |
1.652 |
0.000 |
| y |
1.652 |
-30.166 |
0.000 |
| z |
0.000 |
0.000 |
-35.185 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.692 |
1.652 |
0.000 |
| y |
1.652 |
7.610 |
0.000 |
| z |
0.000 |
0.000 |
0.082 |
|
| Polar |
|---|
| 3z2-r2 | 0.164 |
|---|
| x2-y2 | -10.201 |
|---|
| xy | 1.652 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.768 |
0.711 |
0.000 |
| y |
0.711 |
9.770 |
0.000 |
| z |
0.000 |
0.000 |
5.908 |
<r2> (average value of r
2) Å
2
| <r2> |
150.039 |
| (<r2>)1/2 |
12.249 |