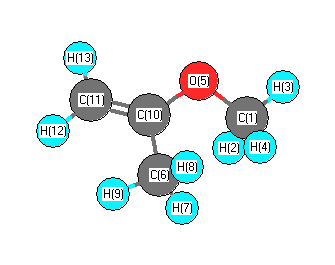
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3212 |
3193 |
5.90 |
|
|
|
| 2 |
A |
3106 |
3088 |
0.15 |
|
|
|
| 3 |
A |
3086 |
3068 |
9.81 |
|
|
|
| 4 |
A |
3069 |
3051 |
19.96 |
|
|
|
| 5 |
A |
3028 |
3010 |
13.42 |
|
|
|
| 6 |
A |
2994 |
2977 |
43.36 |
|
|
|
| 7 |
A |
2966 |
2948 |
20.91 |
|
|
|
| 8 |
A |
2925 |
2908 |
64.74 |
|
|
|
| 9 |
A |
1676 |
1666 |
109.44 |
|
|
|
| 10 |
A |
1427 |
1419 |
9.90 |
|
|
|
| 11 |
A |
1420 |
1412 |
8.14 |
|
|
|
| 12 |
A |
1412 |
1404 |
6.16 |
|
|
|
| 13 |
A |
1400 |
1392 |
17.01 |
|
|
|
| 14 |
A |
1396 |
1387 |
4.37 |
|
|
|
| 15 |
A |
1357 |
1349 |
10.78 |
|
|
|
| 16 |
A |
1333 |
1325 |
38.88 |
|
|
|
| 17 |
A |
1262 |
1254 |
164.06 |
|
|
|
| 18 |
A |
1152 |
1146 |
1.20 |
|
|
|
| 19 |
A |
1113 |
1106 |
8.00 |
|
|
|
| 20 |
A |
1095 |
1089 |
87.45 |
|
|
|
| 21 |
A |
1013 |
1007 |
1.51 |
|
|
|
| 22 |
A |
971 |
965 |
3.36 |
|
|
|
| 23 |
A |
901 |
896 |
4.40 |
|
|
|
| 24 |
A |
796 |
792 |
24.59 |
|
|
|
| 25 |
A |
791 |
786 |
28.62 |
|
|
|
| 26 |
A |
703 |
699 |
3.25 |
|
|
|
| 27 |
A |
518 |
515 |
6.99 |
|
|
|
| 28 |
A |
467 |
464 |
2.27 |
|
|
|
| 29 |
A |
409 |
406 |
0.83 |
|
|
|
| 30 |
A |
303 |
302 |
0.77 |
|
|
|
| 31 |
A |
237 |
235 |
0.56 |
|
|
|
| 32 |
A |
191 |
190 |
1.75 |
|
|
|
| 33 |
A |
75 |
75 |
4.88 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23900.3 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 23761.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.057 |
|
|
|
| 2 |
H |
0.035 |
|
|
|
| 3 |
H |
0.047 |
|
|
|
| 4 |
H |
0.031 |
|
|
|
| 5 |
O |
-0.234 |
|
|
|
| 6 |
C |
-0.033 |
|
|
|
| 7 |
H |
0.049 |
|
|
|
| 8 |
H |
0.055 |
|
|
|
| 9 |
H |
0.043 |
|
|
|
| 10 |
C |
-0.046 |
|
|
|
| 11 |
C |
-0.033 |
|
|
|
| 12 |
H |
0.014 |
|
|
|
| 13 |
H |
0.015 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.050 |
1.519 |
0.375 |
1.884 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-28.691 |
-0.377 |
0.304 |
| y |
-0.377 |
-31.527 |
-0.361 |
| z |
0.304 |
-0.361 |
-33.171 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.658 |
-0.377 |
0.304 |
| y |
-0.377 |
-0.596 |
-0.361 |
| z |
0.304 |
-0.361 |
-3.062 |
|
| Polar |
|---|
| 3z2-r2 | -6.124 |
|---|
| x2-y2 | 2.836 |
|---|
| xy | -0.377 |
|---|
| xz | 0.304 |
|---|
| yz | -0.361 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.883 |
1.027 |
-0.184 |
| y |
1.027 |
7.839 |
-0.132 |
| z |
-0.184 |
-0.132 |
5.033 |
<r2> (average value of r
2) Å
2
| <r2> |
130.388 |
| (<r2>)1/2 |
11.419 |