return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H6O (Cyclobutanone)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-230.583243
Energy at 298.15K-230.590070
HF Energy-229.812085
Nuclear repulsion energy166.437271
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3171 3042 30.04      
2 A' 3150 3022 3.25      
3 A' 3105 2979 19.14      
4 A' 3088 2963 10.80      
5 A' 1890 1813 246.30      
6 A' 1496 1436 0.38      
7 A' 1449 1390 2.28      
8 A' 1252 1202 3.16      
9 A' 1222 1172 0.45      
10 A' 1100 1055 0.13      
11 A' 989 949 1.04      
12 A' 874 839 1.02      
13 A' 734 704 2.14      
14 A' 678 650 2.62      
15 A' 402 386 1.06      
16 A' 94 91 2.81      
17 A" 3155 3027 5.44      
18 A" 3085 2960 16.63      
19 A" 1436 1377 14.69      
20 A" 1282 1230 10.51      
21 A" 1239 1188 3.68      
22 A" 1186 1137 1.70      
23 A" 1103 1058 68.27      
24 A" 952 914 0.31      
25 A" 941 903 0.12      
26 A" 651 624 0.04      
27 A" 460 441 3.86      

Unscaled Zero Point Vibrational Energy (zpe) 20091.5 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 19275.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.35337 0.15874 0.11838

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
O1 -0.197 1.872 0.000
C2 0.000 0.683 0.000
C3 0.122 -0.387 1.111
C4 0.122 -0.387 -1.111
C5 -0.079 -1.469 0.000
H6 -1.103 -1.878 0.000
H7 0.640 -2.303 0.000
H8 1.139 -0.391 1.544
H9 1.139 -0.391 -1.544
H10 -0.616 -0.329 1.929
H11 -0.616 -0.329 -1.929

Atom - Atom Distances (Å)
  O1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11
O11.20582.53782.53783.34333.85824.25823.04803.04802.95672.9567
C21.20581.54721.54722.15302.78803.05342.19882.19882.26352.2635
C32.53781.54722.22301.56422.22712.27491.10542.84391.10303.1292
C42.53781.54722.22301.56422.22712.27492.84391.10543.12921.1030
C53.34332.15301.56421.56421.10301.10132.24302.24302.30412.3041
H63.85822.78802.22712.22711.10301.79473.10243.10242.52122.5212
H74.25823.05342.27492.27491.10131.79472.50802.50803.03253.0325
H83.04802.19881.10542.84392.24303.10242.50803.08861.79823.8921
H93.04802.19882.84391.10542.24303.10242.50803.08863.89211.7982
H102.95672.26351.10303.12922.30412.52123.03251.79823.89213.8577
H112.95672.26353.12921.10302.30412.52123.03253.89211.79823.8577

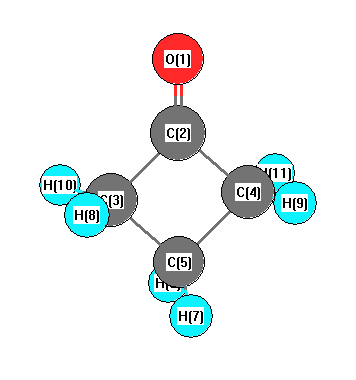
picture of Cyclobutanone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
O1 C2 C3 134.011 O1 C2 C4 134.011
C2 C3 C5 87.569 C2 C3 H8 110.873
C2 C3 H10 116.315 C2 C4 C5 87.569
C2 C4 H9 110.873 C2 C4 H11 116.315
C3 C2 C4 91.839 C3 C5 C4 90.563
C3 C5 H6 112.074 C3 C5 H7 116.101
C4 C5 H6 112.074 C4 C5 H7 116.101
C5 C3 H8 113.203 C5 C3 H10 118.479
C5 C4 H9 113.203 C5 C4 H11 118.479
H6 C5 H7 109.015 H8 C3 H10 109.026
H9 C4 H11 109.026
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability