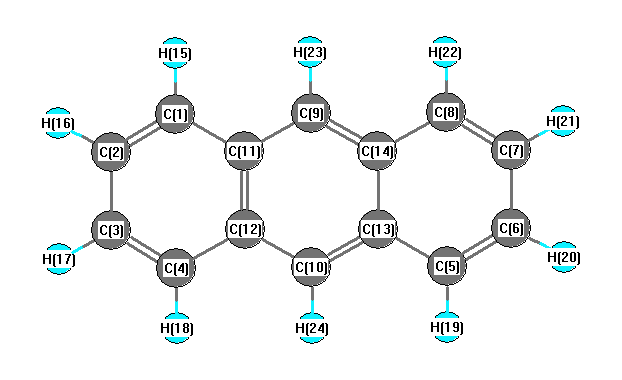
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3135 |
3116 |
0.00 |
|
|
|
| 2 |
Ag |
3108 |
3090 |
0.00 |
|
|
|
| 3 |
Ag |
3098 |
3080 |
0.00 |
|
|
|
| 4 |
Ag |
1562 |
1553 |
0.00 |
|
|
|
| 5 |
Ag |
1478 |
1470 |
0.00 |
|
|
|
| 6 |
Ag |
1423 |
1415 |
0.00 |
|
|
|
| 7 |
Ag |
1268 |
1261 |
0.00 |
|
|
|
| 8 |
Ag |
1138 |
1132 |
0.00 |
|
|
|
| 9 |
Ag |
1015 |
1010 |
0.00 |
|
|
|
| 10 |
Ag |
747 |
743 |
0.00 |
|
|
|
| 11 |
Ag |
616 |
612 |
0.00 |
|
|
|
| 12 |
Ag |
389 |
387 |
0.00 |
|
|
|
| 13 |
Au |
964 |
958 |
0.00 |
|
|
|
| 14 |
Au |
850 |
845 |
0.00 |
|
|
|
| 15 |
Au |
748 |
744 |
0.00 |
|
|
|
| 16 |
Au |
493 |
490 |
0.00 |
|
|
|
| 17 |
Au |
118 |
117 |
0.00 |
|
|
|
| 18 |
B1g |
935 |
930 |
0.00 |
|
|
|
| 19 |
B1g |
750 |
745 |
0.00 |
|
|
|
| 20 |
B1g |
472 |
469 |
0.00 |
|
|
|
| 21 |
B1g |
231 |
229 |
0.00 |
|
|
|
| 22 |
B1u |
3122 |
3104 |
51.84 |
|
|
|
| 23 |
B1u |
3103 |
3085 |
9.63 |
|
|
|
| 24 |
B1u |
3096 |
3078 |
16.17 |
|
|
|
| 25 |
B1u |
1633 |
1624 |
4.51 |
|
|
|
| 26 |
B1u |
1446 |
1437 |
1.48 |
|
|
|
| 27 |
B1u |
1299 |
1292 |
7.79 |
|
|
|
| 28 |
B1u |
1240 |
1233 |
5.59 |
|
|
|
| 29 |
B1u |
1129 |
1122 |
7.56 |
|
|
|
| 30 |
B1u |
884 |
879 |
1.12 |
|
|
|
| 31 |
B1u |
642 |
638 |
0.31 |
|
|
|
| 32 |
B1u |
228 |
227 |
1.23 |
|
|
|
| 33 |
B2g |
964 |
959 |
0.00 |
|
|
|
| 34 |
B2g |
896 |
891 |
0.00 |
|
|
|
| 35 |
B2g |
826 |
821 |
0.00 |
|
|
|
| 36 |
B2g |
769 |
765 |
0.00 |
|
|
|
| 37 |
B2g |
577 |
574 |
0.00 |
|
|
|
| 38 |
B2g |
263 |
262 |
0.00 |
|
|
|
| 39 |
B2u |
3134 |
3116 |
48.98 |
|
|
|
| 40 |
B2u |
3107 |
3089 |
0.02 |
|
|
|
| 41 |
B2u |
1548 |
1539 |
4.89 |
|
|
|
| 42 |
B2u |
1436 |
1428 |
1.03 |
|
|
|
| 43 |
B2u |
1400 |
1392 |
4.18 |
|
|
|
| 44 |
B2u |
1386 |
1378 |
1.08 |
|
|
|
| 45 |
B2u |
1154 |
1147 |
3.62 |
|
|
|
| 46 |
B2u |
1129 |
1123 |
0.42 |
|
|
|
| 47 |
B2u |
1012 |
1006 |
7.16 |
|
|
|
| 48 |
B2u |
809 |
804 |
0.03 |
|
|
|
| 49 |
B2u |
596 |
593 |
7.76 |
|
|
|
| 50 |
B3g |
3122 |
3103 |
0.00 |
|
|
|
| 51 |
B3g |
3102 |
3084 |
0.00 |
|
|
|
| 52 |
B3g |
1633 |
1623 |
0.00 |
|
|
|
| 53 |
B3g |
1586 |
1577 |
0.00 |
|
|
|
| 54 |
B3g |
1359 |
1351 |
0.00 |
|
|
|
| 55 |
B3g |
1239 |
1232 |
0.00 |
|
|
|
| 56 |
B3g |
1162 |
1155 |
0.00 |
|
|
|
| 57 |
B3g |
1095 |
1089 |
0.00 |
|
|
|
| 58 |
B3g |
895 |
890 |
0.00 |
|
|
|
| 59 |
B3g |
515 |
512 |
0.00 |
|
|
|
| 60 |
B3g |
380 |
378 |
0.00 |
|
|
|
| 61 |
B3u |
940 |
935 |
4.64 |
|
|
|
| 62 |
B3u |
871 |
866 |
37.86 |
|
|
|
| 63 |
B3u |
720 |
716 |
54.81 |
|
|
|
| 64 |
B3u |
465 |
463 |
17.24 |
|
|
|
| 65 |
B3u |
376 |
374 |
0.01 |
|
|
|
| 66 |
B3u |
90 |
90 |
0.98 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 41456.8 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 41216.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.009 |
|
|
|
| 2 |
C |
0.030 |
|
|
|
| 3 |
C |
0.030 |
|
|
|
| 4 |
C |
0.009 |
|
|
|
| 5 |
C |
0.009 |
|
|
|
| 6 |
C |
0.030 |
|
|
|
| 7 |
C |
0.030 |
|
|
|
| 8 |
C |
0.009 |
|
|
|
| 9 |
C |
-0.038 |
|
|
|
| 10 |
C |
-0.038 |
|
|
|
| 11 |
C |
0.063 |
|
|
|
| 12 |
C |
0.063 |
|
|
|
| 13 |
C |
0.063 |
|
|
|
| 14 |
C |
0.063 |
|
|
|
| 15 |
H |
-0.037 |
|
|
|
| 16 |
H |
-0.020 |
|
|
|
| 17 |
H |
-0.020 |
|
|
|
| 18 |
H |
-0.037 |
|
|
|
| 19 |
H |
-0.037 |
|
|
|
| 20 |
H |
-0.020 |
|
|
|
| 21 |
H |
-0.020 |
|
|
|
| 22 |
H |
-0.037 |
|
|
|
| 23 |
H |
-0.050 |
|
|
|
| 24 |
H |
-0.050 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-86.964 |
0.000 |
0.000 |
| y |
0.000 |
-70.482 |
0.000 |
| z |
0.000 |
0.000 |
-71.554 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-15.946 |
0.000 |
0.000 |
| y |
0.000 |
8.777 |
0.000 |
| z |
0.000 |
0.000 |
7.170 |
|
| Polar |
|---|
| 3z2-r2 | 14.339 |
|---|
| x2-y2 | -16.482 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.593 |
0.000 |
0.000 |
| y |
0.000 |
43.117 |
0.000 |
| z |
0.000 |
0.000 |
23.954 |
<r2> (average value of r
2) Å
2
| <r2> |
809.902 |
| (<r2>)1/2 |
28.459 |