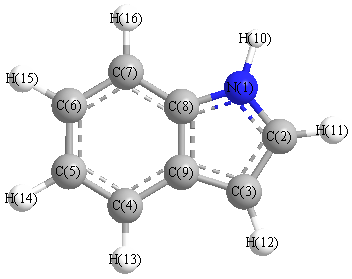
Vibrational Frequencies calculated at BLYP/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3545 |
3550 |
47.92 |
|
|
|
| 2 |
A' |
3182 |
3187 |
5.21 |
|
|
|
| 3 |
A' |
3161 |
3167 |
2.18 |
|
|
|
| 4 |
A' |
3113 |
3118 |
25.24 |
|
|
|
| 5 |
A' |
3100 |
3105 |
41.49 |
|
|
|
| 6 |
A' |
3089 |
3094 |
4.38 |
|
|
|
| 7 |
A' |
3083 |
3088 |
3.27 |
|
|
|
| 8 |
A' |
1605 |
1608 |
1.97 |
|
|
|
| 9 |
A' |
1562 |
1564 |
1.02 |
|
|
|
| 10 |
A' |
1497 |
1499 |
4.31 |
|
|
|
| 11 |
A' |
1474 |
1477 |
2.04 |
|
|
|
| 12 |
A' |
1432 |
1434 |
16.33 |
|
|
|
| 13 |
A' |
1404 |
1406 |
5.76 |
|
|
|
| 14 |
A' |
1341 |
1343 |
30.58 |
|
|
|
| 15 |
A' |
1329 |
1332 |
3.90 |
|
|
|
| 16 |
A' |
1258 |
1260 |
12.63 |
|
|
|
| 17 |
A' |
1222 |
1224 |
9.72 |
|
|
|
| 18 |
A' |
1177 |
1179 |
2.05 |
|
|
|
| 19 |
A' |
1133 |
1135 |
1.15 |
|
|
|
| 20 |
A' |
1102 |
1104 |
0.07 |
|
|
|
| 21 |
A' |
1069 |
1071 |
19.27 |
|
|
|
| 22 |
A' |
1043 |
1045 |
6.01 |
|
|
|
| 23 |
A' |
1001 |
1003 |
5.27 |
|
|
|
| 24 |
A' |
878 |
880 |
6.00 |
|
|
|
| 25 |
A' |
857 |
859 |
0.27 |
|
|
|
| 26 |
A' |
751 |
752 |
1.81 |
|
|
|
| 27 |
A' |
597 |
598 |
1.06 |
|
|
|
| 28 |
A' |
536 |
537 |
0.04 |
|
|
|
| 29 |
A' |
393 |
394 |
3.56 |
|
|
|
| 30 |
A" |
948 |
949 |
0.01 |
|
|
|
| 31 |
A" |
904 |
906 |
1.11 |
|
|
|
| 32 |
A" |
834 |
836 |
2.23 |
|
|
|
| 33 |
A" |
818 |
820 |
2.57 |
|
|
|
| 34 |
A" |
762 |
764 |
5.05 |
|
|
|
| 35 |
A" |
730 |
732 |
44.04 |
|
|
|
| 36 |
A" |
703 |
704 |
31.39 |
|
|
|
| 37 |
A" |
598 |
599 |
6.97 |
|
|
|
| 38 |
A" |
569 |
570 |
1.22 |
|
|
|
| 39 |
A" |
421 |
422 |
0.63 |
|
|
|
| 40 |
A" |
382 |
383 |
58.37 |
|
|
|
| 41 |
A" |
238 |
239 |
0.03 |
|
|
|
| 42 |
A" |
209 |
209 |
8.26 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 27525.8 cm
-1
Scaled (by 1.0016) Zero Point Vibrational Energy (zpe) 27569.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.055 |
|
|
|
| 2 |
C |
0.068 |
|
|
|
| 3 |
C |
-0.014 |
|
|
|
| 4 |
C |
-0.011 |
|
|
|
| 5 |
C |
0.060 |
|
|
|
| 6 |
C |
0.054 |
|
|
|
| 7 |
C |
0.035 |
|
|
|
| 8 |
C |
0.066 |
|
|
|
| 9 |
C |
0.106 |
|
|
|
| 10 |
H |
0.066 |
|
|
|
| 11 |
H |
-0.034 |
|
|
|
| 12 |
H |
-0.073 |
|
|
|
| 13 |
H |
-0.074 |
|
|
|
| 14 |
H |
-0.062 |
|
|
|
| 15 |
H |
-0.061 |
|
|
|
| 16 |
H |
-0.070 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.956 |
2.077 |
0.000 |
2.286 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.892 |
-2.286 |
0.000 |
| y |
-2.286 |
-43.009 |
0.000 |
| z |
0.000 |
0.000 |
-56.851 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.038 |
-2.286 |
0.000 |
| y |
-2.286 |
9.362 |
0.000 |
| z |
0.000 |
0.000 |
-11.400 |
|
| Polar |
|---|
| 3z2-r2 | -22.801 |
|---|
| x2-y2 | -4.883 |
|---|
| xy | -2.286 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
19.502 |
-1.565 |
0.000 |
| y |
-1.565 |
15.547 |
0.000 |
| z |
0.000 |
0.000 |
5.054 |
<r2> (average value of r
2) Å
2
| <r2> |
287.674 |
| (<r2>)1/2 |
16.961 |