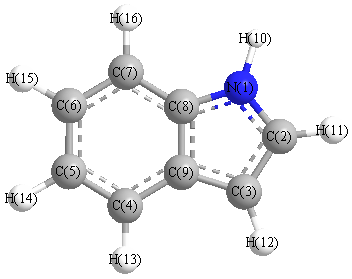
Vibrational Frequencies calculated at B3PW91/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3684 |
3555 |
74.59 |
|
|
|
| 2 |
A' |
3279 |
3164 |
2.58 |
|
|
|
| 3 |
A' |
3258 |
3144 |
0.78 |
|
|
|
| 4 |
A' |
3214 |
3101 |
16.35 |
|
|
|
| 5 |
A' |
3202 |
3090 |
28.72 |
|
|
|
| 6 |
A' |
3191 |
3080 |
3.22 |
|
|
|
| 7 |
A' |
3184 |
3073 |
1.12 |
|
|
|
| 8 |
A' |
1686 |
1627 |
2.09 |
|
|
|
| 9 |
A' |
1640 |
1582 |
1.24 |
|
|
|
| 10 |
A' |
1567 |
1512 |
7.77 |
|
|
|
| 11 |
A' |
1533 |
1479 |
4.92 |
|
|
|
| 12 |
A' |
1493 |
1441 |
21.70 |
|
|
|
| 13 |
A' |
1465 |
1413 |
8.01 |
|
|
|
| 14 |
A' |
1403 |
1354 |
38.49 |
|
|
|
| 15 |
A' |
1374 |
1326 |
6.70 |
|
|
|
| 16 |
A' |
1311 |
1265 |
7.43 |
|
|
|
| 17 |
A' |
1259 |
1215 |
9.14 |
|
|
|
| 18 |
A' |
1222 |
1179 |
2.69 |
|
|
|
| 19 |
A' |
1162 |
1122 |
0.93 |
|
|
|
| 20 |
A' |
1138 |
1098 |
0.30 |
|
|
|
| 21 |
A' |
1111 |
1072 |
24.47 |
|
|
|
| 22 |
A' |
1079 |
1041 |
4.66 |
|
|
|
| 23 |
A' |
1039 |
1003 |
6.49 |
|
|
|
| 24 |
A' |
907 |
875 |
6.05 |
|
|
|
| 25 |
A' |
881 |
850 |
0.36 |
|
|
|
| 26 |
A' |
778 |
751 |
2.70 |
|
|
|
| 27 |
A' |
613 |
591 |
1.25 |
|
|
|
| 28 |
A' |
549 |
530 |
0.05 |
|
|
|
| 29 |
A' |
404 |
389 |
4.15 |
|
|
|
| 30 |
A" |
982 |
947 |
0.01 |
|
|
|
| 31 |
A" |
941 |
908 |
1.59 |
|
|
|
| 32 |
A" |
866 |
836 |
2.78 |
|
|
|
| 33 |
A" |
858 |
828 |
0.05 |
|
|
|
| 34 |
A" |
787 |
759 |
8.48 |
|
|
|
| 35 |
A" |
753 |
727 |
70.88 |
|
|
|
| 36 |
A" |
730 |
705 |
27.18 |
|
|
|
| 37 |
A" |
619 |
597 |
7.04 |
|
|
|
| 38 |
A" |
587 |
566 |
0.94 |
|
|
|
| 39 |
A" |
433 |
418 |
0.00 |
|
|
|
| 40 |
A" |
410 |
396 |
69.02 |
|
|
|
| 41 |
A" |
246 |
237 |
0.10 |
|
|
|
| 42 |
A" |
215 |
208 |
9.81 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28525.1 cm
-1
Scaled (by 0.965) Zero Point Vibrational Energy (zpe) 27526.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.158 |
|
|
|
| 2 |
C |
0.064 |
|
|
|
| 3 |
C |
-0.063 |
|
|
|
| 4 |
C |
-0.038 |
|
|
|
| 5 |
C |
-0.001 |
|
|
|
| 6 |
C |
-0.006 |
|
|
|
| 7 |
C |
-0.006 |
|
|
|
| 8 |
C |
0.117 |
|
|
|
| 9 |
C |
0.021 |
|
|
|
| 10 |
H |
0.097 |
|
|
|
| 11 |
H |
0.019 |
|
|
|
| 12 |
H |
-0.015 |
|
|
|
| 13 |
H |
-0.015 |
|
|
|
| 14 |
H |
-0.004 |
|
|
|
| 15 |
H |
-0.003 |
|
|
|
| 16 |
H |
-0.009 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.930 |
2.079 |
0.000 |
2.278 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.428 |
-2.278 |
0.000 |
| y |
-2.278 |
-41.638 |
0.000 |
| z |
0.000 |
0.000 |
-57.050 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.916 |
-2.278 |
0.000 |
| y |
-2.278 |
10.101 |
0.000 |
| z |
0.000 |
0.000 |
-13.017 |
|
| Polar |
|---|
| 3z2-r2 | -26.034 |
|---|
| x2-y2 | -4.790 |
|---|
| xy | -2.278 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
18.512 |
-1.490 |
0.000 |
| y |
-1.490 |
14.931 |
0.000 |
| z |
0.000 |
0.000 |
5.061 |
<r2> (average value of r
2) Å
2
| <r2> |
282.375 |
| (<r2>)1/2 |
16.804 |