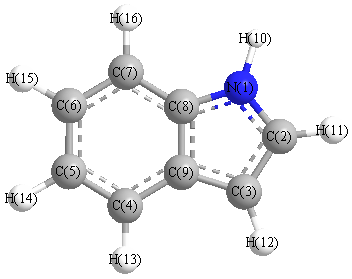
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3578 |
3557 |
58.20 |
|
|
|
| 2 |
A' |
3202 |
3184 |
2.60 |
|
|
|
| 3 |
A' |
3182 |
3164 |
0.92 |
|
|
|
| 4 |
A' |
3136 |
3118 |
17.05 |
|
|
|
| 5 |
A' |
3124 |
3106 |
30.06 |
|
|
|
| 6 |
A' |
3113 |
3095 |
3.33 |
|
|
|
| 7 |
A' |
3106 |
3088 |
2.14 |
|
|
|
| 8 |
A' |
1632 |
1622 |
1.77 |
|
|
|
| 9 |
A' |
1585 |
1576 |
1.28 |
|
|
|
| 10 |
A' |
1515 |
1507 |
5.59 |
|
|
|
| 11 |
A' |
1484 |
1476 |
3.66 |
|
|
|
| 12 |
A' |
1454 |
1445 |
13.29 |
|
|
|
| 13 |
A' |
1421 |
1413 |
4.98 |
|
|
|
| 14 |
A' |
1371 |
1363 |
38.73 |
|
|
|
| 15 |
A' |
1331 |
1323 |
6.35 |
|
|
|
| 16 |
A' |
1275 |
1268 |
6.82 |
|
|
|
| 17 |
A' |
1220 |
1213 |
8.67 |
|
|
|
| 18 |
A' |
1185 |
1178 |
2.33 |
|
|
|
| 19 |
A' |
1128 |
1121 |
0.78 |
|
|
|
| 20 |
A' |
1105 |
1099 |
0.34 |
|
|
|
| 21 |
A' |
1078 |
1072 |
20.31 |
|
|
|
| 22 |
A' |
1044 |
1038 |
4.84 |
|
|
|
| 23 |
A' |
1010 |
1004 |
6.13 |
|
|
|
| 24 |
A' |
880 |
874 |
5.87 |
|
|
|
| 25 |
A' |
857 |
852 |
0.34 |
|
|
|
| 26 |
A' |
757 |
752 |
2.06 |
|
|
|
| 27 |
A' |
595 |
591 |
1.18 |
|
|
|
| 28 |
A' |
535 |
532 |
0.04 |
|
|
|
| 29 |
A' |
392 |
390 |
3.71 |
|
|
|
| 30 |
A" |
943 |
938 |
0.01 |
|
|
|
| 31 |
A" |
901 |
896 |
1.29 |
|
|
|
| 32 |
A" |
830 |
825 |
2.51 |
|
|
|
| 33 |
A" |
817 |
813 |
1.89 |
|
|
|
| 34 |
A" |
760 |
756 |
5.26 |
|
|
|
| 35 |
A" |
726 |
721 |
56.29 |
|
|
|
| 36 |
A" |
701 |
697 |
31.96 |
|
|
|
| 37 |
A" |
601 |
597 |
7.56 |
|
|
|
| 38 |
A" |
570 |
567 |
1.03 |
|
|
|
| 39 |
A" |
419 |
417 |
0.04 |
|
|
|
| 40 |
A" |
398 |
395 |
60.53 |
|
|
|
| 41 |
A" |
238 |
236 |
0.11 |
|
|
|
| 42 |
A" |
208 |
207 |
8.67 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 27702.3 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 27541.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.109 |
|
|
|
| 2 |
C |
0.060 |
|
|
|
| 3 |
C |
-0.044 |
|
|
|
| 4 |
C |
-0.034 |
|
|
|
| 5 |
C |
0.024 |
|
|
|
| 6 |
C |
0.017 |
|
|
|
| 7 |
C |
0.011 |
|
|
|
| 8 |
C |
0.104 |
|
|
|
| 9 |
C |
0.060 |
|
|
|
| 10 |
H |
0.083 |
|
|
|
| 11 |
H |
-0.003 |
|
|
|
| 12 |
H |
-0.040 |
|
|
|
| 13 |
H |
-0.040 |
|
|
|
| 14 |
H |
-0.028 |
|
|
|
| 15 |
H |
-0.027 |
|
|
|
| 16 |
H |
-0.035 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.958 |
2.118 |
0.000 |
2.325 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.077 |
-2.317 |
0.000 |
| y |
-2.317 |
-42.164 |
0.000 |
| z |
0.000 |
0.000 |
-57.040 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.525 |
-2.317 |
0.000 |
| y |
-2.317 |
9.894 |
0.000 |
| z |
0.000 |
0.000 |
-12.419 |
|
| Polar |
|---|
| 3z2-r2 | -24.838 |
|---|
| x2-y2 | -4.913 |
|---|
| xy | -2.317 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
19.326 |
-1.553 |
0.000 |
| y |
-1.553 |
15.396 |
0.000 |
| z |
0.000 |
0.000 |
5.102 |
<r2> (average value of r
2) Å
2
| <r2> |
285.558 |
| (<r2>)1/2 |
16.898 |