return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H4N4 (purine)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-410.806952
Energy at 298.15K-410.814817
HF Energy-409.499904
Nuclear repulsion energy410.037030
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3697 3547 105.22      
2 A' 3274 3141 1.00      
3 A' 3218 3087 17.64      
4 A' 3208 3078 13.79      
5 A' 1690 1622 67.75      
6 A' 1660 1592 83.70      
7 A' 1567 1503 28.50      
8 A' 1514 1452 17.30      
9 A' 1453 1394 78.48      
10 A' 1434 1376 6.85      
11 A' 1381 1325 60.37      
12 A' 1329 1275 9.29      
13 A' 1300 1247 4.19      
14 A' 1246 1196 54.24      
15 A' 1216 1167 4.71      
16 A' 1139 1092 6.41      
17 A' 1088 1044 13.47      
18 A' 950 911 0.51      
19 A' 919 881 14.53      
20 A' 810 777 14.40      
21 A' 660 633 0.41      
22 A' 571 548 3.39      
23 A' 442 424 12.36      
24 A" 1011 970 0.13      
25 A" 953 914 8.35      
26 A" 899 863 7.25      
27 A" 812 779 7.65      
28 A" 665 638 5.68      
29 A" 622 597 23.35      
30 A" 490 470 85.91      
31 A" 416 399 13.86      
32 A" 246 236 0.11      
33 A" 232 223 5.60      

Unscaled Zero Point Vibrational Energy (zpe) 21054.8 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 20200.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.13616 0.05792 0.04063

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 -1.827 -1.268 0.000
C2 -2.109 0.065 0.000
N3 -1.279 1.110 0.000
C4 0.000 0.707 0.000
C5 0.457 -0.625 0.000
C6 -0.542 -1.617 0.000
N7 1.849 -0.695 0.000
C8 2.189 0.573 0.000
N9 1.131 1.476 0.000
H10 -3.175 0.306 0.000
H11 -0.310 -2.688 0.000
H12 3.224 0.914 0.000
H13 1.174 2.488 0.000

Atom - Atom Distances (Å)
  N1 C2 N3 C4 C5 C6 N7 C8 N9 H10 H11 H12 H13
N11.36182.43972.69062.37281.33233.72104.41844.03412.07182.07825.50254.8074
C21.36181.33432.20452.65672.29864.03024.32813.53321.09333.28815.40004.0799
N32.43971.33431.34072.45412.82433.61113.50962.43712.05963.91904.50722.8132
C42.69062.20451.34071.40872.38632.32052.19351.36693.20053.40923.23072.1327
C52.37282.65672.45411.40871.40701.39422.10662.20623.74942.20053.16663.1949
C61.33232.29862.82432.38631.40702.56253.50043.51543.26061.09584.53714.4488
N73.72104.03023.61112.32051.39422.56251.31232.28605.12302.93862.11613.2537
C84.41844.32813.50962.19352.10663.50041.31231.39145.37134.10851.08952.1681
N94.03413.53322.43711.36692.20623.51542.28601.39144.46184.40552.16741.0135
H102.07181.09332.05963.20053.74943.26065.12305.37134.46184.14376.42814.8655
H112.07823.28813.91903.40922.20051.09582.93864.10854.40554.14375.04625.3844
H125.50255.40004.50723.23073.16664.53712.11611.08952.16746.42815.04622.5851
H134.80744.07992.81322.13273.19494.44883.25372.16811.01354.86555.38442.5851

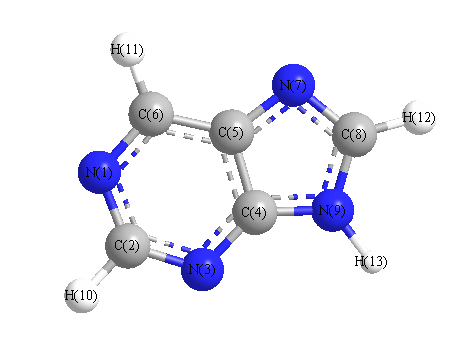
picture of purine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 N3 129.617 N1 C2 H10 114.661
N1 C6 C5 120.010 N1 C6 H11 117.388
C2 N1 C6 117.116 C2 N3 C4 110.992
N3 C2 H10 115.722 N3 C4 C5 126.388
N3 C4 N9 128.336 C4 C5 C6 115.878
C4 C5 N7 111.770 C4 N9 C8 105.354
C4 N9 H13 126.622 C5 C4 N9 105.277
C5 C6 H11 122.602 C5 N7 C8 102.177
C6 C5 N7 132.353 N7 C8 N9 115.422
N7 C8 H12 123.281 C8 N9 H13 128.024
N9 C8 H12 121.297
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability