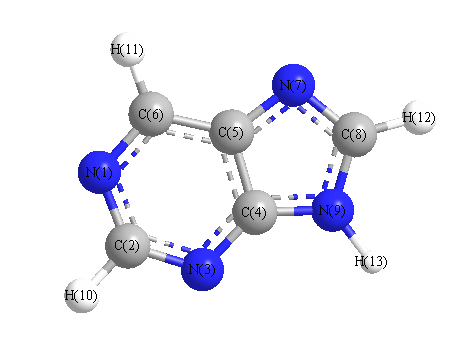
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3542 |
3521 |
67.24 |
|
|
|
| 2 |
A' |
3173 |
3155 |
0.37 |
|
|
|
| 3 |
A' |
3101 |
3083 |
16.27 |
|
|
|
| 4 |
A' |
3089 |
3071 |
16.99 |
|
|
|
| 5 |
A' |
1609 |
1600 |
49.20 |
|
|
|
| 6 |
A' |
1575 |
1566 |
58.60 |
|
|
|
| 7 |
A' |
1474 |
1465 |
22.93 |
|
|
|
| 8 |
A' |
1439 |
1430 |
2.75 |
|
|
|
| 9 |
A' |
1390 |
1382 |
49.75 |
|
|
|
| 10 |
A' |
1368 |
1360 |
12.67 |
|
|
|
| 11 |
A' |
1328 |
1321 |
62.73 |
|
|
|
| 12 |
A' |
1295 |
1287 |
20.81 |
|
|
|
| 13 |
A' |
1271 |
1263 |
8.91 |
|
|
|
| 14 |
A' |
1238 |
1231 |
19.78 |
|
|
|
| 15 |
A' |
1165 |
1158 |
8.45 |
|
|
|
| 16 |
A' |
1102 |
1095 |
5.42 |
|
|
|
| 17 |
A' |
1045 |
1039 |
11.50 |
|
|
|
| 18 |
A' |
914 |
909 |
1.52 |
|
|
|
| 19 |
A' |
882 |
877 |
11.12 |
|
|
|
| 20 |
A' |
785 |
781 |
12.15 |
|
|
|
| 21 |
A' |
637 |
633 |
0.57 |
|
|
|
| 22 |
A' |
552 |
548 |
3.77 |
|
|
|
| 23 |
A' |
430 |
428 |
11.34 |
|
|
|
| 24 |
A" |
950 |
945 |
0.39 |
|
|
|
| 25 |
A" |
900 |
895 |
9.07 |
|
|
|
| 26 |
A" |
831 |
826 |
5.25 |
|
|
|
| 27 |
A" |
791 |
787 |
9.22 |
|
|
|
| 28 |
A" |
645 |
641 |
2.32 |
|
|
|
| 29 |
A" |
606 |
603 |
25.31 |
|
|
|
| 30 |
A" |
522 |
519 |
73.42 |
|
|
|
| 31 |
A" |
398 |
395 |
2.47 |
|
|
|
| 32 |
A" |
233 |
232 |
0.37 |
|
|
|
| 33 |
A" |
224 |
223 |
4.45 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20250.7 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 20133.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.186 |
|
|
|
| 2 |
C |
0.105 |
|
|
|
| 3 |
N |
-0.199 |
|
|
|
| 4 |
C |
0.190 |
|
|
|
| 5 |
C |
-0.011 |
|
|
|
| 6 |
C |
0.113 |
|
|
|
| 7 |
N |
-0.208 |
|
|
|
| 8 |
C |
0.136 |
|
|
|
| 9 |
N |
-0.097 |
|
|
|
| 10 |
H |
0.018 |
|
|
|
| 11 |
H |
0.015 |
|
|
|
| 12 |
H |
0.025 |
|
|
|
| 13 |
H |
0.100 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.157 |
2.960 |
0.000 |
3.663 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-49.904 |
5.987 |
0.000 |
| y |
5.987 |
-46.637 |
0.000 |
| z |
0.000 |
0.000 |
-51.317 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.927 |
5.987 |
0.000 |
| y |
5.987 |
3.974 |
0.000 |
| z |
0.000 |
0.000 |
-3.047 |
|
| Polar |
|---|
| 3z2-r2 | -6.093 |
|---|
| x2-y2 | -3.267 |
|---|
| xy | 5.987 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
15.741 |
1.499 |
0.000 |
| y |
1.499 |
12.003 |
0.000 |
| z |
0.000 |
0.000 |
4.690 |
<r2> (average value of r
2) Å
2
| <r2> |
256.952 |
| (<r2>)1/2 |
16.030 |