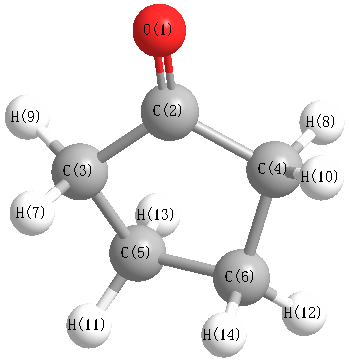
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3049 |
3032 |
5.46 |
|
|
|
| 2 |
A |
3037 |
3020 |
20.86 |
|
|
|
| 3 |
A |
2972 |
2955 |
22.06 |
|
|
|
| 4 |
A |
2963 |
2946 |
5.02 |
|
|
|
| 5 |
A |
1781 |
1770 |
203.43 |
|
|
|
| 6 |
A |
1429 |
1420 |
1.52 |
|
|
|
| 7 |
A |
1363 |
1355 |
0.13 |
|
|
|
| 8 |
A |
1286 |
1278 |
0.92 |
|
|
|
| 9 |
A |
1247 |
1239 |
0.96 |
|
|
|
| 10 |
A |
1167 |
1161 |
0.18 |
|
|
|
| 11 |
A |
1128 |
1121 |
0.14 |
|
|
|
| 12 |
A |
1014 |
1009 |
0.73 |
|
|
|
| 13 |
A |
920 |
914 |
0.22 |
|
|
|
| 14 |
A |
889 |
884 |
0.36 |
|
|
|
| 15 |
A |
790 |
786 |
1.81 |
|
|
|
| 16 |
A |
684 |
680 |
0.79 |
|
|
|
| 17 |
A |
541 |
538 |
5.15 |
|
|
|
| 18 |
A |
236 |
235 |
0.17 |
|
|
|
| 19 |
B |
3050 |
3032 |
18.33 |
|
|
|
| 20 |
B |
3042 |
3024 |
26.70 |
|
|
|
| 21 |
B |
2977 |
2960 |
38.32 |
|
|
|
| 22 |
B |
2963 |
2946 |
5.23 |
|
|
|
| 23 |
B |
1413 |
1405 |
4.71 |
|
|
|
| 24 |
B |
1364 |
1356 |
19.12 |
|
|
|
| 25 |
B |
1284 |
1276 |
5.44 |
|
|
|
| 26 |
B |
1240 |
1233 |
8.15 |
|
|
|
| 27 |
B |
1197 |
1190 |
8.47 |
|
|
|
| 28 |
B |
1115 |
1109 |
59.48 |
|
|
|
| 29 |
B |
1107 |
1100 |
17.42 |
|
|
|
| 30 |
B |
952 |
946 |
9.92 |
|
|
|
| 31 |
B |
886 |
881 |
0.45 |
|
|
|
| 32 |
B |
817 |
813 |
12.61 |
|
|
|
| 33 |
B |
566 |
563 |
2.59 |
|
|
|
| 34 |
B |
455 |
453 |
3.70 |
|
|
|
| 35 |
B |
433 |
431 |
1.97 |
|
|
|
| 36 |
B |
97 |
96 |
4.24 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25725.9 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 25576.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.180 |
|
|
|
| 2 |
C |
0.009 |
|
|
|
| 3 |
C |
-0.019 |
|
|
|
| 4 |
C |
-0.019 |
|
|
|
| 5 |
C |
-0.047 |
|
|
|
| 6 |
C |
-0.047 |
|
|
|
| 7 |
H |
0.057 |
|
|
|
| 8 |
H |
0.057 |
|
|
|
| 9 |
H |
0.040 |
|
|
|
| 10 |
H |
0.040 |
|
|
|
| 11 |
H |
0.021 |
|
|
|
| 12 |
H |
0.021 |
|
|
|
| 13 |
H |
0.032 |
|
|
|
| 14 |
H |
0.032 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-2.655 |
2.655 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.223 |
-0.260 |
0.000 |
| y |
-0.260 |
-35.152 |
0.000 |
| z |
0.000 |
0.000 |
-43.544 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.125 |
-0.260 |
0.000 |
| y |
-0.260 |
4.231 |
0.000 |
| z |
0.000 |
0.000 |
-8.356 |
|
| Polar |
|---|
| 3z2-r2 | -16.713 |
|---|
| x2-y2 | -0.071 |
|---|
| xy | -0.260 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.883 |
0.076 |
0.000 |
| y |
0.076 |
8.674 |
0.000 |
| z |
0.000 |
0.000 |
9.128 |
<r2> (average value of r
2) Å
2
| <r2> |
154.394 |
| (<r2>)1/2 |
12.426 |