return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H15N (triethylamine)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C3 1A
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-291.536942
Energy at 298.15K-291.554228
HF Energy-290.391944
Nuclear repulsion energy335.010764
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3152 3024 45.46      
2 A 3140 3012 5.91      
3 A 3112 2986 0.94      
4 A 3060 2936 7.42      
5 A 2969 2848 185.95      
6 A 1517 1455 8.32      
7 A 1492 1431 0.34      
8 A 1485 1425 2.47      
9 A 1437 1378 14.57      
10 A 1404 1347 0.00      
11 A 1324 1270 5.63      
12 A 1182 1134 9.55      
13 A 1098 1053 0.79      
14 A 1056 1013 8.37      
15 A 821 787 0.53      
16 A 766 735 10.59      
17 A 452 434 3.59      
18 A 308 295 0.76      
19 A 200 191 0.29      
20 A 102 98 0.02      
21 E 3154 3026 36.00      
22 E 3143 3015 46.99      
23 E 3110 2984 23.24      
24 E 3060 2936 29.45      
25 E 2959 2838 12.21      
26 E 1511 1450 2.39      
26 E 1511 1450 2.39      
27 E 1499 1438 0.99      
27 E 1499 1438 0.99      
28 E 1486 1426 2.02      
28 E 1486 1426 2.02      
29 E 1432 1374 26.25      
30 E 1397 1340 6.64      
30 E 1397 1340 6.64      
31 E 1335 1281 24.52      
32 E 1255 1204 16.31      
33 E 1121 1076 16.10      
34 E 1105 1060 4.95      
34 E 1105 1060 4.95      
35 E 945 907 1.91      
35 E 945 907 1.91      
36 E 805 772 1.96      
36 E 805 772 1.96      
37 E 478 459 1.78      
37 E 478 459 1.78      
38 E 320 307 0.46      
38 E 320 307 0.46      
39 E 220 211 0.04      
39 E 220 211 0.04      
40 E 95 91 0.18      
40 E 95 91 0.18      

Unscaled Zero Point Vibrational Energy (zpe) 45465.0 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 43619.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.07656 0.07656 0.04407

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is C3

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.000 0.000 0.005
C2 0.328 1.360 -0.451
C3 1.013 -0.964 -0.451
C4 -1.341 -0.396 -0.451
C5 -0.299 2.438 0.438
C6 2.261 -0.961 0.438
C7 -1.962 -1.478 0.438
H8 1.425 1.476 -0.417
H9 0.030 1.511 -1.516
H10 0.566 -1.973 -0.417
H11 1.293 -0.781 -1.516
H12 -1.991 0.496 -0.417
H13 -1.323 -0.729 -1.516
H14 -1.402 2.395 0.426
H15 0.036 2.306 1.482
H16 0.000 3.445 0.092
H17 2.775 0.017 0.426
H18 1.979 -1.184 1.482
H19 2.984 -1.723 0.092
H20 -1.373 -2.412 0.426
H21 -2.015 -1.122 1.482
H22 -2.984 -1.723 0.092

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 C6 C7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18 H19 H20 H21 H22
N11.47091.47091.47092.49442.49442.49442.09512.14412.09512.14412.09512.14412.80712.73883.44652.80712.73883.44652.80712.73883.4465
C21.47092.42252.42251.53163.14773.75311.10411.11683.34092.57902.47512.86822.19902.17162.18022.92583.59604.10464.22933.92214.5567
C31.47092.42252.42253.75311.53163.14772.47512.86821.10411.11683.34092.57904.22933.92214.55672.19902.17162.18022.92583.59604.1046
C41.47092.42252.42253.14773.75311.53163.34092.57902.47512.86821.10411.11682.92583.59604.10464.22933.92214.55672.19902.17162.1802
C52.49441.53163.75313.14774.25484.25482.15132.18804.57554.08902.71423.86041.10471.10441.10573.91294.40465.31104.96784.08794.9642
C62.49443.14771.53163.75314.25484.25482.71423.86042.15132.18804.57554.08904.96784.08794.96421.10471.10441.10573.91294.40465.3110
C72.49443.75313.14771.53164.25484.25484.57554.08902.71423.86042.15132.18803.91294.40465.31104.96784.08794.96421.10471.10441.1057
H82.09511.10412.47513.34092.15132.71424.57551.77693.55462.51493.55463.69203.09052.49472.48362.15923.31523.59474.86424.71105.4713
H92.14411.11682.86822.57902.18803.86044.08901.77693.69202.61722.51492.61722.57063.10202.51643.67984.47784.66564.59674.48364.7039
H102.09513.34091.10412.47514.57552.15132.71423.55463.69201.77693.55462.51494.86424.71105.47133.09052.49472.48362.15923.31523.5947
H112.14412.57901.11682.86824.08902.18803.86042.51492.61721.77693.69202.61724.59674.48364.70392.57063.10202.51643.67984.47784.6656
H122.09512.47513.34091.10412.71424.57552.15133.55462.51493.55463.69201.77692.15923.31523.59474.86424.71105.47133.09052.49472.4836
H132.14412.86822.57901.11683.86044.08902.18803.69202.61722.51492.61721.77693.67984.47784.66564.59674.48364.70392.57063.10202.5164
H142.80712.19904.22932.92581.10474.96783.91293.09052.57064.86424.59672.15923.67981.78641.78374.80695.03576.02534.80693.72244.4234
H152.73882.17163.92213.59601.10444.08794.40462.49473.10204.71104.48363.31524.47781.78641.79713.72243.99465.18175.03573.99465.2230
H163.44652.18024.55674.10461.10574.96425.31102.48362.51645.47134.70393.59474.66561.78371.79714.42345.22305.96756.02535.18175.9675
H172.80712.92582.19904.22933.91291.10474.96782.15923.67983.09052.57064.86424.59674.80693.72244.42341.78641.78374.80695.03576.0253
H182.73883.59602.17163.92214.40461.10444.08793.31524.47782.49473.10204.71104.48365.03573.99465.22301.78641.79713.72243.99465.1817
H193.44654.10462.18024.55675.31101.10574.96423.59474.66562.48362.51645.47134.70396.02535.18175.96751.78371.79714.42345.22305.9675
H202.80714.22932.92582.19904.96783.91291.10474.86424.59672.15923.67983.09052.57064.80695.03576.02534.80693.72244.42341.78641.7837
H212.73883.92213.59602.17164.08794.40461.10444.71104.48363.31524.47782.49473.10203.72243.99465.18175.03573.99465.22301.78641.7971
H223.44654.55674.10462.18024.96425.31101.10575.47134.70393.59474.66562.48362.51644.42345.22305.96756.02535.18175.96751.78371.7971

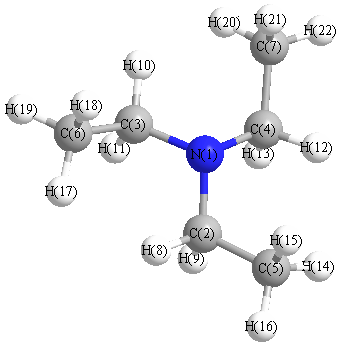
picture of triethylamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C5 112.340 N1 C2 H8 108.059
N1 C2 H9 111.164 N1 C3 C6 112.340
N1 C3 H10 108.059 N1 C3 H11 111.164
N1 C4 C7 112.340 N1 C4 H12 108.059
N1 C4 H13 111.164 C2 N1 C3 110.867
C2 N1 C4 110.867 C2 C5 H14 112.034
C2 C5 H15 109.881 C2 C5 H16 110.473
C3 N1 C4 110.867 C3 C6 H17 112.034
C3 C6 H18 109.881 C3 C6 H19 110.473
C4 C7 H20 112.034 C4 C7 H21 109.881
C4 C7 H22 110.473 C5 C2 H8 108.321
C5 C2 H9 110.427 C6 C3 H10 108.321
C6 C3 H11 110.427 C7 C4 H12 108.321
C7 C4 H13 110.427 H8 C2 H9 106.273
H10 C3 H11 106.273 H12 C4 H13 106.273
H14 C5 H15 107.934 H14 C5 H16 107.604
H15 C5 H16 108.814 H17 C6 H18 107.933
H17 C6 H19 107.604 H18 C6 H19 108.814
H20 C7 H21 107.933 H20 C7 H22 107.604
H21 C7 H22 108.814
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability