return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H15N (triethylamine)

using model chemistry: CCD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C3 1A
Energy calculated at CCD/cc-pVDZ
 hartrees
Energy at 0K-291.529930
Energy at 298.15K-291.547237
HF Energy-290.392155
Nuclear repulsion energy335.128104
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3164 3027 42.82      
2 A 3153 3016 5.16      
3 A 3123 2987 1.05      
4 A 3070 2937 7.38      
5 A 2982 2853 176.54      
6 A 1520 1454 8.89      
7 A 1494 1429 0.29      
8 A 1488 1424 2.43      
9 A 1441 1379 13.82      
10 A 1408 1347 0.03      
11 A 1328 1270 5.54      
12 A 1185 1134 10.00      
13 A 1100 1053 0.88      
14 A 1060 1014 8.31      
15 A 823 787 0.54      
16 A 769 736 10.11      
17 A 454 434 3.70      
18 A 309 295 0.75      
19 A 200 191 0.35      
20 A 103 99 0.02      
21 E 3166 3029 32.99      
22 E 3155 3019 45.35      
23 E 3121 2986 23.41      
24 E 3070 2937 27.11      
25 E 2972 2844 10.87      
26 E 1514 1449 2.67      
26 E 1514 1449 2.67      
27 E 1501 1436 0.96      
27 E 1501 1436 0.96      
28 E 1489 1424 2.15      
28 E 1489 1424 2.15      
29 E 1436 1374 27.11      
30 E 1401 1340 6.02      
30 E 1401 1340 6.02      
31 E 1341 1282 23.34      
32 E 1259 1204 16.48      
33 E 1125 1077 16.02      
34 E 1108 1060 3.94      
34 E 1108 1060 3.94      
35 E 948 907 1.68      
35 E 948 907 1.68      
36 E 807 772 2.01      
36 E 807 772 2.01      
37 E 480 459 1.81      
37 E 480 459 1.81      
38 E 321 307 0.48      
38 E 321 307 0.48      
39 E 221 211 0.04      
39 E 221 211 0.04      
40 E 95 91 0.18      
40 E 95 91 0.18      

Unscaled Zero Point Vibrational Energy (zpe) 45615.4 cm-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 43640.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/cc-pVDZ
ABC
0.07660 0.07660 0.04410

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/cc-pVDZ

Point Group is C3

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.000 0.000 0.003
C2 0.328 1.359 -0.451
C3 1.013 -0.963 -0.451
C4 -1.341 -0.396 -0.451
C5 -0.298 2.438 0.438
C6 2.260 -0.961 0.438
C7 -1.962 -1.477 0.438
H8 1.424 1.476 -0.418
H9 0.029 1.511 -1.516
H10 0.566 -1.972 -0.418
H11 1.294 -0.781 -1.516
H12 -1.991 0.495 -0.418
H13 -1.323 -0.730 -1.516
H14 -1.401 2.394 0.429
H15 0.038 2.306 1.481
H16 0.000 3.444 0.093
H17 2.774 0.016 0.429
H18 1.978 -1.186 1.481
H19 2.983 -1.722 0.093
H20 -1.373 -2.411 0.429
H21 -2.016 -1.120 1.481
H22 -2.983 -1.722 0.093

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 C6 C7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18 H19 H20 H21 H22
N11.46981.46981.46982.49402.49402.49402.09432.14282.09432.14282.09432.14282.80652.73913.44542.80652.73913.44542.80652.73913.4454
C21.46982.42102.42101.53163.14743.75161.10371.11623.33932.57792.47412.86742.19872.17132.17982.92603.59634.10344.22773.92054.5548
C31.46982.42102.42103.75161.53163.14742.47412.86741.10371.11623.33932.57794.22773.92054.55482.19872.17132.17982.92603.59634.1034
C41.46982.42102.42103.14743.75161.53163.33932.57792.47412.86741.10371.11622.92603.59634.10344.22773.92054.55482.19872.17132.1798
C52.49401.53163.75163.14744.25364.25362.15032.18724.57424.08782.71493.86051.10421.10391.10513.91154.40445.30914.96604.08614.9630
C62.49403.14741.53163.75164.25364.25362.71493.86052.15032.18724.57424.08784.96604.08614.96301.10421.10391.10513.91154.40445.3091
C72.49403.75163.14741.53164.25364.25364.57424.08782.71493.86052.15032.18723.91154.40445.30914.96604.08614.96301.10421.10391.1051
H82.09431.10372.47413.33932.15032.71494.57421.77563.55332.51333.55333.69093.08932.49352.48242.16093.31683.59434.86314.71005.4694
H92.14281.11622.86742.57792.18723.86054.08781.77563.69092.61752.51332.61752.57043.10072.51503.68074.47804.66504.59624.48184.7023
H102.09433.33931.10372.47414.57422.15032.71493.55333.69091.77563.55332.51334.86314.71005.46943.08932.49352.48242.16093.31683.5943
H112.14282.57791.11622.86744.08782.18723.86052.51332.61751.77563.69092.61754.59624.48184.70232.57043.10072.51503.68074.47804.6650
H122.09432.47413.33931.10372.71494.57422.15033.55332.51333.55333.69091.77562.16093.31683.59434.86314.71005.46943.08932.49352.4824
H132.14282.86742.57791.11623.86054.08782.18723.69092.61752.51332.61751.77563.68074.47804.66504.59624.48184.70232.57043.10072.5150
H142.80652.19874.22772.92601.10424.96603.91153.08932.57044.86314.59622.16093.68071.78551.78294.80495.03436.02304.80493.71964.4225
H152.73912.17133.92053.59631.10394.08614.40442.49353.10074.71004.48183.31684.47801.78551.79613.71963.99445.17925.03433.99445.2230
H163.44542.17984.55484.10341.10514.96305.30912.48242.51505.46944.70233.59434.66501.78291.79614.42255.22305.96566.02305.17925.9656
H172.80652.92602.19874.22773.91151.10424.96602.16093.68073.08932.57044.86314.59624.80493.71964.42251.78551.78294.80495.03436.0230
H182.73913.59632.17133.92054.40441.10394.08613.31684.47802.49353.10074.71004.48185.03433.99445.22301.78551.79613.71963.99445.1792
H193.44544.10342.17984.55485.30911.10514.96303.59434.66502.48242.51505.46944.70236.02305.17925.96561.78291.79614.42255.22305.9656
H202.80654.22772.92602.19874.96603.91151.10424.86314.59622.16093.68073.08932.57044.80495.03436.02304.80493.71964.42251.78551.7829
H212.73913.92053.59632.17134.08614.40441.10394.71004.48183.31684.47802.49353.10073.71963.99445.17925.03433.99445.22301.78551.7961
H223.44544.55484.10342.17984.96305.30911.10515.46944.70233.59434.66502.48242.51504.42255.22305.96566.02305.17925.96561.78291.7961

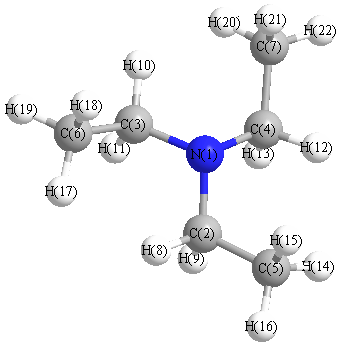
picture of triethylamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C5 112.380 N1 C2 H8 108.096
N1 C2 H9 111.181 N1 C3 C6 112.380
N1 C3 H10 108.096 N1 C3 H11 111.181
N1 C4 C7 112.380 N1 C4 H12 108.096
N1 C4 H13 111.181 C2 N1 C3 110.894
C2 N1 C4 110.894 C2 C5 H14 112.044
C2 C5 H15 109.887 C2 C5 H16 110.482
C3 N1 C4 110.894 C3 C6 H17 112.044
C3 C6 H18 109.887 C3 C6 H19 110.482
C4 C7 H20 112.044 C4 C7 H21 109.887
C4 C7 H22 110.482 C5 C2 H8 108.275
C5 C2 H9 110.410 C6 C3 H10 108.275
C6 C3 H11 110.410 C7 C4 H12 108.275
C7 C4 H13 110.410 H8 C2 H9 106.237
H10 C3 H11 106.237 H12 C4 H13 106.237
H14 C5 H15 107.923 H14 C5 H16 107.608
H15 C5 H16 108.795 H17 C6 H18 107.923
H17 C6 H19 107.608 H18 C6 H19 108.795
H20 C7 H21 107.922 H20 C7 H22 107.608
H21 C7 H22 108.795
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability