return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C7H8 (Norbornadiene)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-270.674580
Energy at 298.15K-270.683956
Nuclear repulsion energy285.838021
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3258 3126 7.08      
2 A1 3152 3024 9.70      
3 A1 3083 2958 39.52      
4 A1 1658 1590 0.19      
5 A1 1497 1436 0.42      
6 A1 1277 1225 1.78      
7 A1 1130 1084 0.20      
8 A1 956 918 2.53      
9 A1 912 875 6.35      
10 A1 794 762 1.37      
11 A1 749 719 48.32      
12 A1 428 411 4.69      
13 A2 3229 3098 0.00      
14 A2 1325 1271 0.00      
15 A2 1268 1216 0.00      
16 A2 1138 1091 0.00      
17 A2 934 896 0.00      
18 A2 906 869 0.00      
19 A2 718 689 0.00      
20 A2 445 427 0.00      
21 B1 3256 3124 18.23      
22 B1 3149 3021 34.48      
23 B1 1621 1555 4.70      
24 B1 1258 1207 0.39      
25 B1 1084 1040 0.36      
26 B1 1038 996 0.23      
27 B1 941 903 1.90      
28 B1 675 648 28.82      
29 B1 511 490 4.69      
30 B2 3231 3100 9.92      
31 B2 3148 3020 57.44      
32 B2 1355 1300 18.60      
33 B2 1281 1229 0.01      
34 B2 1202 1153 0.42      
35 B2 976 936 0.05      
36 B2 925 887 2.23      
37 B2 897 860 2.14      
38 B2 821 788 6.98      
39 B2 554 531 1.35      

Unscaled Zero Point Vibrational Energy (zpe) 28389.5 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 27236.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.14126 0.11907 0.10488

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.357
C2 0.000 1.127 0.277
C3 0.000 -1.127 0.277
C4 1.247 0.676 -0.523
C5 1.247 -0.676 -0.523
C6 -1.247 0.676 -0.523
C7 -1.247 -0.676 -0.523
H8 -0.910 0.000 1.983
H9 0.910 0.000 1.983
H10 0.000 2.172 0.624
H11 0.000 -2.172 0.624
H12 1.943 1.348 -1.033
H13 1.943 -1.348 -1.033
H14 -1.943 1.348 -1.033
H15 -1.943 -1.348 -1.033

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 H8 H9 H10 H11 H12 H13 H14 H15
C11.56161.56162.35522.35522.35522.35521.10401.10402.29222.29223.36243.36243.36243.3624
C21.56162.25481.54862.33351.54862.33352.23822.23821.10053.31732.35373.40842.35373.4084
C31.56162.25482.33351.54862.33351.54862.23822.23823.31731.10053.40842.35373.40842.3537
C42.35521.54862.33351.35152.49332.83603.37442.61752.26003.31331.09392.19993.29943.8117
C52.35522.33351.54861.35152.83602.49333.37442.61753.31332.26002.19991.09393.81173.2994
C62.35521.54862.33352.49332.83601.35152.61753.37442.26003.31333.29943.81171.09392.1999
C72.35522.33351.54862.83602.49331.35152.61753.37443.31332.26003.81173.29942.19991.0939
H81.10402.23822.23823.37443.37442.61752.61751.81902.71862.71864.36474.36473.46143.4614
H91.10402.23822.23822.61752.61753.37443.37441.81902.71862.71863.46143.46144.36474.3647
H102.29221.10053.31732.26003.31332.26003.31332.71862.71864.34342.68324.34832.68324.3483
H112.29223.31731.10053.31332.26003.31332.26002.71862.71864.34344.34832.68324.34832.6832
H123.36242.35373.40841.09392.19993.29943.81174.36473.46142.68324.34832.69553.88624.7295
H133.36243.40842.35372.19991.09393.81173.29944.36473.46144.34832.68322.69554.72953.8862
H143.36242.35373.40843.29943.81171.09392.19993.46144.36472.68324.34833.88624.72952.6955
H153.36243.40842.35373.81173.29942.19991.09393.46144.36474.34832.68324.72953.88622.6955

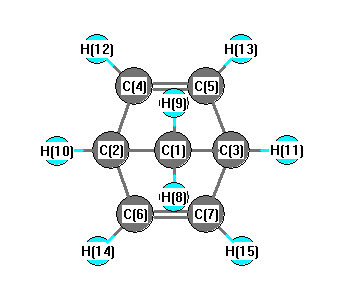
picture of Norbornadiene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 98.447 C1 C2 C6 98.447
C1 C2 H10 117.831 C1 C3 C5 98.447
C1 C3 C7 98.447 C1 C3 H11 117.831
C2 C1 C3 92.432 C2 C1 H8 113.091
C2 C1 H9 113.091 C2 C4 C5 106.957
C2 C4 H12 125.039 C2 C6 C7 106.957
C2 C6 H14 125.039 C3 C1 H8 113.091
C3 C1 H9 113.091 C3 C5 C4 106.957
C3 C5 H13 125.039 C3 C7 C6 106.957
C3 C7 H15 125.039 C4 C2 C6 107.227
C4 C2 H10 116.084 C4 C5 H13 127.904
C5 C3 C7 107.227 C5 C3 H11 116.084
C5 C4 H12 127.904 C6 C2 H10 116.084
C6 C7 H15 127.904 C7 C3 H11 116.084
C7 C6 H14 127.904 H8 C1 H9 110.945
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability