Jump to
S1C2
S1C3
Energy calculated at BLYP/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -209.137596 |
| Energy at 298.15K | -209.143553 |
| HF Energy | -209.137596 |
| Nuclear repulsion energy | 117.775920 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at BLYP/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3461 |
3466 |
9.90 |
|
|
|
| 2 |
A' |
3034 |
3038 |
14.79 |
|
|
|
| 3 |
A' |
2931 |
2936 |
64.69 |
|
|
|
| 4 |
A' |
2792 |
2796 |
115.39 |
|
|
|
| 5 |
A' |
1742 |
1745 |
371.52 |
|
|
|
| 6 |
A' |
1466 |
1468 |
14.25 |
|
|
|
| 7 |
A' |
1417 |
1420 |
4.60 |
|
|
|
| 8 |
A' |
1402 |
1404 |
6.58 |
|
|
|
| 9 |
A' |
1335 |
1337 |
8.40 |
|
|
|
| 10 |
A' |
1249 |
1251 |
89.67 |
|
|
|
| 11 |
A' |
1115 |
1117 |
30.55 |
|
|
|
| 12 |
A' |
969 |
970 |
41.29 |
|
|
|
| 13 |
A' |
588 |
589 |
12.43 |
|
|
|
| 14 |
A' |
328 |
329 |
7.22 |
|
|
|
| 15 |
A" |
2977 |
2982 |
36.73 |
|
|
|
| 16 |
A" |
1413 |
1415 |
4.65 |
|
|
|
| 17 |
A" |
1101 |
1102 |
0.09 |
|
|
|
| 18 |
A" |
985 |
987 |
1.37 |
|
|
|
| 19 |
A" |
600 |
601 |
88.94 |
|
|
|
| 20 |
A" |
204 |
205 |
0.96 |
|
|
|
| 21 |
A" |
95 |
95 |
0.62 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15601.3 cm
-1
Scaled (by 1.0016) Zero Point Vibrational Energy (zpe) 15626.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at BLYP/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.283 |
-0.772 |
0.000 |
| O2 |
1.412 |
-1.255 |
0.000 |
| N3 |
0.000 |
0.575 |
0.000 |
| C4 |
-1.338 |
1.157 |
0.000 |
| H5 |
-0.663 |
-1.388 |
0.000 |
| H6 |
0.815 |
1.196 |
0.000 |
| H7 |
-2.082 |
0.335 |
0.000 |
| H8 |
-1.519 |
1.783 |
0.901 |
| H9 |
-1.519 |
1.783 |
-0.901 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2284 | 1.3767 | 2.5198 | 1.1290 | 2.0390 | 2.6112 | 3.2535 | 3.2535 |
O2 | 1.2284 | | 2.3120 | 3.6585 | 2.0799 | 2.5232 | 3.8391 | 4.3166 | 4.3166 | N3 | 1.3767 | 2.3120 | | 1.4592 | 2.0721 | 1.0248 | 2.0954 | 2.1392 | 2.1392 | C4 | 2.5198 | 3.6585 | 1.4592 | | 2.6328 | 2.1538 | 1.1078 | 1.1115 | 1.1115 | H5 | 1.1290 | 2.0799 | 2.0721 | 2.6328 | | 2.9771 | 2.2320 | 3.4053 | 3.4053 | H6 | 2.0390 | 2.5232 | 1.0248 | 2.1538 | 2.9771 | | 3.0219 | 2.5696 | 2.5696 | H7 | 2.6112 | 3.8391 | 2.0954 | 1.1078 | 2.2320 | 3.0219 | | 1.7951 | 1.7951 | H8 | 3.2535 | 4.3166 | 2.1392 | 1.1115 | 3.4053 | 2.5696 | 1.7951 | | 1.8011 | H9 | 3.2535 | 4.3166 | 2.1392 | 1.1115 | 3.4053 | 2.5696 | 1.7951 | 1.8011 | |
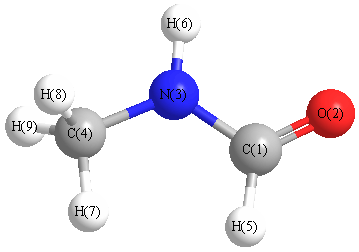 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
125.355 |
|
C1 |
N3 |
H6 |
115.442 |
| O2 |
C1 |
N3 |
125.022 |
|
O2 |
C1 |
H5 |
123.786 |
| N3 |
C1 |
H5 |
111.192 |
|
N3 |
C4 |
H7 |
108.654 |
| N3 |
C4 |
H8 |
111.932 |
|
N3 |
C4 |
H9 |
111.932 |
| C4 |
N3 |
H6 |
119.203 |
|
H7 |
C4 |
H8 |
107.968 |
| H7 |
C4 |
H9 |
107.968 |
|
H8 |
C4 |
H9 |
108.239 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.131 |
|
|
|
| 2 |
O |
-0.230 |
|
|
|
| 3 |
N |
-0.084 |
|
|
|
| 4 |
C |
0.070 |
|
|
|
| 5 |
H |
-0.038 |
|
|
|
| 6 |
H |
0.071 |
|
|
|
| 7 |
H |
0.029 |
|
|
|
| 8 |
H |
0.025 |
|
|
|
| 9 |
H |
0.025 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.619 |
2.699 |
0.000 |
3.761 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.080 |
3.518 |
0.000 |
| y |
3.518 |
-24.409 |
0.000 |
| z |
0.000 |
0.000 |
-24.584 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.583 |
3.518 |
0.000 |
| y |
3.518 |
0.923 |
0.000 |
| z |
0.000 |
0.000 |
0.660 |
|
| Polar |
|---|
| 3z2-r2 | 1.320 |
|---|
| x2-y2 | -1.670 |
|---|
| xy | 3.518 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.991 |
-1.089 |
0.000 |
| y |
-1.089 |
6.122 |
0.000 |
| z |
0.000 |
0.000 |
3.202 |
<r2> (average value of r
2) Å
2
| <r2> |
90.844 |
| (<r2>)1/2 |
9.531 |
Jump to
S1C1
S1C3
Energy calculated at BLYP/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -209.137596 |
| Energy at 298.15K | -209.143553 |
| HF Energy | -209.137596 |
| Nuclear repulsion energy | 117.775920 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at BLYP/cc-pVDZ
Geometric Data calculated at BLYP/cc-pVDZ
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at BLYP/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -209.138936 |
| Energy at 298.15K | -209.144926 |
| HF Energy | -209.138936 |
| Nuclear repulsion energy | 119.903989 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at BLYP/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3486 |
3492 |
8.30 |
|
|
|
| 2 |
A |
3068 |
3073 |
1.41 |
|
|
|
| 3 |
A |
2991 |
2996 |
35.66 |
|
|
|
| 4 |
A |
2940 |
2945 |
52.49 |
|
|
|
| 5 |
A |
2810 |
2814 |
150.32 |
|
|
|
| 6 |
A |
1728 |
1731 |
265.78 |
|
|
|
| 7 |
A |
1491 |
1494 |
61.34 |
|
|
|
| 8 |
A |
1431 |
1433 |
5.44 |
|
|
|
| 9 |
A |
1421 |
1423 |
32.59 |
|
|
|
| 10 |
A |
1374 |
1376 |
18.53 |
|
|
|
| 11 |
A |
1361 |
1363 |
0.95 |
|
|
|
| 12 |
A |
1166 |
1168 |
81.08 |
|
|
|
| 13 |
A |
1111 |
1113 |
20.76 |
|
|
|
| 14 |
A |
1105 |
1107 |
0.37 |
|
|
|
| 15 |
A |
962 |
964 |
0.00 |
|
|
|
| 16 |
A |
925 |
927 |
14.68 |
|
|
|
| 17 |
A |
739 |
740 |
0.07 |
|
|
|
| 18 |
A |
523 |
524 |
33.24 |
|
|
|
| 19 |
A |
288 |
289 |
12.13 |
|
|
|
| 20 |
A |
242 |
242 |
54.03 |
|
|
|
| 21 |
A |
108 |
108 |
0.47 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15634.7 cm
-1
Scaled (by 1.0016) Zero Point Vibrational Energy (zpe) 15659.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at BLYP/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.875 |
0.440 |
0.005 |
| O2 |
1.397 |
-0.678 |
-0.001 |
| N3 |
-0.478 |
0.662 |
-0.021 |
| C4 |
-1.443 |
-0.444 |
0.005 |
| H5 |
1.464 |
1.398 |
0.018 |
| H6 |
-0.813 |
1.624 |
0.057 |
| H7 |
-2.453 |
-0.053 |
-0.219 |
| H8 |
-1.167 |
-1.199 |
-0.757 |
| H9 |
-1.461 |
-0.952 |
0.992 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2340 | 1.3714 | 2.4807 | 1.1246 | 2.0625 | 3.3715 | 2.7268 | 2.8929 |
O2 | 1.2340 | | 2.3050 | 2.8496 | 2.0773 | 3.1917 | 3.9064 | 2.7235 | 3.0377 | N3 | 1.3714 | 2.3050 | | 1.4679 | 2.0771 | 1.0212 | 2.1095 | 2.1164 | 2.1442 | C4 | 2.4807 | 2.8496 | 1.4679 | | 3.4412 | 2.1619 | 1.1056 | 1.1070 | 1.1105 | H5 | 1.1246 | 2.0773 | 2.0771 | 3.4412 | | 2.2886 | 4.1835 | 3.7768 | 3.8764 | H6 | 2.0625 | 3.1917 | 1.0212 | 2.1619 | 2.2886 | | 2.3612 | 2.9585 | 2.8159 | H7 | 3.3715 | 3.9064 | 2.1095 | 1.1056 | 4.1835 | 2.3612 | | 1.8042 | 1.8048 | H8 | 2.7268 | 2.7235 | 2.1164 | 1.1070 | 3.7768 | 2.9585 | 1.8042 | | 1.7900 | H9 | 2.8929 | 3.0377 | 2.1442 | 1.1105 | 3.8764 | 2.8159 | 1.8048 | 1.7900 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
121.739 |
|
C1 |
N3 |
H6 |
118.354 |
| O2 |
C1 |
N3 |
124.342 |
|
O2 |
C1 |
H5 |
123.390 |
| N3 |
C1 |
H5 |
112.263 |
|
N3 |
C4 |
H7 |
109.302 |
| N3 |
C4 |
H8 |
109.766 |
|
N3 |
C4 |
H9 |
111.773 |
| C4 |
N3 |
H6 |
119.502 |
|
H7 |
C4 |
H8 |
109.260 |
| H7 |
C4 |
H9 |
109.049 |
|
H8 |
C4 |
H9 |
107.646 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.150 |
|
|
|
| 2 |
O |
-0.235 |
|
|
|
| 3 |
N |
-0.098 |
|
|
|
| 4 |
C |
0.068 |
|
|
|
| 5 |
H |
-0.037 |
|
|
|
| 6 |
H |
0.068 |
|
|
|
| 7 |
H |
0.022 |
|
|
|
| 8 |
H |
0.041 |
|
|
|
| 9 |
H |
0.022 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.783 |
2.307 |
0.003 |
3.615 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.486 |
2.000 |
-0.002 |
| y |
2.000 |
-22.341 |
0.004 |
| z |
-0.002 |
0.004 |
-24.582 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.025 |
2.000 |
-0.002 |
| y |
2.000 |
2.693 |
0.004 |
| z |
-0.002 |
0.004 |
-0.668 |
|
| Polar |
|---|
| 3z2-r2 | -1.337 |
|---|
| x2-y2 | -3.145 |
|---|
| xy | 2.000 |
|---|
| xz | -0.002 |
|---|
| yz | 0.004 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.516 |
0.069 |
0.000 |
| y |
0.069 |
5.403 |
0.001 |
| z |
0.000 |
0.001 |
3.189 |
<r2> (average value of r
2) Å
2
| <r2> |
80.121 |
| (<r2>)1/2 |
8.951 |
