Jump to
S1C2
S1C3
Energy calculated at B3PW91/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -209.134690 |
| Energy at 298.15K | -209.140745 |
| HF Energy | -209.134690 |
| Nuclear repulsion energy | 119.144222 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3606 |
3480 |
24.49 |
|
|
|
| 2 |
A' |
3139 |
3029 |
12.53 |
|
|
|
| 3 |
A' |
3028 |
2922 |
57.27 |
|
|
|
| 4 |
A' |
2919 |
2817 |
99.57 |
|
|
|
| 5 |
A' |
1838 |
1773 |
462.30 |
|
|
|
| 6 |
A' |
1519 |
1466 |
24.68 |
|
|
|
| 7 |
A' |
1473 |
1421 |
9.11 |
|
|
|
| 8 |
A' |
1441 |
1391 |
4.01 |
|
|
|
| 9 |
A' |
1375 |
1327 |
9.36 |
|
|
|
| 10 |
A' |
1305 |
1259 |
98.64 |
|
|
|
| 11 |
A' |
1173 |
1132 |
20.58 |
|
|
|
| 12 |
A' |
1011 |
976 |
44.52 |
|
|
|
| 13 |
A' |
615 |
593 |
14.59 |
|
|
|
| 14 |
A' |
336 |
325 |
8.00 |
|
|
|
| 15 |
A" |
3093 |
2985 |
28.47 |
|
|
|
| 16 |
A" |
1453 |
1402 |
6.38 |
|
|
|
| 17 |
A" |
1135 |
1095 |
0.10 |
|
|
|
| 18 |
A" |
1037 |
1000 |
0.56 |
|
|
|
| 19 |
A" |
623 |
601 |
108.16 |
|
|
|
| 20 |
A" |
210 |
203 |
0.79 |
|
|
|
| 21 |
A" |
100 |
97 |
0.68 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16214.6 cm
-1
Scaled (by 0.965) Zero Point Vibrational Energy (zpe) 15647.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.282 |
-0.760 |
0.000 |
| O2 |
1.399 |
-1.235 |
0.000 |
| N3 |
0.000 |
0.571 |
0.000 |
| C4 |
-1.326 |
1.135 |
0.000 |
| H5 |
-0.651 |
-1.378 |
0.000 |
| H6 |
0.805 |
1.189 |
0.000 |
| H7 |
-2.060 |
0.315 |
0.000 |
| H8 |
-1.511 |
1.754 |
0.894 |
| H9 |
-1.511 |
1.754 |
-0.894 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2139 | 1.3599 | 2.4850 | 1.1193 | 2.0177 | 2.5765 | 3.2141 | 3.2141 |
O2 | 1.2139 | | 2.2844 | 3.6115 | 2.0549 | 2.4959 | 3.7902 | 4.2662 | 4.2662 | N3 | 1.3599 | 2.2844 | | 1.4410 | 2.0544 | 1.0153 | 2.0753 | 2.1170 | 2.1170 | C4 | 2.4850 | 3.6115 | 1.4410 | | 2.6018 | 2.1320 | 1.1000 | 1.1030 | 1.1030 | H5 | 1.1193 | 2.0549 | 2.0544 | 2.6018 | | 2.9512 | 2.2024 | 3.3681 | 3.3681 | H6 | 2.0177 | 2.4959 | 1.0153 | 2.1320 | 2.9512 | | 2.9952 | 2.5465 | 2.5465 | H7 | 2.5765 | 3.7902 | 2.0753 | 1.1000 | 2.2024 | 2.9952 | | 1.7801 | 1.7801 | H8 | 3.2141 | 4.2662 | 2.1170 | 1.1030 | 3.3681 | 2.5465 | 1.7801 | | 1.7886 | H9 | 3.2141 | 4.2662 | 2.1170 | 1.1030 | 3.3681 | 2.5465 | 1.7801 | 1.7886 | |
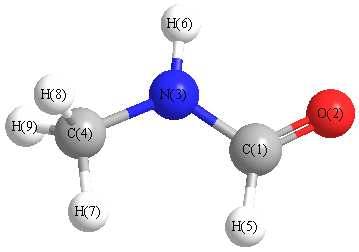 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
125.022 |
|
C1 |
N3 |
H6 |
115.540 |
| O2 |
C1 |
N3 |
125.039 |
|
O2 |
C1 |
H5 |
123.405 |
| N3 |
C1 |
H5 |
111.556 |
|
N3 |
C4 |
H7 |
108.778 |
| N3 |
C4 |
H8 |
111.964 |
|
N3 |
C4 |
H9 |
111.964 |
| C4 |
N3 |
H6 |
119.438 |
|
H7 |
C4 |
H8 |
107.809 |
| H7 |
C4 |
H9 |
107.809 |
|
H8 |
C4 |
H9 |
108.351 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.182 |
|
|
|
| 2 |
O |
-0.279 |
|
|
|
| 3 |
N |
-0.188 |
|
|
|
| 4 |
C |
0.007 |
|
|
|
| 5 |
H |
0.000 |
|
|
|
| 6 |
H |
0.105 |
|
|
|
| 7 |
H |
0.058 |
|
|
|
| 8 |
H |
0.057 |
|
|
|
| 9 |
H |
0.057 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.822 |
2.732 |
0.000 |
3.928 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.876 |
3.786 |
0.000 |
| y |
3.786 |
-24.049 |
0.000 |
| z |
0.000 |
0.000 |
-24.353 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.675 |
3.786 |
0.000 |
| y |
3.786 |
1.066 |
0.000 |
| z |
0.000 |
0.000 |
0.609 |
|
| Polar |
|---|
| 3z2-r2 | 1.219 |
|---|
| x2-y2 | -1.828 |
|---|
| xy | 3.786 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.647 |
-1.023 |
0.000 |
| y |
-1.023 |
5.693 |
0.000 |
| z |
0.000 |
0.000 |
3.125 |
<r2> (average value of r
2) Å
2
| <r2> |
88.865 |
| (<r2>)1/2 |
9.427 |
Jump to
S1C1
S1C3
Energy calculated at B3PW91/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -209.134690 |
| Energy at 298.15K | -209.140745 |
| HF Energy | -209.134690 |
| Nuclear repulsion energy | 119.144222 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/cc-pVDZ
Geometric Data calculated at B3PW91/cc-pVDZ
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at B3PW91/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -209.136286 |
| Energy at 298.15K | -209.142388 |
| HF Energy | -209.136286 |
| Nuclear repulsion energy | 121.338047 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3641 |
3513 |
24.93 |
|
|
|
| 2 |
A |
3172 |
3061 |
0.29 |
|
|
|
| 3 |
A |
3106 |
2997 |
28.32 |
|
|
|
| 4 |
A |
3037 |
2931 |
46.78 |
|
|
|
| 5 |
A |
2942 |
2839 |
127.84 |
|
|
|
| 6 |
A |
1822 |
1758 |
329.45 |
|
|
|
| 7 |
A |
1555 |
1500 |
98.28 |
|
|
|
| 8 |
A |
1471 |
1419 |
7.66 |
|
|
|
| 9 |
A |
1467 |
1416 |
24.77 |
|
|
|
| 10 |
A |
1413 |
1364 |
19.84 |
|
|
|
| 11 |
A |
1404 |
1355 |
0.74 |
|
|
|
| 12 |
A |
1228 |
1185 |
69.38 |
|
|
|
| 13 |
A |
1147 |
1107 |
20.41 |
|
|
|
| 14 |
A |
1139 |
1099 |
0.34 |
|
|
|
| 15 |
A |
1017 |
981 |
0.11 |
|
|
|
| 16 |
A |
981 |
947 |
21.28 |
|
|
|
| 17 |
A |
767 |
740 |
0.50 |
|
|
|
| 18 |
A |
538 |
519 |
38.80 |
|
|
|
| 19 |
A |
298 |
288 |
13.67 |
|
|
|
| 20 |
A |
263 |
254 |
66.13 |
|
|
|
| 21 |
A |
110 |
106 |
0.20 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16258.5 cm
-1
Scaled (by 0.965) Zero Point Vibrational Energy (zpe) 15689.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.865 |
0.435 |
0.005 |
| O2 |
1.374 |
-0.672 |
-0.000 |
| N3 |
-0.472 |
0.658 |
-0.021 |
| C4 |
-1.420 |
-0.440 |
0.005 |
| H5 |
1.457 |
1.379 |
0.019 |
| H6 |
-0.805 |
1.610 |
0.057 |
| H7 |
-2.423 |
-0.055 |
-0.221 |
| H8 |
-1.139 |
-1.188 |
-0.751 |
| H9 |
-1.437 |
-0.940 |
0.988 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2193 | 1.3553 | 2.4469 | 1.1145 | 2.0419 | 3.3321 | 2.6874 | 2.8559 |
O2 | 1.2193 | | 2.2751 | 2.8035 | 2.0536 | 3.1557 | 3.8532 | 2.6728 | 2.9915 | N3 | 1.3553 | 2.2751 | | 1.4508 | 2.0602 | 1.0118 | 2.0871 | 2.0940 | 2.1213 | C4 | 2.4469 | 2.8035 | 1.4508 | | 3.4044 | 2.1411 | 1.0981 | 1.0995 | 1.1025 | H5 | 1.1145 | 2.0536 | 2.0602 | 3.4044 | | 2.2740 | 4.1442 | 3.7318 | 3.8333 | H6 | 2.0419 | 3.1557 | 1.0118 | 2.1411 | 2.2740 | | 2.3388 | 2.9314 | 2.7869 | H7 | 3.3321 | 3.8532 | 2.0871 | 1.0981 | 4.1442 | 2.3388 | | 1.7922 | 1.7935 | H8 | 2.6874 | 2.6728 | 2.0940 | 1.0995 | 3.7318 | 2.9314 | 1.7922 | | 1.7809 | H9 | 2.8559 | 2.9915 | 2.1213 | 1.1025 | 3.8333 | 2.7869 | 1.7935 | 1.7809 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
121.344 |
|
C1 |
N3 |
H6 |
118.504 |
| O2 |
C1 |
N3 |
124.099 |
|
O2 |
C1 |
H5 |
123.206 |
| N3 |
C1 |
H5 |
112.692 |
|
N3 |
C4 |
H7 |
109.151 |
| N3 |
C4 |
H8 |
109.616 |
|
N3 |
C4 |
H9 |
111.635 |
| C4 |
N3 |
H6 |
119.734 |
|
H7 |
C4 |
H8 |
109.280 |
| H7 |
C4 |
H9 |
109.174 |
|
H8 |
C4 |
H9 |
107.949 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.193 |
|
|
|
| 2 |
O |
-0.284 |
|
|
|
| 3 |
N |
-0.199 |
|
|
|
| 4 |
C |
-0.000 |
|
|
|
| 5 |
H |
0.006 |
|
|
|
| 6 |
H |
0.100 |
|
|
|
| 7 |
H |
0.054 |
|
|
|
| 8 |
H |
0.077 |
|
|
|
| 9 |
H |
0.054 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.818 |
2.486 |
-0.000 |
3.758 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.190 |
2.294 |
0.000 |
| y |
2.294 |
-21.841 |
-0.000 |
| z |
0.000 |
-0.000 |
-24.354 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.092 |
2.294 |
0.000 |
| y |
2.294 |
2.931 |
-0.000 |
| z |
0.000 |
-0.000 |
-0.839 |
|
| Polar |
|---|
| 3z2-r2 | -1.677 |
|---|
| x2-y2 | -3.348 |
|---|
| xy | 2.294 |
|---|
| xz | 0.000 |
|---|
| yz | -0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.067 |
0.019 |
0.000 |
| y |
0.019 |
5.050 |
0.000 |
| z |
0.000 |
0.000 |
3.115 |
<r2> (average value of r
2) Å
2
| <r2> |
78.253 |
| (<r2>)1/2 |
8.846 |
