Jump to
S1C2
S1C3
Energy calculated at MP3=FULL/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -208.632703 |
| Energy at 298.15K | -208.638635 |
| HF Energy | -207.977168 |
| Nuclear repulsion energy | 119.099459 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3=FULL/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3695 |
3474 |
32.73 |
|
|
|
| 2 |
A' |
3189 |
2999 |
17.15 |
|
|
|
| 3 |
A' |
3084 |
2900 |
54.97 |
|
|
|
| 4 |
A' |
3014 |
2834 |
83.91 |
|
|
|
| 5 |
A' |
1898 |
1785 |
514.57 |
|
|
|
| 6 |
A' |
1559 |
1466 |
32.02 |
|
|
|
| 7 |
A' |
1519 |
1428 |
14.97 |
|
|
|
| 8 |
A' |
1483 |
1395 |
6.38 |
|
|
|
| 9 |
A' |
1422 |
1337 |
9.23 |
|
|
|
| 10 |
A' |
1329 |
1250 |
134.17 |
|
|
|
| 11 |
A' |
1200 |
1129 |
22.44 |
|
|
|
| 12 |
A' |
1031 |
970 |
44.85 |
|
|
|
| 13 |
A' |
627 |
590 |
14.12 |
|
|
|
| 14 |
A' |
348 |
327 |
8.05 |
|
|
|
| 15 |
A" |
3153 |
2965 |
32.60 |
|
|
|
| 16 |
A" |
1491 |
1402 |
4.88 |
|
|
|
| 17 |
A" |
1164 |
1095 |
0.93 |
|
|
|
| 18 |
A" |
1073 |
1009 |
0.11 |
|
|
|
| 19 |
A" |
591 |
556 |
119.68 |
|
|
|
| 20 |
A" |
198 |
186 |
1.14 |
|
|
|
| 21 |
A" |
31 |
29 |
2.27 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16549.9 cm
-1
Scaled (by 0.9403) Zero Point Vibrational Energy (zpe) 15561.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP3=FULL/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.291 |
-0.758 |
0.000 |
| O2 |
1.406 |
-1.226 |
0.000 |
| N3 |
0.000 |
0.574 |
0.000 |
| C4 |
-1.339 |
1.124 |
0.000 |
| H5 |
-0.634 |
-1.383 |
0.000 |
| H6 |
0.797 |
1.200 |
0.000 |
| H7 |
-2.061 |
0.293 |
0.000 |
| H8 |
-1.530 |
1.739 |
0.895 |
| H9 |
-1.530 |
1.739 |
-0.895 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2096 | 1.3634 | 2.4887 | 1.1160 | 2.0216 | 2.5759 | 3.2167 | 3.2167 |
O2 | 1.2096 | | 2.2841 | 3.6129 | 2.0461 | 2.5007 | 3.7853 | 4.2672 | 4.2672 | N3 | 1.3634 | 2.2841 | | 1.4471 | 2.0571 | 1.0127 | 2.0804 | 2.1208 | 2.1208 | C4 | 2.4887 | 3.6129 | 1.4471 | | 2.6033 | 2.1369 | 1.1008 | 1.1029 | 1.1029 | H5 | 1.1160 | 2.0461 | 2.0571 | 2.6033 | | 2.9521 | 2.2010 | 3.3683 | 3.3683 | H6 | 2.0216 | 2.5007 | 1.0127 | 2.1369 | 2.9521 | | 2.9983 | 2.5507 | 2.5507 | H7 | 2.5759 | 3.7853 | 2.0804 | 1.1008 | 2.2010 | 2.9983 | | 1.7814 | 1.7814 | H8 | 3.2167 | 4.2672 | 2.1208 | 1.1029 | 3.3683 | 2.5507 | 1.7814 | | 1.7907 | H9 | 3.2167 | 4.2672 | 2.1208 | 1.1029 | 3.3683 | 2.5507 | 1.7814 | 1.7907 | |
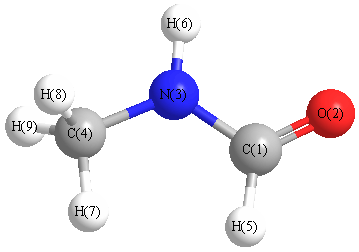 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
124.605 |
|
C1 |
N3 |
H6 |
115.819 |
| O2 |
C1 |
N3 |
125.071 |
|
O2 |
C1 |
H5 |
123.181 |
| N3 |
C1 |
H5 |
111.748 |
|
N3 |
C4 |
H7 |
108.718 |
| N3 |
C4 |
H8 |
111.838 |
|
N3 |
C4 |
H9 |
111.838 |
| C4 |
N3 |
H6 |
119.576 |
|
H7 |
C4 |
H8 |
107.876 |
| H7 |
C4 |
H9 |
107.876 |
|
H8 |
C4 |
H9 |
108.544 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at MP3=FULL/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -208.632703 |
| Energy at 298.15K | -208.638635 |
| HF Energy | -207.977168 |
| Nuclear repulsion energy | 119.099459 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3=FULL/cc-pVDZ
Geometric Data calculated at MP3=FULL/cc-pVDZ
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at MP3=FULL/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -208.634694 |
| Energy at 298.15K | -208.640629 |
| HF Energy | -207.978918 |
| Nuclear repulsion energy | 121.260186 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3=FULL/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3728 |
3506 |
36.97 |
|
|
|
| 2 |
A |
3227 |
3034 |
1.07 |
|
|
|
| 3 |
A |
3161 |
2972 |
33.27 |
|
|
|
| 4 |
A |
3091 |
2906 |
45.14 |
|
|
|
| 5 |
A |
3033 |
2852 |
116.08 |
|
|
|
| 6 |
A |
1884 |
1771 |
375.56 |
|
|
|
| 7 |
A |
1594 |
1499 |
129.91 |
|
|
|
| 8 |
A |
1511 |
1420 |
29.56 |
|
|
|
| 9 |
A |
1509 |
1419 |
6.04 |
|
|
|
| 10 |
A |
1460 |
1373 |
16.19 |
|
|
|
| 11 |
A |
1448 |
1362 |
2.94 |
|
|
|
| 12 |
A |
1256 |
1181 |
86.22 |
|
|
|
| 13 |
A |
1175 |
1104 |
14.17 |
|
|
|
| 14 |
A |
1168 |
1098 |
1.16 |
|
|
|
| 15 |
A |
1055 |
992 |
0.50 |
|
|
|
| 16 |
A |
999 |
939 |
25.07 |
|
|
|
| 17 |
A |
782 |
735 |
1.55 |
|
|
|
| 18 |
A |
515 |
484 |
32.28 |
|
|
|
| 19 |
A |
309 |
291 |
13.09 |
|
|
|
| 20 |
A |
212 |
200 |
82.74 |
|
|
|
| 21 |
A |
42 |
40 |
3.94 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16578.5 cm
-1
Scaled (by 0.9403) Zero Point Vibrational Energy (zpe) 15588.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP3=FULL/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.867 |
0.433 |
0.004 |
| O2 |
1.366 |
-0.674 |
-0.000 |
| N3 |
-0.470 |
0.663 |
-0.023 |
| C4 |
-1.418 |
-0.440 |
0.006 |
| H5 |
1.469 |
1.368 |
0.023 |
| H6 |
-0.799 |
1.613 |
0.065 |
| H7 |
-2.425 |
-0.053 |
-0.209 |
| H8 |
-1.146 |
-1.183 |
-0.759 |
| H9 |
-1.426 |
-0.947 |
0.986 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2146 | 1.3575 | 2.4464 | 1.1119 | 2.0430 | 3.3343 | 2.6917 | 2.8508 |
O2 | 1.2146 | | 2.2720 | 2.7939 | 2.0451 | 3.1506 | 3.8469 | 2.6726 | 2.9734 | N3 | 1.3575 | 2.2720 | | 1.4549 | 2.0636 | 1.0091 | 2.0897 | 2.0994 | 2.1268 | C4 | 2.4464 | 2.7939 | 1.4549 | | 3.4064 | 2.1456 | 1.0996 | 1.1006 | 1.1034 | H5 | 1.1119 | 2.0451 | 2.0636 | 3.4064 | | 2.2816 | 4.1510 | 3.7355 | 3.8295 | H6 | 2.0430 | 3.1506 | 1.0091 | 2.1456 | 2.2816 | | 2.3438 | 2.9358 | 2.7920 | H7 | 3.3343 | 3.8469 | 2.0897 | 1.0996 | 4.1510 | 2.3438 | | 1.7933 | 1.7954 | H8 | 2.6917 | 2.6726 | 2.0994 | 1.1006 | 3.7355 | 2.9358 | 1.7933 | | 1.7833 | H9 | 2.8508 | 2.9734 | 2.1268 | 1.1034 | 3.8295 | 2.7920 | 1.7954 | 1.7833 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
120.848 |
|
C1 |
N3 |
H6 |
118.630 |
| O2 |
C1 |
N3 |
124.005 |
|
O2 |
C1 |
H5 |
122.992 |
| N3 |
C1 |
H5 |
113.001 |
|
N3 |
C4 |
H7 |
108.988 |
| N3 |
C4 |
H8 |
109.696 |
|
N3 |
C4 |
H9 |
111.735 |
| C4 |
N3 |
H6 |
120.000 |
|
H7 |
C4 |
H8 |
109.194 |
| H7 |
C4 |
H9 |
109.173 |
|
H8 |
C4 |
H9 |
108.019 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
