Jump to
S1C2
S1C3
Energy calculated at B2PLYP=FULLultrafine/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -209.004155 |
| Energy at 298.15K | |
| HF Energy | -208.793289 |
| Nuclear repulsion energy | 118.859501 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3618 |
3467 |
26.04 |
89.71 |
0.23 |
0.37 |
| 2 |
A' |
3158 |
3026 |
15.02 |
51.15 |
0.74 |
0.85 |
| 3 |
A' |
3051 |
2923 |
57.78 |
180.77 |
0.03 |
0.05 |
| 4 |
A' |
2952 |
2829 |
98.46 |
97.13 |
0.32 |
0.48 |
| 5 |
A' |
1818 |
1742 |
429.11 |
8.11 |
0.50 |
0.67 |
| 6 |
A' |
1530 |
1466 |
17.86 |
12.18 |
0.75 |
0.86 |
| 7 |
A' |
1484 |
1422 |
11.52 |
16.16 |
0.40 |
0.57 |
| 8 |
A' |
1461 |
1400 |
5.50 |
3.38 |
0.75 |
0.86 |
| 9 |
A' |
1392 |
1334 |
10.85 |
7.73 |
0.64 |
0.78 |
| 10 |
A' |
1307 |
1252 |
104.58 |
5.26 |
0.39 |
0.56 |
| 11 |
A' |
1176 |
1127 |
26.41 |
4.17 |
0.72 |
0.84 |
| 12 |
A' |
1014 |
972 |
42.45 |
5.37 |
0.27 |
0.43 |
| 13 |
A' |
613 |
587 |
13.58 |
5.55 |
0.38 |
0.55 |
| 14 |
A' |
342 |
327 |
7.70 |
0.35 |
0.20 |
0.33 |
| 15 |
A" |
3115 |
2985 |
31.74 |
106.91 |
0.75 |
0.86 |
| 16 |
A" |
1475 |
1413 |
5.46 |
15.81 |
0.75 |
0.86 |
| 17 |
A" |
1149 |
1101 |
0.34 |
1.60 |
0.75 |
0.86 |
| 18 |
A" |
1042 |
999 |
0.56 |
2.13 |
0.75 |
0.86 |
| 19 |
A" |
610 |
584 |
107.66 |
1.15 |
0.75 |
0.86 |
| 20 |
A" |
204 |
196 |
1.00 |
3.37 |
0.75 |
0.86 |
| 21 |
A" |
81 |
78 |
1.20 |
0.14 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 16295.2 cm
-1
Scaled (by 0.9582) Zero Point Vibrational Energy (zpe) 15614.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULLultrafine/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.284 |
-0.761 |
0.000 |
| O2 |
1.404 |
-1.238 |
0.000 |
| N3 |
0.000 |
0.574 |
0.000 |
| C4 |
-1.333 |
1.136 |
0.000 |
| H5 |
-0.648 |
-1.377 |
0.000 |
| H6 |
0.805 |
1.192 |
0.000 |
| H7 |
-2.062 |
0.314 |
0.000 |
| H8 |
-1.518 |
1.753 |
0.894 |
| H9 |
-1.518 |
1.753 |
-0.894 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2179 | 1.3642 | 2.4923 | 1.1167 | 2.0212 | 2.5799 | 3.2192 | 3.2192 |
O2 | 1.2179 | | 2.2920 | 3.6232 | 2.0567 | 2.5029 | 3.7974 | 4.2757 | 4.2757 | N3 | 1.3642 | 2.2920 | | 1.4467 | 2.0550 | 1.0150 | 2.0781 | 2.1197 | 2.1197 | C4 | 2.4923 | 3.6232 | 1.4467 | | 2.6045 | 2.1383 | 1.0990 | 1.1017 | 1.1017 | H5 | 1.1167 | 2.0567 | 2.0550 | 2.6045 | | 2.9510 | 2.2037 | 3.3691 | 3.3691 | H6 | 2.0212 | 2.5029 | 1.0150 | 2.1383 | 2.9510 | | 2.9981 | 2.5509 | 2.5509 | H7 | 2.5799 | 3.7974 | 2.0781 | 1.0990 | 2.2037 | 2.9981 | | 1.7796 | 1.7796 | H8 | 3.2192 | 4.2757 | 2.1197 | 1.1017 | 3.3691 | 2.5509 | 1.7796 | | 1.7877 | H9 | 3.2192 | 4.2757 | 2.1197 | 1.1017 | 3.3691 | 2.5509 | 1.7796 | 1.7877 | |
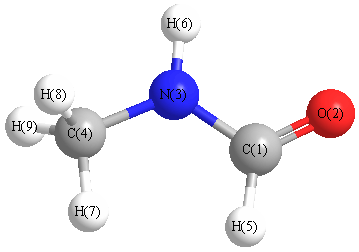 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
124.887 |
|
C1 |
N3 |
H6 |
115.541 |
| O2 |
C1 |
N3 |
125.067 |
|
O2 |
C1 |
H5 |
123.467 |
| N3 |
C1 |
H5 |
111.466 |
|
N3 |
C4 |
H7 |
108.667 |
| N3 |
C4 |
H8 |
111.856 |
|
N3 |
C4 |
H9 |
111.856 |
| C4 |
N3 |
H6 |
119.572 |
|
H7 |
C4 |
H8 |
107.930 |
| H7 |
C4 |
H9 |
107.930 |
|
H8 |
C4 |
H9 |
108.454 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP=FULLultrafine/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.192 |
|
|
|
| 2 |
O |
-0.269 |
|
|
|
| 3 |
N |
-0.188 |
|
|
|
| 4 |
C |
0.055 |
|
|
|
| 5 |
H |
-0.014 |
|
|
|
| 6 |
H |
0.097 |
|
|
|
| 7 |
H |
0.043 |
|
|
|
| 8 |
H |
0.042 |
|
|
|
| 9 |
H |
0.042 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.792 |
2.680 |
0.000 |
3.871 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.176 |
3.745 |
0.012 |
| y |
3.745 |
-24.353 |
-0.011 |
| z |
0.012 |
-0.011 |
-24.610 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.694 |
3.745 |
0.012 |
| y |
3.745 |
1.040 |
-0.011 |
| z |
0.012 |
-0.011 |
0.654 |
|
| Polar |
|---|
| 3z2-r2 | 1.309 |
|---|
| x2-y2 | -1.823 |
|---|
| xy | 3.745 |
|---|
| xz | 0.012 |
|---|
| yz | -0.011 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.645 |
-1.036 |
-0.001 |
| y |
-1.036 |
5.694 |
0.002 |
| z |
-0.001 |
0.002 |
3.105 |
<r2> (average value of r
2) Å
2
| <r2> |
89.423 |
| (<r2>)1/2 |
9.456 |
Jump to
S1C1
S1C3
Energy calculated at B2PLYP=FULLultrafine/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -209.004155 |
| Energy at 298.15K | |
| HF Energy | -208.793289 |
| Nuclear repulsion energy | 118.859501 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/cc-pVDZ
Geometric Data calculated at B2PLYP=FULLultrafine/cc-pVDZ
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at B2PLYP=FULLultrafine/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -209.005949 |
| Energy at 298.15K | |
| HF Energy | -208.794922 |
| Nuclear repulsion energy | 121.055114 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3651 |
3498 |
25.63 |
|
|
|
| 2 |
A |
3192 |
3059 |
0.59 |
|
|
|
| 3 |
A |
3125 |
2995 |
31.92 |
|
|
|
| 4 |
A |
3058 |
2930 |
48.05 |
|
|
|
| 5 |
A |
2972 |
2847 |
129.43 |
|
|
|
| 6 |
A |
1801 |
1726 |
304.25 |
|
|
|
| 7 |
A |
1561 |
1496 |
90.72 |
|
|
|
| 8 |
A |
1493 |
1431 |
6.49 |
|
|
|
| 9 |
A |
1488 |
1426 |
32.50 |
|
|
|
| 10 |
A |
1431 |
1372 |
18.99 |
|
|
|
| 11 |
A |
1417 |
1358 |
1.53 |
|
|
|
| 12 |
A |
1228 |
1176 |
82.03 |
|
|
|
| 13 |
A |
1158 |
1109 |
16.69 |
|
|
|
| 14 |
A |
1153 |
1105 |
0.67 |
|
|
|
| 15 |
A |
1022 |
979 |
0.15 |
|
|
|
| 16 |
A |
976 |
935 |
18.67 |
|
|
|
| 17 |
A |
767 |
735 |
0.49 |
|
|
|
| 18 |
A |
527 |
505 |
34.62 |
|
|
|
| 19 |
A |
302 |
290 |
13.05 |
|
|
|
| 20 |
A |
240 |
230 |
70.25 |
|
|
|
| 21 |
A |
93 |
89 |
1.13 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16327.7 cm
-1
Scaled (by 0.9582) Zero Point Vibrational Energy (zpe) 15645.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULLultrafine/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.866 |
0.438 |
0.004 |
| O2 |
1.376 |
-0.674 |
-0.000 |
| N3 |
-0.474 |
0.661 |
-0.022 |
| C4 |
-1.422 |
-0.443 |
0.005 |
| H5 |
1.457 |
1.381 |
0.021 |
| H6 |
-0.807 |
1.613 |
0.060 |
| H7 |
-2.426 |
-0.060 |
-0.217 |
| H8 |
-1.142 |
-1.188 |
-0.752 |
| H9 |
-1.434 |
-0.945 |
0.986 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2233 | 1.3591 | 2.4512 | 1.1131 | 2.0454 | 3.3363 | 2.6917 | 2.8570 |
O2 | 1.2233 | | 2.2818 | 2.8067 | 2.0568 | 3.1623 | 3.8567 | 2.6768 | 2.9896 | N3 | 1.3591 | 2.2818 | | 1.4551 | 2.0615 | 1.0115 | 2.0892 | 2.0972 | 2.1247 | C4 | 2.4512 | 2.8067 | 1.4551 | | 3.4077 | 2.1464 | 1.0975 | 1.0986 | 1.1017 | H5 | 1.1131 | 2.0568 | 2.0615 | 3.4077 | | 2.2764 | 4.1480 | 3.7347 | 3.8334 | H6 | 2.0454 | 3.1623 | 1.0115 | 2.1464 | 2.2764 | | 2.3437 | 2.9353 | 2.7912 | H7 | 3.3363 | 3.8567 | 2.0892 | 1.0975 | 4.1480 | 2.3437 | | 1.7914 | 1.7927 | H8 | 2.6917 | 2.6768 | 2.0972 | 1.0986 | 3.7347 | 2.9353 | 1.7914 | | 1.7792 | H9 | 2.8570 | 2.9896 | 2.1247 | 1.1017 | 3.8334 | 2.7912 | 1.7927 | 1.7792 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
121.118 |
|
C1 |
N3 |
H6 |
118.534 |
| O2 |
C1 |
N3 |
124.078 |
|
O2 |
C1 |
H5 |
123.299 |
| N3 |
C1 |
H5 |
112.620 |
|
N3 |
C4 |
H7 |
109.064 |
| N3 |
C4 |
H8 |
109.630 |
|
N3 |
C4 |
H9 |
111.665 |
| C4 |
N3 |
H6 |
119.889 |
|
H7 |
C4 |
H8 |
109.318 |
| H7 |
C4 |
H9 |
109.207 |
|
H8 |
C4 |
H9 |
107.924 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
