return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8O2 (Acetylacetone)

using model chemistry: CCD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A
Energy calculated at CCD/cc-pVDZ
 hartrees
Energy at 0K-344.844063
Energy at 298.15K-344.852629
HF Energy-343.753174
Nuclear repulsion energy295.289769
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3201 3062 6.40      
2 A 3157 3020 6.76      
3 A 3109 2974 3.98      
4 A 3073 2940 0.07      
5 A 1872 1791 16.94      
6 A 1482 1417 11.00      
7 A 1479 1415 11.61      
8 A 1466 1402 6.48      
9 A 1405 1344 8.30      
10 A 1291 1235 21.00      
11 A 1152 1102 1.06      
12 A 1087 1040 0.18      
13 A 958 916 1.01      
14 A 810 775 0.19      
15 A 633 606 1.47      
16 A 494 473 7.45      
17 A 321 307 0.58      
18 A 182 174 0.01      
19 A 149 143 0.84      
20 A 58 56 6.33      
21 B 3201 3062 7.16      
22 B 3183 3045 6.65      
23 B 3157 3020 1.58      
24 B 3073 2940 5.87      
25 B 1854 1774 248.06      
26 B 1479 1415 1.22      
27 B 1471 1407 19.61      
28 B 1405 1344 84.85      
29 B 1294 1238 121.45      
30 B 1215 1162 77.33      
31 B 1074 1027 1.55      
32 B 1007 963 0.51      
33 B 923 883 6.75      
34 B 830 794 3.41      
35 B 552 528 26.20      
36 B 504 482 3.48      
37 B 412 395 1.00      
38 B 181 173 0.20      
39 B 61 59 9.93      

Unscaled Zero Point Vibrational Energy (zpe) 27125.4 cm-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 25950.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/cc-pVDZ
ABC
0.13609 0.06517 0.05129

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/cc-pVDZ

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.016
C2 0.000 1.237 0.105
C3 0.000 -1.237 0.105
C4 -1.367 1.747 -0.312
C5 1.367 -1.747 -0.312
O6 1.040 1.727 -0.281
O7 -1.040 -1.727 -0.281
H8 -0.907 -0.029 1.641
H9 0.907 0.029 1.641
H10 -1.264 2.513 -1.095
H11 -1.984 0.903 -0.669
H12 -1.879 2.180 0.567
H13 1.264 -2.513 -1.095
H14 1.984 -0.903 -0.669
H15 1.879 -2.180 0.567

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 O6 O7 H8 H9 H10 H11 H12 H13 H14 H15
C11.53671.53672.58572.58572.39732.39731.10181.10183.51742.75582.91273.51742.75582.9127
C21.53672.47411.51763.30861.21283.16452.18812.15462.16002.15592.15244.13533.01953.9267
C31.53672.47413.30861.51763.16451.21282.15462.18814.13533.01953.92672.16002.15592.1524
C42.58571.51763.30864.43662.40793.48892.68013.45511.10051.10451.10545.06784.28775.1700
C52.58573.30861.51764.43663.48892.40793.45512.68015.06784.28775.17001.10051.10451.1054
O62.39731.21283.16452.40793.48894.03163.25142.56792.56723.15883.07364.32302.82104.0844
O72.39733.16451.21283.48892.40794.03162.56793.25144.32302.82104.08442.56723.15883.0736
H81.10182.18812.15462.68013.45513.25142.56791.81483.75252.71452.64194.28603.80283.6796
H91.10182.15462.18813.45512.68012.56793.25141.81484.28603.80283.67963.75252.71452.6419
H103.51742.16004.13531.10055.06782.56724.32303.75254.28601.81461.80355.62604.73335.8875
H112.75582.15593.01951.10454.28773.15882.82102.71453.80281.81461.77994.73334.36075.0949
H122.91272.15243.92671.10545.17003.07364.08442.64193.67961.80351.77995.88755.09495.7556
H133.51744.13532.16005.06781.10054.32302.56724.28603.75255.62604.73335.88751.81461.8035
H142.75583.01952.15594.28771.10452.82103.15883.80282.71454.73334.36075.09491.81461.7799
H152.91273.92672.15245.17001.10544.08443.07363.67962.64195.88755.09495.75561.80351.7799

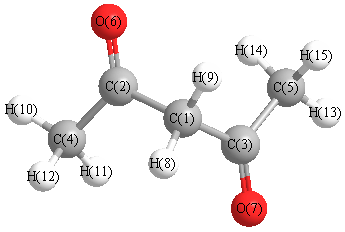
picture of Acetylacetone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 115.683 C1 C2 O6 120.906
C1 C3 C5 115.683 C1 C3 O7 120.906
C2 C1 C3 107.219 C2 C1 H8 110.978
C2 C1 H9 108.363 C2 C4 H10 110.159
C2 C4 H11 109.610 C2 C4 H12 109.278
C3 C1 H8 108.363 C3 C1 H9 110.978
C3 C5 H13 110.159 C3 C5 H14 109.610
C3 C5 H15 109.278 C4 C2 O6 123.360
C5 C3 O7 123.360 H8 C1 H9 110.888
H10 C4 H11 110.761 H10 C4 H12 109.676
H11 C4 H12 107.300 H13 C5 H14 110.761
H13 C5 H15 109.676 H14 C5 H15 107.300
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability