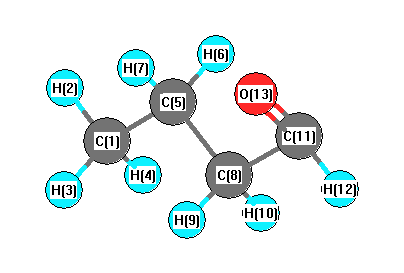
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3052 |
3034 |
28.03 |
|
|
|
| 2 |
A' |
2984 |
2967 |
17.17 |
|
|
|
| 3 |
A' |
2964 |
2947 |
27.25 |
|
|
|
| 4 |
A' |
2934 |
2917 |
22.36 |
|
|
|
| 5 |
A' |
2755 |
2739 |
184.21 |
|
|
|
| 6 |
A' |
1767 |
1757 |
143.71 |
|
|
|
| 7 |
A' |
1432 |
1423 |
5.12 |
|
|
|
| 8 |
A' |
1412 |
1404 |
2.61 |
|
|
|
| 9 |
A' |
1377 |
1369 |
5.88 |
|
|
|
| 10 |
A' |
1355 |
1347 |
6.04 |
|
|
|
| 11 |
A' |
1348 |
1341 |
5.03 |
|
|
|
| 12 |
A' |
1331 |
1323 |
28.01 |
|
|
|
| 13 |
A' |
1253 |
1246 |
4.32 |
|
|
|
| 14 |
A' |
1103 |
1097 |
9.36 |
|
|
|
| 15 |
A' |
1044 |
1038 |
0.20 |
|
|
|
| 16 |
A' |
939 |
933 |
7.80 |
|
|
|
| 17 |
A' |
841 |
836 |
10.66 |
|
|
|
| 18 |
A' |
680 |
676 |
11.74 |
|
|
|
| 19 |
A' |
337 |
335 |
0.62 |
|
|
|
| 20 |
A' |
189 |
187 |
7.56 |
|
|
|
| 21 |
A" |
3044 |
3026 |
45.32 |
|
|
|
| 22 |
A" |
3017 |
3000 |
1.85 |
|
|
|
| 23 |
A" |
2961 |
2944 |
7.77 |
|
|
|
| 24 |
A" |
1423 |
1415 |
6.53 |
|
|
|
| 25 |
A" |
1268 |
1260 |
0.22 |
|
|
|
| 26 |
A" |
1196 |
1189 |
0.15 |
|
|
|
| 27 |
A" |
1099 |
1093 |
0.05 |
|
|
|
| 28 |
A" |
913 |
907 |
0.00 |
|
|
|
| 29 |
A" |
760 |
755 |
2.43 |
|
|
|
| 30 |
A" |
643 |
639 |
3.93 |
|
|
|
| 31 |
A" |
254 |
253 |
0.05 |
|
|
|
| 32 |
A" |
190 |
189 |
0.58 |
|
|
|
| 33 |
A" |
74 |
73 |
1.29 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23968.2 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 23829.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.053 |
|
|
|
| 2 |
H |
0.032 |
|
|
|
| 3 |
H |
0.031 |
|
|
|
| 4 |
H |
0.031 |
|
|
|
| 5 |
C |
-0.066 |
|
|
|
| 6 |
H |
0.035 |
|
|
|
| 7 |
H |
0.035 |
|
|
|
| 8 |
C |
-0.084 |
|
|
|
| 9 |
H |
0.048 |
|
|
|
| 10 |
H |
0.048 |
|
|
|
| 11 |
C |
0.123 |
|
|
|
| 12 |
H |
-0.013 |
|
|
|
| 13 |
O |
-0.167 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.659 |
1.586 |
0.000 |
2.296 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.328 |
-2.943 |
0.000 |
| y |
-2.943 |
-31.025 |
0.000 |
| z |
0.000 |
0.000 |
-30.809 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.411 |
-2.943 |
0.000 |
| y |
-2.943 |
2.543 |
0.000 |
| z |
0.000 |
0.000 |
2.868 |
|
| Polar |
|---|
| 3z2-r2 | 5.736 |
|---|
| x2-y2 | -5.303 |
|---|
| xy | -2.943 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.498 |
-0.237 |
0.000 |
| y |
-0.237 |
7.605 |
0.000 |
| z |
0.000 |
0.000 |
5.751 |
<r2> (average value of r
2) Å
2
| <r2> |
156.622 |
| (<r2>)1/2 |
12.515 |