return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H9N (Pyrrolidine)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-211.956576
Energy at 298.15K-211.968128
HF Energy-211.160038
Nuclear repulsion energy189.426878
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3532 3388 0.80      
2 A' 3147 3019 61.00      
3 A' 3113 2986 8.71      
4 A' 3092 2967 43.43      
5 A' 2968 2848 135.41      
6 A' 1530 1468 0.76      
7 A' 1503 1442 3.73      
8 A' 1414 1357 3.61      
9 A' 1331 1277 0.88      
10 A' 1264 1212 2.10      
11 A' 1237 1187 8.58      
12 A' 1087 1043 0.46      
13 A' 1021 979 4.39      
14 A' 961 922 3.49      
15 A' 929 892 29.82      
16 A' 915 878 38.38      
17 A' 777 745 19.35      
18 A' 587 563 31.39      
19 A' 315 302 4.20      
20 A" 3128 3001 1.70      
21 A" 3107 2981 72.70      
22 A" 3080 2955 17.72      
23 A" 2964 2844 35.22      
24 A" 1513 1451 1.85      
25 A" 1479 1419 0.55      
26 A" 1450 1391 3.61      
27 A" 1338 1284 14.62      
28 A" 1314 1261 10.72      
29 A" 1252 1201 5.37      
30 A" 1212 1163 0.34      
31 A" 1153 1106 10.20      
32 A" 1115 1069 0.01      
33 A" 951 912 0.07      
34 A" 877 841 1.91      
35 A" 632 606 0.63      
36 A" 61 59 0.11      

Unscaled Zero Point Vibrational Energy (zpe) 28673.3 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 27509.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.22816 0.22493 0.12990

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.562 -1.090 0.000
H2 0.423 -2.103 0.000
C3 -0.107 -0.474 1.154
C4 -0.107 -0.474 -1.154
C5 -0.107 1.025 0.779
C6 -0.107 1.025 -0.779
H7 -1.157 -0.827 1.281
H8 -1.157 -0.827 -1.281
H9 0.439 -0.687 2.090
H10 0.439 -0.687 -2.090
H11 0.804 1.515 1.166
H12 0.804 1.515 -1.166
H13 -0.978 1.553 1.206
H14 -0.978 1.553 -1.206

Atom - Atom Distances (Å)
  N1 H2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14
N11.02311.46951.46952.35122.35122.15992.15992.13212.13212.86382.86383.28763.2876
H21.02312.06622.06623.26763.26762.40142.40142.52492.52493.82033.82034.09704.0970
C31.46952.06622.30901.54532.44691.11552.67601.10413.29722.18703.18872.20653.2307
C41.46952.06622.30902.44691.54532.67601.11553.29721.10413.18872.18703.23072.2065
C52.35123.26761.54532.44691.55902.18802.96352.22443.38601.10362.20311.10392.2312
C62.35123.26762.44691.54531.55902.96352.18803.38602.22442.20311.10362.23121.1039
H72.15992.40141.11552.67602.18802.96352.56251.79503.73293.05673.91402.38843.4474
H82.15992.40142.67601.11552.96352.18802.56253.73291.79503.91403.05673.44742.3884
H92.13212.52491.10413.29722.22443.38601.79503.73294.18032.41543.94762.79414.2295
H102.13212.52493.29721.10413.38602.22443.73291.79504.18033.94762.41544.22952.7941
H112.86383.82032.18703.18871.10362.20313.05673.91402.41543.94762.33211.78222.9666
H122.86383.82033.18872.18702.20311.10363.91403.05673.94762.41542.33212.96661.7822
H133.28764.09702.20653.23071.10392.23122.38843.44742.79414.22951.78222.96662.4118
H143.28764.09703.23072.20652.23121.10393.44742.38844.22952.79412.96661.78222.4118

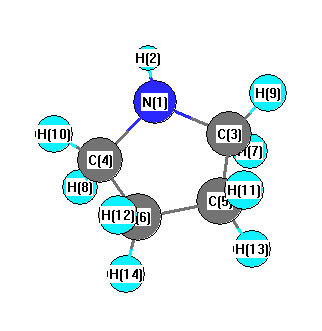
picture of Pyrrolidine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C3 C5 102.478 N1 C3 H7 112.627
N1 C3 H9 111.093 N1 C4 C6 102.478
N1 C4 H8 112.627 N1 C4 H10 111.093
H2 N1 C3 110.711 H2 N1 C4 110.711
C3 N1 C4 103.563 C3 C5 C6 104.045
C3 C5 H11 110.186 C3 C5 H13 111.718
C4 C6 C5 104.045 C4 C6 H12 110.186
C4 C6 H14 111.718 C5 C3 H7 109.579
C5 C3 H9 113.147 C5 C6 H12 110.503
C5 C6 H14 112.723 C6 C4 H8 109.579
C6 C4 H10 113.147 C6 C5 H11 110.503
C6 C5 H13 112.723 H7 C3 H9 107.940
H8 C4 H10 107.940 H11 C5 H13 107.675
H12 C6 H14 107.675
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability