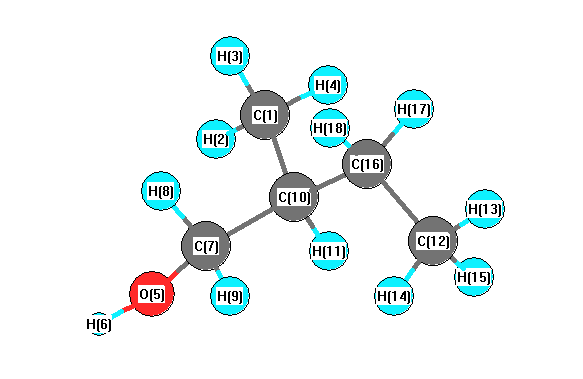
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3681 |
3659 |
17.57 |
|
|
|
| 2 |
A |
3074 |
3056 |
13.86 |
|
|
|
| 3 |
A |
3052 |
3034 |
36.58 |
|
|
|
| 4 |
A |
3046 |
3028 |
34.98 |
|
|
|
| 5 |
A |
3039 |
3021 |
28.17 |
|
|
|
| 6 |
A |
2989 |
2972 |
20.25 |
|
|
|
| 7 |
A |
2969 |
2952 |
31.08 |
|
|
|
| 8 |
A |
2963 |
2946 |
43.95 |
|
|
|
| 9 |
A |
2949 |
2931 |
50.03 |
|
|
|
| 10 |
A |
2939 |
2922 |
4.08 |
|
|
|
| 11 |
A |
2914 |
2897 |
53.94 |
|
|
|
| 12 |
A |
2866 |
2849 |
58.42 |
|
|
|
| 13 |
A |
1440 |
1432 |
2.25 |
|
|
|
| 14 |
A |
1433 |
1425 |
6.64 |
|
|
|
| 15 |
A |
1430 |
1421 |
3.51 |
|
|
|
| 16 |
A |
1422 |
1414 |
7.14 |
|
|
|
| 17 |
A |
1414 |
1406 |
5.38 |
|
|
|
| 18 |
A |
1405 |
1397 |
2.09 |
|
|
|
| 19 |
A |
1393 |
1385 |
3.58 |
|
|
|
| 20 |
A |
1349 |
1341 |
1.91 |
|
|
|
| 21 |
A |
1337 |
1329 |
6.23 |
|
|
|
| 22 |
A |
1328 |
1320 |
1.77 |
|
|
|
| 23 |
A |
1305 |
1297 |
3.36 |
|
|
|
| 24 |
A |
1272 |
1265 |
3.61 |
|
|
|
| 25 |
A |
1250 |
1242 |
13.92 |
|
|
|
| 26 |
A |
1217 |
1210 |
7.67 |
|
|
|
| 27 |
A |
1200 |
1193 |
13.25 |
|
|
|
| 28 |
A |
1146 |
1139 |
1.99 |
|
|
|
| 29 |
A |
1124 |
1118 |
2.36 |
|
|
|
| 30 |
A |
1097 |
1091 |
2.37 |
|
|
|
| 31 |
A |
1052 |
1045 |
71.46 |
|
|
|
| 32 |
A |
1032 |
1026 |
12.16 |
|
|
|
| 33 |
A |
988 |
982 |
19.57 |
|
|
|
| 34 |
A |
941 |
935 |
5.25 |
|
|
|
| 35 |
A |
897 |
892 |
2.21 |
|
|
|
| 36 |
A |
892 |
887 |
1.57 |
|
|
|
| 37 |
A |
808 |
804 |
3.96 |
|
|
|
| 38 |
A |
744 |
740 |
5.08 |
|
|
|
| 39 |
A |
481 |
478 |
5.94 |
|
|
|
| 40 |
A |
435 |
433 |
1.36 |
|
|
|
| 41 |
A |
376 |
374 |
1.70 |
|
|
|
| 42 |
A |
272 |
270 |
4.85 |
|
|
|
| 43 |
A |
261 |
260 |
14.23 |
|
|
|
| 44 |
A |
246 |
245 |
61.23 |
|
|
|
| 45 |
A |
227 |
225 |
5.92 |
|
|
|
| 46 |
A |
216 |
215 |
10.43 |
|
|
|
| 47 |
A |
117 |
116 |
3.21 |
|
|
|
| 48 |
A |
80 |
79 |
3.40 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35051.1 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 34847.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.006 |
|
|
|
| 2 |
H |
0.034 |
|
|
|
| 3 |
H |
0.019 |
|
|
|
| 4 |
H |
0.022 |
|
|
|
| 5 |
O |
-0.237 |
|
|
|
| 6 |
H |
0.137 |
|
|
|
| 7 |
C |
0.125 |
|
|
|
| 8 |
H |
0.003 |
|
|
|
| 9 |
H |
0.009 |
|
|
|
| 10 |
C |
-0.146 |
|
|
|
| 11 |
H |
0.006 |
|
|
|
| 12 |
C |
-0.049 |
|
|
|
| 13 |
H |
0.027 |
|
|
|
| 14 |
H |
0.032 |
|
|
|
| 15 |
H |
0.032 |
|
|
|
| 16 |
C |
-0.038 |
|
|
|
| 17 |
H |
0.016 |
|
|
|
| 18 |
H |
0.013 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.214 |
-0.938 |
0.916 |
1.329 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.432 |
3.086 |
-2.368 |
| y |
3.086 |
-37.935 |
-1.503 |
| z |
-2.368 |
-1.503 |
-40.204 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.363 |
3.086 |
-2.368 |
| y |
3.086 |
1.883 |
-1.503 |
| z |
-2.368 |
-1.503 |
-1.520 |
|
| Polar |
|---|
| 3z2-r2 | -3.040 |
|---|
| x2-y2 | -1.498 |
|---|
| xy | 3.086 |
|---|
| xz | -2.368 |
|---|
| yz | -1.503 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.447 |
-0.108 |
-0.062 |
| y |
-0.108 |
9.573 |
-0.130 |
| z |
-0.062 |
-0.130 |
8.359 |
<r2> (average value of r
2) Å
2
| <r2> |
217.668 |
| (<r2>)1/2 |
14.754 |