Jump to
S1C2
Energy calculated at mPW1PW91/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -1195.843154 |
| Energy at 298.15K | -1195.845528 |
| HF Energy | -1195.843154 |
| Nuclear repulsion energy | 194.099954 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
2671 |
2560 |
0.67 |
|
|
|
| 2 |
A |
869 |
833 |
0.05 |
|
|
|
| 3 |
A |
498 |
477 |
0.34 |
|
|
|
| 4 |
A |
324 |
310 |
27.26 |
|
|
|
| 5 |
A |
205 |
197 |
0.01 |
|
|
|
| 6 |
B |
2670 |
2558 |
3.11 |
|
|
|
| 7 |
B |
858 |
823 |
6.28 |
|
|
|
| 8 |
B |
481 |
461 |
30.37 |
|
|
|
| 9 |
B |
347 |
333 |
19.62 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4461.2 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 4275.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/cc-pVDZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
0.000 |
0.843 |
| S2 |
0.000 |
1.679 |
-0.386 |
| S3 |
0.000 |
-1.679 |
-0.386 |
| H4 |
-1.342 |
1.774 |
-0.576 |
| H5 |
1.342 |
-1.774 |
-0.576 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
H4 |
H5 |
| S1 | | 2.0810 | 2.0810 | 2.6384 | 2.6384 |
S2 | 2.0810 | | 3.3583 | 1.3587 | 3.7095 | S3 | 2.0810 | 3.3583 | | 3.7095 | 1.3587 | H4 | 2.6384 | 1.3587 | 3.7095 | | 4.4486 | H5 | 2.6384 | 3.7095 | 1.3587 | 4.4486 | |
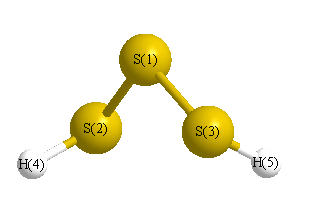 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
H4 |
97.972 |
|
S1 |
S3 |
H5 |
97.972 |
| S2 |
S1 |
S3 |
107.589 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
-0.064 |
|
|
|
| 2 |
S |
-0.090 |
|
|
|
| 3 |
S |
-0.090 |
|
|
|
| 4 |
H |
0.122 |
|
|
|
| 5 |
H |
0.122 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.639 |
0.639 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.914 |
-4.109 |
0.000 |
| y |
-4.109 |
-38.891 |
0.000 |
| z |
0.000 |
0.000 |
-41.032 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.048 |
-4.109 |
0.000 |
| y |
-4.109 |
0.082 |
0.000 |
| z |
0.000 |
0.000 |
-3.130 |
|
| Polar |
|---|
| 3z2-r2 | -6.260 |
|---|
| x2-y2 | 1.977 |
|---|
| xy | -4.109 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.546 |
-0.762 |
0.000 |
| y |
-0.762 |
10.869 |
0.000 |
| z |
0.000 |
0.000 |
5.698 |
<r2> (average value of r
2) Å
2
| <r2> |
141.253 |
| (<r2>)1/2 |
11.885 |
Jump to
S1C1
Energy calculated at mPW1PW91/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -1195.842777 |
| Energy at 298.15K | -1195.845127 |
| HF Energy | -1195.842777 |
| Nuclear repulsion energy | 194.122605 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
2660 |
2549 |
6.38 |
|
|
|
| 2 |
A' |
872 |
836 |
2.22 |
|
|
|
| 3 |
A' |
498 |
477 |
0.37 |
|
|
|
| 4 |
A' |
335 |
321 |
20.33 |
|
|
|
| 5 |
A' |
206 |
197 |
0.09 |
|
|
|
| 6 |
A" |
2662 |
2551 |
0.97 |
|
|
|
| 7 |
A" |
859 |
823 |
4.99 |
|
|
|
| 8 |
A" |
481 |
461 |
34.47 |
|
|
|
| 9 |
A" |
313 |
300 |
9.94 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4442.7 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 4257.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
-0.054 |
0.841 |
0.000 |
| S2 |
-0.054 |
-0.389 |
1.678 |
| S3 |
-0.054 |
-0.389 |
-1.678 |
| H4 |
1.292 |
-0.506 |
1.825 |
| H5 |
1.292 |
-0.506 |
-1.825 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
H4 |
H5 |
| S1 | | 2.0806 | 2.0806 | 2.6375 | 2.6375 |
S2 | 2.0806 | | 3.3561 | 1.3593 | 3.7543 | S3 | 2.0806 | 3.3561 | | 3.7543 | 1.3593 | H4 | 2.6375 | 1.3593 | 3.7543 | | 3.6491 | H5 | 2.6375 | 3.7543 | 1.3593 | 3.6491 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
H4 |
97.925 |
|
S1 |
S3 |
H5 |
97.925 |
| S2 |
S1 |
S3 |
107.517 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
-0.064 |
|
|
|
| 2 |
S |
-0.083 |
|
|
|
| 3 |
S |
-0.083 |
|
|
|
| 4 |
H |
0.115 |
|
|
|
| 5 |
H |
0.115 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.970 |
-0.525 |
0.000 |
2.039 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.170 |
-1.401 |
0.000 |
| y |
-1.401 |
-41.192 |
0.000 |
| z |
0.000 |
0.000 |
-38.592 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.722 |
-1.401 |
0.000 |
| y |
-1.401 |
-3.310 |
0.000 |
| z |
0.000 |
0.000 |
0.589 |
|
| Polar |
|---|
| 3z2-r2 | 1.177 |
|---|
| x2-y2 | 4.021 |
|---|
| xy | -1.401 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.523 |
-0.351 |
0.000 |
| y |
-0.351 |
5.661 |
0.000 |
| z |
0.000 |
0.000 |
10.912 |
<r2> (average value of r
2) Å
2
| <r2> |
141.265 |
| (<r2>)1/2 |
11.885 |
