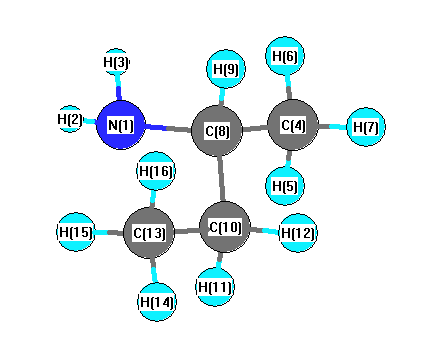
Vibrational Frequencies calculated at BLYP/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3396 |
3401 |
2.94 |
|
|
|
| 2 |
A |
3327 |
3333 |
7.14 |
|
|
|
| 3 |
A |
3027 |
3032 |
38.77 |
|
|
|
| 4 |
A |
3014 |
3018 |
43.28 |
|
|
|
| 5 |
A |
3008 |
3013 |
52.58 |
|
|
|
| 6 |
A |
3008 |
3012 |
26.79 |
|
|
|
| 7 |
A |
2973 |
2978 |
12.13 |
|
|
|
| 8 |
A |
2946 |
2951 |
27.82 |
|
|
|
| 9 |
A |
2942 |
2947 |
42.40 |
|
|
|
| 10 |
A |
2937 |
2942 |
21.73 |
|
|
|
| 11 |
A |
2803 |
2807 |
88.59 |
|
|
|
| 12 |
A |
1598 |
1601 |
22.92 |
|
|
|
| 13 |
A |
1441 |
1443 |
4.62 |
|
|
|
| 14 |
A |
1433 |
1435 |
1.70 |
|
|
|
| 15 |
A |
1430 |
1432 |
3.59 |
|
|
|
| 16 |
A |
1422 |
1424 |
2.70 |
|
|
|
| 17 |
A |
1407 |
1410 |
0.47 |
|
|
|
| 18 |
A |
1362 |
1365 |
4.51 |
|
|
|
| 19 |
A |
1353 |
1355 |
1.22 |
|
|
|
| 20 |
A |
1351 |
1354 |
5.63 |
|
|
|
| 21 |
A |
1326 |
1328 |
14.56 |
|
|
|
| 22 |
A |
1288 |
1290 |
8.26 |
|
|
|
| 23 |
A |
1254 |
1256 |
1.71 |
|
|
|
| 24 |
A |
1223 |
1225 |
0.17 |
|
|
|
| 25 |
A |
1149 |
1151 |
1.30 |
|
|
|
| 26 |
A |
1113 |
1115 |
7.44 |
|
|
|
| 27 |
A |
1025 |
1027 |
0.90 |
|
|
|
| 28 |
A |
995 |
997 |
2.87 |
|
|
|
| 29 |
A |
985 |
986 |
6.11 |
|
|
|
| 30 |
A |
951 |
952 |
2.04 |
|
|
|
| 31 |
A |
921 |
922 |
19.89 |
|
|
|
| 32 |
A |
848 |
849 |
83.52 |
|
|
|
| 33 |
A |
790 |
791 |
1.34 |
|
|
|
| 34 |
A |
745 |
746 |
2.37 |
|
|
|
| 35 |
A |
467 |
468 |
6.09 |
|
|
|
| 36 |
A |
440 |
440 |
4.26 |
|
|
|
| 37 |
A |
362 |
362 |
1.10 |
|
|
|
| 38 |
A |
276 |
277 |
27.39 |
|
|
|
| 39 |
A |
245 |
245 |
0.95 |
|
|
|
| 40 |
A |
224 |
225 |
0.82 |
|
|
|
| 41 |
A |
220 |
220 |
15.89 |
|
|
|
| 42 |
A |
92 |
93 |
2.34 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31557.6 cm
-1
Scaled (by 1.0016) Zero Point Vibrational Energy (zpe) 31608.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.117 |
|
|
|
| 2 |
H |
0.055 |
|
|
|
| 3 |
H |
0.055 |
|
|
|
| 4 |
C |
0.065 |
|
|
|
| 5 |
H |
0.007 |
|
|
|
| 6 |
H |
-0.010 |
|
|
|
| 7 |
H |
-0.003 |
|
|
|
| 8 |
C |
-0.050 |
|
|
|
| 9 |
H |
-0.043 |
|
|
|
| 10 |
C |
0.047 |
|
|
|
| 11 |
H |
-0.010 |
|
|
|
| 12 |
H |
-0.018 |
|
|
|
| 13 |
C |
0.011 |
|
|
|
| 14 |
H |
0.002 |
|
|
|
| 15 |
H |
0.009 |
|
|
|
| 16 |
H |
0.000 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.012 |
0.273 |
1.092 |
1.125 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.749 |
0.431 |
0.947 |
| y |
0.431 |
-32.691 |
1.875 |
| z |
0.947 |
1.875 |
-35.795 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.494 |
0.431 |
0.947 |
| y |
0.431 |
2.081 |
1.875 |
| z |
0.947 |
1.875 |
-2.575 |
|
| Polar |
|---|
| 3z2-r2 | -5.150 |
|---|
| x2-y2 | -1.057 |
|---|
| xy | 0.431 |
|---|
| xz | 0.947 |
|---|
| yz | 1.875 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.545 |
-0.031 |
0.038 |
| y |
-0.031 |
8.401 |
0.073 |
| z |
0.038 |
0.073 |
7.341 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |