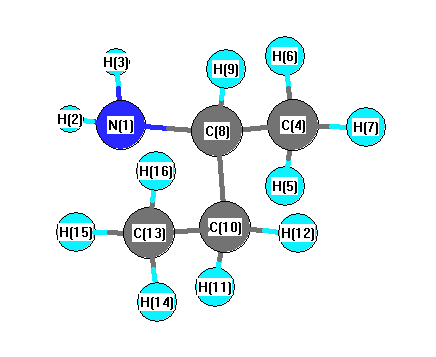
Vibrational Frequencies calculated at B3PW91/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3559 |
3435 |
0.07 |
|
|
|
| 2 |
A |
3482 |
3361 |
1.93 |
|
|
|
| 3 |
A |
3134 |
3024 |
29.34 |
|
|
|
| 4 |
A |
3122 |
3012 |
33.36 |
|
|
|
| 5 |
A |
3115 |
3006 |
29.21 |
|
|
|
| 6 |
A |
3115 |
3006 |
37.41 |
|
|
|
| 7 |
A |
3080 |
2972 |
11.46 |
|
|
|
| 8 |
A |
3038 |
2931 |
25.72 |
|
|
|
| 9 |
A |
3034 |
2928 |
39.39 |
|
|
|
| 10 |
A |
3033 |
2927 |
20.11 |
|
|
|
| 11 |
A |
2909 |
2808 |
82.58 |
|
|
|
| 12 |
A |
1639 |
1581 |
31.97 |
|
|
|
| 13 |
A |
1479 |
1427 |
6.62 |
|
|
|
| 14 |
A |
1470 |
1419 |
1.43 |
|
|
|
| 15 |
A |
1467 |
1416 |
7.39 |
|
|
|
| 16 |
A |
1459 |
1408 |
3.68 |
|
|
|
| 17 |
A |
1446 |
1395 |
0.82 |
|
|
|
| 18 |
A |
1416 |
1366 |
4.69 |
|
|
|
| 19 |
A |
1388 |
1339 |
13.69 |
|
|
|
| 20 |
A |
1387 |
1338 |
7.73 |
|
|
|
| 21 |
A |
1367 |
1319 |
7.14 |
|
|
|
| 22 |
A |
1319 |
1273 |
3.84 |
|
|
|
| 23 |
A |
1288 |
1243 |
1.56 |
|
|
|
| 24 |
A |
1255 |
1211 |
0.33 |
|
|
|
| 25 |
A |
1193 |
1151 |
2.19 |
|
|
|
| 26 |
A |
1156 |
1116 |
11.28 |
|
|
|
| 27 |
A |
1076 |
1039 |
0.52 |
|
|
|
| 28 |
A |
1038 |
1002 |
2.27 |
|
|
|
| 29 |
A |
1011 |
976 |
6.80 |
|
|
|
| 30 |
A |
974 |
939 |
3.33 |
|
|
|
| 31 |
A |
953 |
919 |
14.42 |
|
|
|
| 32 |
A |
881 |
850 |
96.62 |
|
|
|
| 33 |
A |
817 |
789 |
1.53 |
|
|
|
| 34 |
A |
769 |
742 |
0.84 |
|
|
|
| 35 |
A |
478 |
461 |
6.58 |
|
|
|
| 36 |
A |
452 |
436 |
5.10 |
|
|
|
| 37 |
A |
370 |
357 |
1.67 |
|
|
|
| 38 |
A |
283 |
273 |
29.04 |
|
|
|
| 39 |
A |
249 |
241 |
0.51 |
|
|
|
| 40 |
A |
231 |
223 |
3.82 |
|
|
|
| 41 |
A |
224 |
216 |
14.63 |
|
|
|
| 42 |
A |
99 |
95 |
2.17 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32627.1 cm
-1
Scaled (by 0.965) Zero Point Vibrational Energy (zpe) 31485.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.180 |
|
|
|
| 2 |
H |
0.081 |
|
|
|
| 3 |
H |
0.083 |
|
|
|
| 4 |
C |
-0.027 |
|
|
|
| 5 |
H |
0.045 |
|
|
|
| 6 |
H |
0.029 |
|
|
|
| 7 |
H |
0.035 |
|
|
|
| 8 |
C |
-0.119 |
|
|
|
| 9 |
H |
0.003 |
|
|
|
| 10 |
C |
-0.046 |
|
|
|
| 11 |
H |
0.034 |
|
|
|
| 12 |
H |
0.025 |
|
|
|
| 13 |
C |
-0.086 |
|
|
|
| 14 |
H |
0.042 |
|
|
|
| 15 |
H |
0.044 |
|
|
|
| 16 |
H |
0.037 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.019 |
0.251 |
1.141 |
1.169 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.043 |
0.434 |
0.977 |
| y |
0.434 |
-32.022 |
1.923 |
| z |
0.977 |
1.923 |
-35.210 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.573 |
0.434 |
0.977 |
| y |
0.434 |
2.104 |
1.923 |
| z |
0.977 |
1.923 |
-2.678 |
|
| Polar |
|---|
| 3z2-r2 | -5.356 |
|---|
| x2-y2 | -1.021 |
|---|
| xy | 0.434 |
|---|
| xz | 0.977 |
|---|
| yz | 1.923 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
150.883 |
| (<r2>)1/2 |
12.283 |