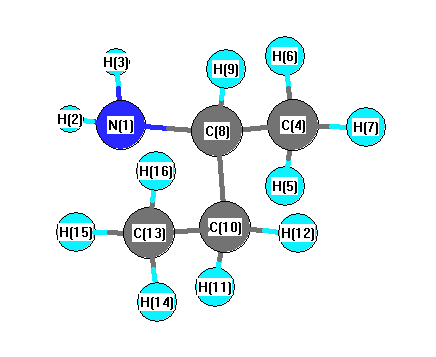
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3442 |
3422 |
0.91 |
|
|
|
| 2 |
A |
3366 |
3346 |
4.80 |
|
|
|
| 3 |
A |
3059 |
3041 |
27.21 |
|
|
|
| 4 |
A |
3047 |
3030 |
30.68 |
|
|
|
| 5 |
A |
3040 |
3023 |
26.58 |
|
|
|
| 6 |
A |
3039 |
3022 |
38.69 |
|
|
|
| 7 |
A |
3003 |
2985 |
11.53 |
|
|
|
| 8 |
A |
2964 |
2947 |
24.75 |
|
|
|
| 9 |
A |
2961 |
2943 |
41.37 |
|
|
|
| 10 |
A |
2957 |
2940 |
18.24 |
|
|
|
| 11 |
A |
2818 |
2801 |
86.13 |
|
|
|
| 12 |
A |
1587 |
1578 |
27.68 |
|
|
|
| 13 |
A |
1431 |
1423 |
6.12 |
|
|
|
| 14 |
A |
1422 |
1414 |
1.54 |
|
|
|
| 15 |
A |
1419 |
1411 |
6.26 |
|
|
|
| 16 |
A |
1411 |
1402 |
3.54 |
|
|
|
| 17 |
A |
1395 |
1387 |
1.03 |
|
|
|
| 18 |
A |
1365 |
1357 |
4.61 |
|
|
|
| 19 |
A |
1339 |
1331 |
4.55 |
|
|
|
| 20 |
A |
1338 |
1331 |
13.50 |
|
|
|
| 21 |
A |
1319 |
1311 |
8.30 |
|
|
|
| 22 |
A |
1274 |
1267 |
4.61 |
|
|
|
| 23 |
A |
1248 |
1240 |
1.44 |
|
|
|
| 24 |
A |
1217 |
1210 |
0.26 |
|
|
|
| 25 |
A |
1155 |
1149 |
1.37 |
|
|
|
| 26 |
A |
1116 |
1110 |
9.22 |
|
|
|
| 27 |
A |
1049 |
1043 |
0.24 |
|
|
|
| 28 |
A |
1011 |
1005 |
1.76 |
|
|
|
| 29 |
A |
980 |
974 |
6.48 |
|
|
|
| 30 |
A |
943 |
938 |
1.75 |
|
|
|
| 31 |
A |
932 |
926 |
19.31 |
|
|
|
| 32 |
A |
852 |
848 |
89.75 |
|
|
|
| 33 |
A |
795 |
791 |
0.60 |
|
|
|
| 34 |
A |
746 |
742 |
1.46 |
|
|
|
| 35 |
A |
466 |
464 |
6.11 |
|
|
|
| 36 |
A |
442 |
439 |
4.96 |
|
|
|
| 37 |
A |
361 |
359 |
1.62 |
|
|
|
| 38 |
A |
280 |
278 |
29.55 |
|
|
|
| 39 |
A |
246 |
244 |
0.11 |
|
|
|
| 40 |
A |
226 |
225 |
5.80 |
|
|
|
| 41 |
A |
221 |
220 |
10.90 |
|
|
|
| 42 |
A |
96 |
95 |
2.23 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31689.3 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 31505.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.141 |
|
|
|
| 2 |
H |
0.070 |
|
|
|
| 3 |
H |
0.071 |
|
|
|
| 4 |
C |
-0.002 |
|
|
|
| 5 |
H |
0.033 |
|
|
|
| 6 |
H |
0.018 |
|
|
|
| 7 |
H |
0.024 |
|
|
|
| 8 |
C |
-0.109 |
|
|
|
| 9 |
H |
-0.012 |
|
|
|
| 10 |
C |
-0.015 |
|
|
|
| 11 |
H |
0.020 |
|
|
|
| 12 |
H |
0.012 |
|
|
|
| 13 |
C |
-0.061 |
|
|
|
| 14 |
H |
0.030 |
|
|
|
| 15 |
H |
0.035 |
|
|
|
| 16 |
H |
0.027 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.012 |
0.288 |
1.102 |
1.139 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.395 |
0.453 |
0.937 |
| y |
0.453 |
-32.363 |
1.885 |
| z |
0.937 |
1.885 |
-35.575 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.574 |
0.453 |
0.937 |
| y |
0.453 |
2.122 |
1.885 |
| z |
0.937 |
1.885 |
-2.696 |
|
| Polar |
|---|
| 3z2-r2 | -5.392 |
|---|
| x2-y2 | -1.032 |
|---|
| xy | 0.453 |
|---|
| xz | 0.937 |
|---|
| yz | 1.885 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.497 |
-0.040 |
0.031 |
| y |
-0.040 |
8.344 |
0.068 |
| z |
0.031 |
0.068 |
7.328 |
<r2> (average value of r
2) Å
2
| <r2> |
152.120 |
| (<r2>)1/2 |
12.334 |