Jump to
S2C1
Energy calculated at mPW1PW91/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -489.216361 |
| Energy at 298.15K | |
| HF Energy | -489.216361 |
| Nuclear repulsion energy | 97.763468 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
843 |
808 |
111.45 |
4.56 |
0.45 |
0.62 |
| 2 |
A1 |
319 |
306 |
17.12 |
0.59 |
0.49 |
0.66 |
| 3 |
B2 |
862 |
826 |
132.46 |
5.32 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 1012.2 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 970.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/cc-pVDZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Si1 |
0.000 |
0.000 |
0.598 |
| F2 |
0.000 |
1.262 |
-0.465 |
| F3 |
0.000 |
-1.262 |
-0.465 |
Atom - Atom Distances (Å)
| |
Si1 |
F2 |
F3 |
| Si1 | | 1.6507 | 1.6507 |
F2 | 1.6507 | | 2.5246 | F3 | 1.6507 | 2.5246 | |
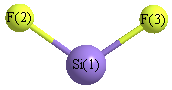 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| F2 |
Si1 |
F3 |
99.764 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
Si |
0.654 |
|
|
|
| 2 |
F |
-0.327 |
|
|
|
| 3 |
F |
-0.327 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
1.450 |
1.450 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-19.454 |
0.000 |
0.000 |
| y |
0.000 |
-24.875 |
0.000 |
| z |
0.000 |
0.000 |
-22.864 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.415 |
0.000 |
0.000 |
| y |
0.000 |
-3.716 |
0.000 |
| z |
0.000 |
0.000 |
-0.699 |
|
| Polar |
|---|
| 3z2-r2 | -1.397 |
|---|
| x2-y2 | 5.421 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.406 |
0.000 |
0.000 |
| y |
0.000 |
3.698 |
0.000 |
| z |
0.000 |
0.000 |
3.107 |
<r2> (average value of r
2) Å
2
| <r2> |
51.581 |
| (<r2>)1/2 |
7.182 |
Jump to
S1C1
Energy calculated at mPW1PW91/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -489.106464 |
| Energy at 298.15K | -488.986655 |
| HF Energy | -489.106464 |
| Nuclear repulsion energy | 96.211196 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at mPW1PW91/cc-pVDZ
Point Group is C2v
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
Si |
0.605 |
|
|
|
| 2 |
F |
-0.302 |
|
|
|
| 3 |
F |
-0.302 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
1.703 |
1.703 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.559 |
0.000 |
0.000 |
| y |
0.000 |
-25.119 |
0.000 |
| z |
0.000 |
0.000 |
-20.252 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.127 |
0.000 |
0.000 |
| y |
0.000 |
-3.714 |
0.000 |
| z |
0.000 |
0.000 |
3.587 |
|
| Polar |
|---|
| 3z2-r2 | 7.173 |
|---|
| x2-y2 | 2.560 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.713 |
0.000 |
0.000 |
| y |
0.000 |
3.399 |
0.000 |
| z |
0.000 |
0.000 |
2.776 |
<r2> (average value of r
2) Å
2
| <r2> |
55.202 |
| (<r2>)1/2 |
7.430 |
