Jump to
S1C2
S1C3
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
Geometric Data calculated at QCISD(T)/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
Geometric Data calculated at QCISD(T)/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at QCISD(T)/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -209.836080 |
| Energy at 298.15K | -209.845311 |
| HF Energy | -209.121188 |
| Nuclear repulsion energy | 134.146535 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3736 |
3561 |
|
|
|
|
| 2 |
A |
3562 |
3395 |
|
|
|
|
| 3 |
A |
3473 |
3310 |
|
|
|
|
| 4 |
A |
3097 |
2952 |
|
|
|
|
| 5 |
A |
3087 |
2943 |
|
|
|
|
| 6 |
A |
3001 |
2860 |
|
|
|
|
| 7 |
A |
2957 |
2819 |
|
|
|
|
| 8 |
A |
1647 |
1570 |
|
|
|
|
| 9 |
A |
1508 |
1437 |
|
|
|
|
| 10 |
A |
1490 |
1420 |
|
|
|
|
| 11 |
A |
1471 |
1402 |
|
|
|
|
| 12 |
A |
1415 |
1349 |
|
|
|
|
| 13 |
A |
1378 |
1313 |
|
|
|
|
| 14 |
A |
1330 |
1268 |
|
|
|
|
| 15 |
A |
1261 |
1202 |
|
|
|
|
| 16 |
A |
1194 |
1138 |
|
|
|
|
| 17 |
A |
1129 |
1076 |
|
|
|
|
| 18 |
A |
1075 |
1025 |
|
|
|
|
| 19 |
A |
1024 |
976 |
|
|
|
|
| 20 |
A |
967 |
922 |
|
|
|
|
| 21 |
A |
896 |
854 |
|
|
|
|
| 22 |
A |
852 |
812 |
|
|
|
|
| 23 |
A |
589 |
561 |
|
|
|
|
| 24 |
A |
539 |
513 |
|
|
|
|
| 25 |
A |
332 |
316 |
|
|
|
|
| 26 |
A |
280 |
267 |
|
|
|
|
| 27 |
A |
200 |
191 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21743.9 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 20724.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
-1.346 |
-0.577 |
0.118 |
| C2 |
-0.635 |
0.659 |
-0.276 |
| C3 |
0.793 |
0.557 |
0.272 |
| O4 |
1.400 |
-0.642 |
-0.173 |
| H5 |
-1.633 |
-0.488 |
1.100 |
| H6 |
-2.222 |
-0.648 |
-0.408 |
| H7 |
-1.120 |
1.601 |
0.066 |
| H8 |
-0.586 |
0.680 |
-1.381 |
| H9 |
1.406 |
1.407 |
-0.081 |
| H10 |
0.758 |
0.612 |
1.387 |
| H11 |
0.674 |
-1.287 |
-0.107 |
Atom - Atom Distances (Å)
| |
N1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| N1 | | 1.4789 | 2.4256 | 2.7629 | 1.0269 | 1.0244 | 2.1896 | 2.0992 | 3.3988 | 2.7296 | 2.1537 |
C2 | 1.4789 | | 1.5327 | 2.4180 | 2.0507 | 2.0602 | 1.1128 | 1.1069 | 2.1831 | 2.1698 | 2.3518 | C3 | 2.4256 | 1.5327 | | 1.4162 | 2.7683 | 3.3172 | 2.1882 | 2.1560 | 1.1064 | 1.1169 | 1.8869 | O4 | 2.7629 | 2.4180 | 1.4162 | | 3.2938 | 3.6302 | 3.3820 | 2.6744 | 2.0515 | 2.1026 | 0.9739 | H5 | 1.0269 | 2.0507 | 2.7683 | 3.2938 | | 1.6271 | 2.3869 | 2.9361 | 3.7719 | 2.6479 | 2.7243 | H6 | 1.0244 | 2.0602 | 3.3172 | 3.6302 | 1.6271 | | 2.5492 | 2.3217 | 4.1831 | 3.7004 | 2.9811 | H7 | 2.1896 | 1.1128 | 2.1882 | 3.3820 | 2.3869 | 2.5492 | | 1.7962 | 2.5376 | 2.4996 | 3.4043 | H8 | 2.0992 | 1.1069 | 2.1560 | 2.6744 | 2.9361 | 2.3217 | 1.7962 | | 2.4875 | 3.0779 | 2.6611 | H9 | 3.3988 | 2.1831 | 1.1064 | 2.0515 | 3.7719 | 4.1831 | 2.5376 | 2.4875 | | 1.7909 | 2.7926 | H10 | 2.7296 | 2.1698 | 1.1169 | 2.1026 | 2.6479 | 3.7004 | 2.4996 | 3.0779 | 1.7909 | | 2.4186 | H11 | 2.1537 | 2.3518 | 1.8869 | 0.9739 | 2.7243 | 2.9811 | 3.4043 | 2.6611 | 2.7926 | 2.4186 | |
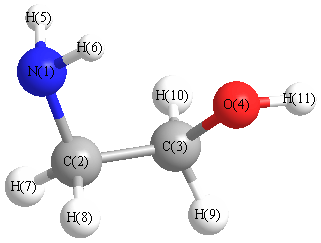 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C2 |
C3 |
107.289 |
|
N1 |
C2 |
H7 |
114.576 |
| N1 |
C2 |
H8 |
107.686 |
|
C2 |
N1 |
H5 |
108.501 |
| C2 |
N1 |
H6 |
109.434 |
|
C2 |
C3 |
O4 |
110.103 |
| C2 |
C3 |
H9 |
110.591 |
|
C2 |
C3 |
H10 |
108.943 |
| C3 |
C2 |
H7 |
110.613 |
|
C3 |
C2 |
H8 |
108.455 |
| C3 |
O4 |
H11 |
102.699 |
|
O4 |
C3 |
H9 |
108.200 |
| O4 |
C3 |
H10 |
111.658 |
|
H5 |
N1 |
H6 |
104.979 |
| H7 |
C2 |
H8 |
108.041 |
|
H9 |
C3 |
H10 |
107.314 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
