Jump to
S1C2
S1C3
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Geometric Data calculated at QCISD/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Geometric Data calculated at QCISD/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -209.818276 |
| Energy at 298.15K | |
| HF Energy | -209.121962 |
| Nuclear repulsion energy | 134.248308 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
-1.351 |
-0.574 |
0.118 |
| C2 |
-0.635 |
0.655 |
-0.275 |
| C3 |
0.790 |
0.555 |
0.272 |
| O4 |
1.406 |
-0.637 |
-0.173 |
| H5 |
-1.639 |
-0.489 |
1.098 |
| H6 |
-2.223 |
-0.648 |
-0.411 |
| H7 |
-1.117 |
1.596 |
0.065 |
| H8 |
-0.585 |
0.677 |
-1.379 |
| H9 |
1.400 |
1.407 |
-0.078 |
| H10 |
0.756 |
0.608 |
1.385 |
| H11 |
0.692 |
-1.293 |
-0.106 |
Atom - Atom Distances (Å)
| |
N1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| N1 | | 1.4753 | 2.4259 | 2.7728 | 1.0248 | 1.0224 | 2.1836 | 2.0957 | 3.3961 | 2.7283 | 2.1782 |
C2 | 1.4753 | | 1.5304 | 2.4173 | 2.0498 | 2.0583 | 1.1112 | 1.1054 | 2.1791 | 2.1671 | 2.3631 | C3 | 2.4259 | 1.5304 | | 1.4132 | 2.7701 | 3.3159 | 2.1830 | 2.1531 | 1.1049 | 1.1150 | 1.8889 | O4 | 2.7728 | 2.4173 | 1.4132 | | 3.3021 | 3.6365 | 3.3775 | 2.6732 | 2.0463 | 2.0972 | 0.9712 | H5 | 1.0248 | 2.0498 | 2.7701 | 3.3021 | | 1.6259 | 2.3855 | 2.9336 | 3.7703 | 2.6499 | 2.7443 | H6 | 1.0224 | 2.0583 | 3.3159 | 3.6365 | 1.6259 | | 2.5472 | 2.3182 | 4.1790 | 3.6987 | 3.0015 | H7 | 2.1836 | 1.1112 | 2.1830 | 3.3775 | 2.3855 | 2.5472 | | 1.7927 | 2.5283 | 2.4961 | 3.4135 | H8 | 2.0957 | 1.1054 | 2.1531 | 2.6732 | 2.9336 | 2.3182 | 1.7927 | | 2.4841 | 3.0739 | 2.6707 | H9 | 3.3961 | 2.1791 | 1.1049 | 2.0463 | 3.7703 | 4.1790 | 2.5283 | 2.4841 | | 1.7876 | 2.7915 | H10 | 2.7283 | 2.1671 | 1.1150 | 2.0972 | 2.6499 | 3.6987 | 2.4961 | 3.0739 | 1.7876 | | 2.4171 | H11 | 2.1782 | 2.3631 | 1.8889 | 0.9712 | 2.7443 | 3.0015 | 3.4135 | 2.6707 | 2.7915 | 2.4171 | |
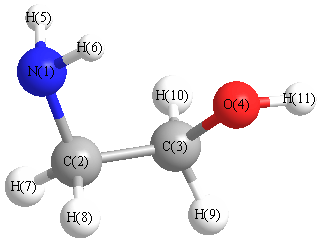 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C2 |
C3 |
107.612 |
|
N1 |
C2 |
H7 |
114.449 |
| N1 |
C2 |
H8 |
107.737 |
|
C2 |
N1 |
H5 |
108.810 |
| C2 |
N1 |
H6 |
109.663 |
|
C2 |
C3 |
O4 |
110.349 |
| C2 |
C3 |
H9 |
110.526 |
|
C2 |
C3 |
H10 |
109.000 |
| C3 |
C2 |
H7 |
110.453 |
|
C3 |
C2 |
H8 |
108.467 |
| C3 |
O4 |
H11 |
103.220 |
|
O4 |
C3 |
H9 |
108.086 |
| O4 |
C3 |
H10 |
111.561 |
|
H5 |
N1 |
H6 |
105.158 |
| H7 |
C2 |
H8 |
107.952 |
|
H9 |
C3 |
H10 |
107.276 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
