Jump to
S1C2
S1C3
Vibrational Frequencies calculated at CCD/cc-pVDZ
Geometric Data calculated at CCD/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Vibrational Frequencies calculated at CCD/cc-pVDZ
Geometric Data calculated at CCD/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at CCD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -209.813660 |
| Energy at 298.15K | -209.822912 |
| HF Energy | -209.122273 |
| Nuclear repulsion energy | 134.292541 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3811 |
3646 |
67.55 |
|
|
|
| 2 |
A |
3604 |
3448 |
0.85 |
|
|
|
| 3 |
A |
3517 |
3365 |
0.26 |
|
|
|
| 4 |
A |
3124 |
2989 |
49.13 |
|
|
|
| 5 |
A |
3115 |
2980 |
33.48 |
|
|
|
| 6 |
A |
3031 |
2899 |
66.54 |
|
|
|
| 7 |
A |
2995 |
2865 |
74.27 |
|
|
|
| 8 |
A |
1666 |
1594 |
23.79 |
|
|
|
| 9 |
A |
1523 |
1457 |
0.06 |
|
|
|
| 10 |
A |
1505 |
1440 |
13.21 |
|
|
|
| 11 |
A |
1487 |
1423 |
79.24 |
|
|
|
| 12 |
A |
1433 |
1371 |
15.70 |
|
|
|
| 13 |
A |
1398 |
1338 |
0.38 |
|
|
|
| 14 |
A |
1347 |
1289 |
2.25 |
|
|
|
| 15 |
A |
1277 |
1222 |
19.03 |
|
|
|
| 16 |
A |
1211 |
1158 |
3.72 |
|
|
|
| 17 |
A |
1154 |
1104 |
60.21 |
|
|
|
| 18 |
A |
1098 |
1051 |
19.22 |
|
|
|
| 19 |
A |
1036 |
991 |
23.07 |
|
|
|
| 20 |
A |
977 |
934 |
58.27 |
|
|
|
| 21 |
A |
908 |
868 |
11.23 |
|
|
|
| 22 |
A |
863 |
825 |
51.80 |
|
|
|
| 23 |
A |
573 |
548 |
123.71 |
|
|
|
| 24 |
A |
541 |
518 |
14.29 |
|
|
|
| 25 |
A |
333 |
319 |
0.62 |
|
|
|
| 26 |
A |
284 |
272 |
14.51 |
|
|
|
| 27 |
A |
200 |
191 |
6.23 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22003.9 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 21051.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
-1.354 |
-0.573 |
0.117 |
| C2 |
-0.636 |
0.652 |
-0.275 |
| C3 |
0.789 |
0.553 |
0.273 |
| O4 |
1.408 |
-0.634 |
-0.173 |
| H5 |
-1.639 |
-0.490 |
1.098 |
| H6 |
-2.226 |
-0.645 |
-0.409 |
| H7 |
-1.116 |
1.595 |
0.063 |
| H8 |
-0.585 |
0.674 |
-1.379 |
| H9 |
1.397 |
1.408 |
-0.075 |
| H10 |
0.755 |
0.603 |
1.385 |
| H11 |
0.702 |
-1.295 |
-0.109 |
Atom - Atom Distances (Å)
| |
N1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| N1 | | 1.4737 | 2.4263 | 2.7779 | 1.0240 | 1.0217 | 2.1817 | 2.0945 | 3.3953 | 2.7276 | 2.1904 |
C2 | 1.4737 | | 1.5301 | 2.4170 | 2.0485 | 2.0569 | 1.1107 | 1.1050 | 2.1777 | 2.1667 | 2.3681 | C3 | 2.4263 | 1.5301 | | 1.4111 | 2.7685 | 3.3160 | 2.1815 | 2.1520 | 1.1045 | 1.1142 | 1.8894 | O4 | 2.7779 | 2.4170 | 1.4111 | | 3.3041 | 3.6423 | 3.3755 | 2.6717 | 2.0441 | 2.0940 | 0.9695 | H5 | 1.0240 | 2.0485 | 2.7685 | 3.3041 | | 1.6249 | 2.3858 | 2.9327 | 3.7671 | 2.6474 | 2.7532 | H6 | 1.0217 | 2.0569 | 3.3160 | 3.6423 | 1.6249 | | 2.5441 | 2.3185 | 4.1777 | 3.6971 | 3.0144 | H7 | 2.1817 | 1.1107 | 2.1815 | 3.3755 | 2.3858 | 2.5441 | | 1.7912 | 2.5235 | 2.4966 | 3.4182 | H8 | 2.0945 | 1.1050 | 2.1520 | 2.6717 | 2.9327 | 2.3185 | 1.7912 | | 2.4831 | 3.0728 | 2.6731 | H9 | 3.3953 | 2.1777 | 1.1045 | 2.0441 | 3.7671 | 4.1777 | 2.5235 | 2.4831 | | 1.7863 | 2.7909 | H10 | 2.7276 | 2.1667 | 1.1142 | 2.0940 | 2.6474 | 3.6971 | 2.4966 | 3.0728 | 1.7863 | | 2.4165 | H11 | 2.1904 | 2.3681 | 1.8894 | 0.9695 | 2.7532 | 3.0144 | 3.4182 | 2.6731 | 2.7909 | 2.4165 | |
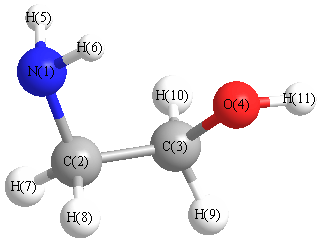 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C2 |
C3 |
107.741 |
|
N1 |
C2 |
H7 |
114.444 |
| N1 |
C2 |
H8 |
107.776 |
|
C2 |
N1 |
H5 |
108.870 |
| C2 |
N1 |
H6 |
109.699 |
|
C2 |
C3 |
O4 |
110.464 |
| C2 |
C3 |
H9 |
110.462 |
|
C2 |
C3 |
H10 |
109.040 |
| C3 |
C2 |
H7 |
110.391 |
|
C3 |
C2 |
H8 |
108.432 |
| C3 |
O4 |
H11 |
103.499 |
|
O4 |
C3 |
H9 |
108.081 |
| O4 |
C3 |
H10 |
111.498 |
|
H5 |
N1 |
H6 |
105.174 |
| H7 |
C2 |
H8 |
107.887 |
|
H9 |
C3 |
H10 |
107.246 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
