Jump to
S1C2
S1C3
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
Geometric Data calculated at PBEPBE/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
Geometric Data calculated at PBEPBE/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -210.134999 |
| Energy at 298.15K | -210.144185 |
| HF Energy | -210.134999 |
| Nuclear repulsion energy | 134.360226 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3474 |
3454 |
1.66 |
|
|
|
| 2 |
A |
3440 |
3420 |
113.35 |
|
|
|
| 3 |
A |
3385 |
3365 |
0.35 |
|
|
|
| 4 |
A |
3009 |
2992 |
37.14 |
|
|
|
| 5 |
A |
2991 |
2973 |
42.69 |
|
|
|
| 6 |
A |
2906 |
2889 |
79.55 |
|
|
|
| 7 |
A |
2831 |
2814 |
101.21 |
|
|
|
| 8 |
A |
1584 |
1575 |
20.39 |
|
|
|
| 9 |
A |
1438 |
1430 |
0.39 |
|
|
|
| 10 |
A |
1432 |
1424 |
97.14 |
|
|
|
| 11 |
A |
1412 |
1403 |
18.43 |
|
|
|
| 12 |
A |
1349 |
1341 |
5.86 |
|
|
|
| 13 |
A |
1314 |
1306 |
0.43 |
|
|
|
| 14 |
A |
1273 |
1266 |
2.87 |
|
|
|
| 15 |
A |
1211 |
1204 |
15.57 |
|
|
|
| 16 |
A |
1152 |
1145 |
2.60 |
|
|
|
| 17 |
A |
1087 |
1081 |
58.51 |
|
|
|
| 18 |
A |
1047 |
1041 |
25.73 |
|
|
|
| 19 |
A |
974 |
969 |
6.86 |
|
|
|
| 20 |
A |
909 |
904 |
58.91 |
|
|
|
| 21 |
A |
872 |
867 |
5.22 |
|
|
|
| 22 |
A |
805 |
800 |
62.05 |
|
|
|
| 23 |
A |
646 |
642 |
95.01 |
|
|
|
| 24 |
A |
539 |
535 |
7.91 |
|
|
|
| 25 |
A |
326 |
324 |
1.06 |
|
|
|
| 26 |
A |
287 |
286 |
12.25 |
|
|
|
| 27 |
A |
200 |
199 |
8.22 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20945.4 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 20823.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
-1.325 |
-0.571 |
0.116 |
| C2 |
-0.634 |
0.678 |
-0.255 |
| C3 |
0.810 |
0.561 |
0.251 |
| O4 |
1.367 |
-0.670 |
-0.166 |
| H5 |
-1.624 |
-0.524 |
1.101 |
| H6 |
-2.188 |
-0.681 |
-0.431 |
| H7 |
-1.113 |
1.614 |
0.123 |
| H8 |
-0.618 |
0.730 |
-1.364 |
| H9 |
1.432 |
1.390 |
-0.149 |
| H10 |
0.811 |
0.665 |
1.371 |
| H11 |
0.581 |
-1.271 |
-0.113 |
Atom - Atom Distances (Å)
| |
N1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| N1 | | 1.4753 | 2.4200 | 2.7080 | 1.0307 | 1.0278 | 2.1950 | 2.0938 | 3.3935 | 2.7688 | 2.0433 |
C2 | 1.4753 | | 1.5342 | 2.4142 | 2.0655 | 2.0727 | 1.1171 | 1.1103 | 2.1874 | 2.1752 | 2.3016 | C3 | 2.4200 | 1.5342 | | 1.4143 | 2.7967 | 3.3161 | 2.1962 | 2.1627 | 1.1111 | 1.1243 | 1.8824 | O4 | 2.7080 | 2.4142 | 1.4143 | | 3.2510 | 3.5649 | 3.3839 | 2.7087 | 2.0615 | 2.1104 | 0.9907 | H5 | 1.0307 | 2.0655 | 2.7967 | 3.2510 | | 1.6400 | 2.4060 | 2.9435 | 3.8163 | 2.7233 | 2.6257 | H6 | 1.0278 | 2.0727 | 3.3161 | 3.5649 | 1.6400 | | 2.5937 | 2.3087 | 4.1804 | 3.7490 | 2.8493 | H7 | 2.1950 | 1.1171 | 2.1962 | 3.3839 | 2.4060 | 2.5937 | | 1.7990 | 2.5696 | 2.4823 | 3.3543 | H8 | 2.0938 | 1.1103 | 2.1627 | 2.7087 | 2.9435 | 2.3087 | 1.7990 | | 2.4728 | 3.0866 | 2.6475 | H9 | 3.3935 | 2.1874 | 1.1111 | 2.0615 | 3.8163 | 4.1804 | 2.5696 | 2.4728 | | 1.7943 | 2.7945 | H10 | 2.7688 | 2.1752 | 1.1243 | 2.1104 | 2.7233 | 3.7490 | 2.4823 | 3.0866 | 1.7943 | | 2.4510 | H11 | 2.0433 | 2.3016 | 1.8824 | 0.9907 | 2.6257 | 2.8493 | 3.3543 | 2.6475 | 2.7945 | 2.4510 | |
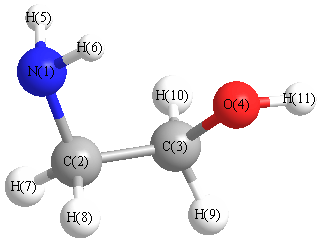 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C2 |
C3 |
107.031 |
|
N1 |
C2 |
H7 |
115.015 |
| N1 |
C2 |
H8 |
107.308 |
|
C2 |
N1 |
H5 |
109.743 |
| C2 |
N1 |
H6 |
110.517 |
|
C2 |
C3 |
O4 |
109.859 |
| C2 |
C3 |
H9 |
110.541 |
|
C2 |
C3 |
H10 |
108.838 |
| C3 |
C2 |
H7 |
110.885 |
|
C3 |
C2 |
H8 |
108.669 |
| C3 |
O4 |
H11 |
101.560 |
|
O4 |
C3 |
H9 |
108.839 |
| O4 |
C3 |
H10 |
111.961 |
|
H5 |
N1 |
H6 |
105.629 |
| H7 |
C2 |
H8 |
107.738 |
|
H9 |
C3 |
H10 |
106.767 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.181 |
|
|
|
| 2 |
C |
-0.042 |
|
|
|
| 3 |
C |
0.113 |
|
|
|
| 4 |
O |
-0.256 |
|
|
|
| 5 |
H |
0.086 |
|
|
|
| 6 |
H |
0.088 |
|
|
|
| 7 |
H |
0.012 |
|
|
|
| 8 |
H |
0.037 |
|
|
|
| 9 |
H |
0.022 |
|
|
|
| 10 |
H |
-0.002 |
|
|
|
| 11 |
H |
0.125 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.931 |
0.889 |
0.482 |
3.101 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.500 |
2.241 |
0.075 |
| y |
2.241 |
-25.677 |
-0.357 |
| z |
0.075 |
-0.357 |
-24.650 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.336 |
2.241 |
0.075 |
| y |
2.241 |
-0.602 |
-0.357 |
| z |
0.075 |
-0.357 |
0.939 |
|
| Polar |
|---|
| 3z2-r2 | 1.877 |
|---|
| x2-y2 | 0.177 |
|---|
| xy | 2.241 |
|---|
| xz | 0.075 |
|---|
| yz | -0.357 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.824 |
0.212 |
0.032 |
| y |
0.212 |
5.419 |
0.079 |
| z |
0.032 |
0.079 |
4.986 |
<r2> (average value of r
2) Å
2
| <r2> |
86.118 |
| (<r2>)1/2 |
9.280 |
