Jump to
S1C2
S1C3
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
Geometric Data calculated at PBEPBEultrafine/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
Geometric Data calculated at PBEPBEultrafine/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at PBEPBEultrafine/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -210.134996 |
| Energy at 298.15K | -210.144184 |
| HF Energy | -210.134996 |
| Nuclear repulsion energy | 134.344553 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3473 |
3453 |
1.66 |
|
|
|
| 2 |
A |
3438 |
3418 |
113.15 |
|
|
|
| 3 |
A |
3385 |
3365 |
0.35 |
|
|
|
| 4 |
A |
3010 |
2992 |
37.50 |
|
|
|
| 5 |
A |
2992 |
2975 |
42.23 |
|
|
|
| 6 |
A |
2907 |
2890 |
79.68 |
|
|
|
| 7 |
A |
2831 |
2815 |
101.23 |
|
|
|
| 8 |
A |
1585 |
1576 |
20.34 |
|
|
|
| 9 |
A |
1438 |
1430 |
0.87 |
|
|
|
| 10 |
A |
1431 |
1423 |
93.11 |
|
|
|
| 11 |
A |
1411 |
1403 |
20.89 |
|
|
|
| 12 |
A |
1350 |
1342 |
6.00 |
|
|
|
| 13 |
A |
1314 |
1306 |
0.46 |
|
|
|
| 14 |
A |
1273 |
1266 |
2.88 |
|
|
|
| 15 |
A |
1211 |
1204 |
15.82 |
|
|
|
| 16 |
A |
1152 |
1145 |
2.78 |
|
|
|
| 17 |
A |
1087 |
1081 |
58.80 |
|
|
|
| 18 |
A |
1047 |
1041 |
25.61 |
|
|
|
| 19 |
A |
975 |
969 |
6.99 |
|
|
|
| 20 |
A |
910 |
904 |
59.21 |
|
|
|
| 21 |
A |
872 |
867 |
5.34 |
|
|
|
| 22 |
A |
805 |
801 |
61.75 |
|
|
|
| 23 |
A |
648 |
644 |
94.79 |
|
|
|
| 24 |
A |
539 |
536 |
7.99 |
|
|
|
| 25 |
A |
326 |
324 |
1.00 |
|
|
|
| 26 |
A |
288 |
286 |
12.46 |
|
|
|
| 27 |
A |
200 |
198 |
8.21 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20948.5 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 20827.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
-1.325 |
-0.571 |
0.116 |
| C2 |
-0.634 |
0.678 |
-0.255 |
| C3 |
0.810 |
0.561 |
0.251 |
| O4 |
1.368 |
-0.670 |
-0.166 |
| H5 |
-1.624 |
-0.524 |
1.101 |
| H6 |
-2.189 |
-0.681 |
-0.430 |
| H7 |
-1.114 |
1.613 |
0.122 |
| H8 |
-0.618 |
0.730 |
-1.364 |
| H9 |
1.432 |
1.390 |
-0.148 |
| H10 |
0.811 |
0.666 |
1.371 |
| H11 |
0.582 |
-1.271 |
-0.112 |
Atom - Atom Distances (Å)
| |
N1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| N1 | | 1.4754 | 2.4206 | 2.7094 | 1.0308 | 1.0279 | 2.1948 | 2.0938 | 3.3941 | 2.7694 | 2.0444 |
C2 | 1.4754 | | 1.5343 | 2.4146 | 2.0653 | 2.0730 | 1.1171 | 1.1103 | 2.1878 | 2.1753 | 2.3016 | C3 | 2.4206 | 1.5343 | | 1.4144 | 2.7969 | 3.3169 | 2.1965 | 2.1628 | 1.1111 | 1.1243 | 1.8817 | O4 | 2.7094 | 2.4146 | 1.4144 | | 3.2521 | 3.5669 | 3.3844 | 2.7090 | 2.0610 | 2.1107 | 0.9907 | H5 | 1.0308 | 2.0653 | 2.7969 | 3.2521 | | 1.6399 | 2.4055 | 2.9433 | 3.8164 | 2.7235 | 2.6263 | H6 | 1.0279 | 2.0730 | 3.3169 | 3.5669 | 1.6399 | | 2.5932 | 2.3092 | 4.1813 | 3.7496 | 2.8513 | H7 | 2.1948 | 1.1171 | 2.1965 | 3.3844 | 2.4055 | 2.5932 | | 1.7988 | 2.5703 | 2.4826 | 3.3542 | H8 | 2.0938 | 1.1103 | 2.1628 | 2.7090 | 2.9433 | 2.3092 | 1.7988 | | 2.4733 | 3.0867 | 2.6479 | H9 | 3.3941 | 2.1878 | 1.1111 | 2.0610 | 3.8164 | 4.1813 | 2.5703 | 2.4733 | | 1.7941 | 2.7938 | H10 | 2.7694 | 2.1753 | 1.1243 | 2.1107 | 2.7235 | 3.7496 | 2.4826 | 3.0867 | 1.7941 | | 2.4502 | H11 | 2.0444 | 2.3016 | 1.8817 | 0.9907 | 2.6263 | 2.8513 | 3.3542 | 2.6479 | 2.7938 | 2.4502 | |
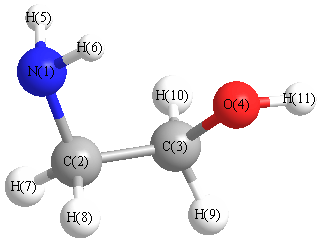 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C2 |
C3 |
107.063 |
|
N1 |
C2 |
H7 |
114.989 |
| N1 |
C2 |
H8 |
107.303 |
|
C2 |
N1 |
H5 |
109.715 |
| C2 |
N1 |
H6 |
110.534 |
|
C2 |
C3 |
O4 |
109.878 |
| C2 |
C3 |
H9 |
110.569 |
|
C2 |
C3 |
H10 |
108.832 |
| C3 |
C2 |
H7 |
110.901 |
|
C3 |
C2 |
H8 |
108.670 |
| C3 |
O4 |
H11 |
101.502 |
|
O4 |
C3 |
H9 |
108.794 |
| O4 |
C3 |
H10 |
111.977 |
|
H5 |
N1 |
H6 |
105.603 |
| H7 |
C2 |
H8 |
107.721 |
|
H9 |
C3 |
H10 |
106.755 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.181 |
|
|
|
| 2 |
C |
-0.042 |
|
|
|
| 3 |
C |
0.113 |
|
|
|
| 4 |
O |
-0.256 |
|
|
|
| 5 |
H |
0.086 |
|
|
|
| 6 |
H |
0.088 |
|
|
|
| 7 |
H |
0.012 |
|
|
|
| 8 |
H |
0.037 |
|
|
|
| 9 |
H |
0.022 |
|
|
|
| 10 |
H |
-0.002 |
|
|
|
| 11 |
H |
0.125 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.930 |
0.890 |
0.484 |
3.100 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.496 |
2.237 |
0.075 |
| y |
2.237 |
-25.681 |
-0.359 |
| z |
0.075 |
-0.359 |
-24.653 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.329 |
2.237 |
0.075 |
| y |
2.237 |
-0.607 |
-0.359 |
| z |
0.075 |
-0.359 |
0.936 |
|
| Polar |
|---|
| 3z2-r2 | 1.872 |
|---|
| x2-y2 | 0.185 |
|---|
| xy | 2.237 |
|---|
| xz | 0.075 |
|---|
| yz | -0.359 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.826 |
0.212 |
0.032 |
| y |
0.212 |
5.418 |
0.079 |
| z |
0.032 |
0.079 |
4.986 |
<r2> (average value of r
2) Å
2
| <r2> |
86.149 |
| (<r2>)1/2 |
9.282 |
