Jump to
S1C2
S1C3
Vibrational Frequencies calculated at MP3=FULL/cc-pVDZ
Geometric Data calculated at MP3=FULL/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Vibrational Frequencies calculated at MP3=FULL/cc-pVDZ
Geometric Data calculated at MP3=FULL/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at MP3=FULL/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -209.818389 |
| Energy at 298.15K | -209.827658 |
| HF Energy | -209.122635 |
| Nuclear repulsion energy | 134.526348 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3=FULL/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3829 |
3600 |
72.22 |
|
|
|
| 2 |
A |
3631 |
3414 |
1.24 |
|
|
|
| 3 |
A |
3546 |
3334 |
0.26 |
|
|
|
| 4 |
A |
3146 |
2959 |
48.72 |
|
|
|
| 5 |
A |
3137 |
2950 |
33.35 |
|
|
|
| 6 |
A |
3053 |
2871 |
65.64 |
|
|
|
| 7 |
A |
3015 |
2835 |
74.91 |
|
|
|
| 8 |
A |
1670 |
1570 |
24.75 |
|
|
|
| 9 |
A |
1527 |
1436 |
0.07 |
|
|
|
| 10 |
A |
1510 |
1420 |
17.59 |
|
|
|
| 11 |
A |
1494 |
1404 |
79.29 |
|
|
|
| 12 |
A |
1436 |
1350 |
14.00 |
|
|
|
| 13 |
A |
1402 |
1318 |
0.29 |
|
|
|
| 14 |
A |
1351 |
1270 |
2.30 |
|
|
|
| 15 |
A |
1281 |
1204 |
19.04 |
|
|
|
| 16 |
A |
1215 |
1143 |
3.47 |
|
|
|
| 17 |
A |
1159 |
1090 |
62.45 |
|
|
|
| 18 |
A |
1103 |
1037 |
18.61 |
|
|
|
| 19 |
A |
1037 |
975 |
21.56 |
|
|
|
| 20 |
A |
978 |
919 |
58.63 |
|
|
|
| 21 |
A |
911 |
857 |
10.51 |
|
|
|
| 22 |
A |
864 |
812 |
54.15 |
|
|
|
| 23 |
A |
581 |
547 |
124.86 |
|
|
|
| 24 |
A |
545 |
512 |
12.01 |
|
|
|
| 25 |
A |
334 |
314 |
0.62 |
|
|
|
| 26 |
A |
285 |
268 |
14.63 |
|
|
|
| 27 |
A |
201 |
189 |
6.32 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22119.6 cm
-1
Scaled (by 0.9403) Zero Point Vibrational Energy (zpe) 20799.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP3=FULL/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
-1.350 |
-0.572 |
0.117 |
| C2 |
-0.634 |
0.654 |
-0.274 |
| C3 |
0.790 |
0.553 |
0.271 |
| O4 |
1.402 |
-0.636 |
-0.172 |
| H5 |
-1.632 |
-0.492 |
1.096 |
| H6 |
-2.221 |
-0.645 |
-0.408 |
| H7 |
-1.113 |
1.593 |
0.066 |
| H8 |
-0.584 |
0.677 |
-1.375 |
| H9 |
1.397 |
1.404 |
-0.078 |
| H10 |
0.757 |
0.605 |
1.382 |
| H11 |
0.691 |
-1.290 |
-0.112 |
Atom - Atom Distances (Å)
| |
N1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| N1 | | 1.4723 | 2.4233 | 2.7687 | 1.0217 | 1.0194 | 2.1790 | 2.0918 | 3.3901 | 2.7253 | 2.1766 |
C2 | 1.4723 | | 1.5289 | 2.4128 | 2.0459 | 2.0544 | 1.1082 | 1.1027 | 2.1748 | 2.1634 | 2.3584 | C3 | 2.4233 | 1.5289 | | 1.4089 | 2.7642 | 3.3113 | 2.1789 | 2.1489 | 1.1021 | 1.1119 | 1.8853 | O4 | 2.7687 | 2.4128 | 1.4089 | | 3.2917 | 3.6310 | 3.3696 | 2.6685 | 2.0423 | 2.0906 | 0.9679 | H5 | 1.0217 | 2.0459 | 2.7642 | 3.2917 | | 1.6228 | 2.3835 | 2.9282 | 3.7618 | 2.6448 | 2.7376 | H6 | 1.0194 | 2.0544 | 3.3113 | 3.6310 | 1.6228 | | 2.5420 | 2.3156 | 4.1712 | 3.6927 | 2.9977 | H7 | 2.1790 | 1.1082 | 2.1789 | 3.3696 | 2.3835 | 2.5420 | | 1.7877 | 2.5216 | 2.4910 | 3.4063 | H8 | 2.0918 | 1.1027 | 2.1489 | 2.6685 | 2.9282 | 2.3156 | 1.7877 | | 2.4777 | 3.0670 | 2.6638 | H9 | 3.3901 | 2.1748 | 1.1021 | 2.0423 | 3.7618 | 4.1712 | 2.5216 | 2.4777 | | 1.7825 | 2.7854 | H10 | 2.7253 | 2.1634 | 1.1119 | 2.0906 | 2.6448 | 3.6927 | 2.4910 | 3.0670 | 1.7825 | | 2.4142 | H11 | 2.1766 | 2.3584 | 1.8853 | 0.9679 | 2.7376 | 2.9977 | 3.4063 | 2.6638 | 2.7854 | 2.4142 | |
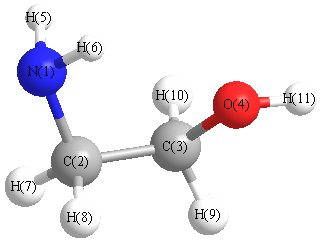 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C2 |
C3 |
107.679 |
|
N1 |
C2 |
H7 |
114.477 |
| N1 |
C2 |
H8 |
107.791 |
|
C2 |
N1 |
H5 |
108.894 |
| C2 |
N1 |
H6 |
109.744 |
|
C2 |
C3 |
O4 |
110.362 |
| C2 |
C3 |
H9 |
110.449 |
|
C2 |
C3 |
H10 |
108.990 |
| C3 |
C2 |
H7 |
110.411 |
|
C3 |
C2 |
H8 |
108.394 |
| C3 |
O4 |
H11 |
103.410 |
|
O4 |
C3 |
H9 |
108.227 |
| O4 |
C3 |
H10 |
111.524 |
|
H5 |
N1 |
H6 |
105.321 |
| H7 |
C2 |
H8 |
107.915 |
|
H9 |
C3 |
H10 |
107.243 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
