return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H8O2 (Ethyl acetate)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-306.875830
Energy at 298.15K 
HF Energy-305.896456
Nuclear repulsion energy242.786399
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3205 3075 7.31      
2 A' 3157 3028 23.86      
3 A' 3084 2959 7.74      
4 A' 3083 2958 13.50      
5 A' 3068 2943 18.92      
6 A' 1852 1776 222.84      
7 A' 1524 1463 3.27      
8 A' 1499 1438 2.19      
9 A' 1472 1412 12.06      
10 A' 1447 1388 4.74      
11 A' 1415 1357 91.92      
12 A' 1392 1335 16.24      
13 A' 1301 1248 392.52      
14 A' 1152 1105 15.44      
15 A' 1112 1067 77.81      
16 A' 1020 979 11.57      
17 A' 966 927 7.14      
18 A' 879 843 10.41      
19 A' 648 621 9.18      
20 A' 427 410 0.61      
21 A' 376 361 9.52      
22 A' 199 190 4.06      
23 A" 3167 3039 7.57      
24 A" 3163 3035 35.05      
25 A" 3129 3002 12.59      
26 A" 1484 1424 4.94      
27 A" 1478 1418 6.88      
28 A" 1302 1249 1.28      
29 A" 1190 1142 4.30      
30 A" 1070 1026 5.33      
31 A" 814 781 0.16      
32 A" 607 583 5.95      
33 A" 274 263 1.04      
34 A" 153 146 3.96      
35 A" 83 79 0.02      
36 A" 69 66 0.76      

Unscaled Zero Point Vibrational Energy (zpe) 26128.9 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 25068.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.27811 0.06971 0.05757

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -2.317 0.036 0.000
C2 -0.907 -0.518 0.000
O3 0.000 0.489 0.000
O4 -0.608 -1.692 0.000
C5 1.377 0.058 0.000
C6 2.237 1.312 0.000
H7 -3.038 -0.793 0.000
H8 -2.465 0.671 0.890
H9 -2.465 0.671 -0.890
H10 1.562 -0.568 0.891
H11 1.562 -0.568 -0.891
H12 3.306 1.035 0.000
H13 2.032 1.924 -0.896
H14 2.032 1.924 0.896

Atom - Atom Distances (Å)
  C1 C2 O3 O4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14
C11.51442.36042.43043.69354.72861.09941.10331.10334.02544.02545.71064.82414.8241
C21.51441.35601.21142.35593.63822.14842.15262.15262.62582.62584.49073.92513.9251
O32.36041.35602.26471.44282.38333.29772.62732.62732.08622.08623.35082.64362.6436
O42.43041.21142.26472.64654.13752.59113.13463.13462.60132.60134.77074.56604.5660
C53.69352.35591.44282.64651.52044.49633.99113.99111.10471.10472.16252.17082.1708
C64.72863.63822.38334.13751.52045.67954.82824.82822.18732.18731.10451.10391.1039
H71.09942.14843.29772.59114.49635.67951.80671.80674.69124.69126.60235.82135.8213
H81.10332.15262.62733.13463.99114.82821.80671.78064.21374.57465.85074.99814.6682
H91.10332.15262.62733.13463.99114.82821.80671.78064.57464.21375.85074.66824.9981
H104.02542.62582.08622.60131.10472.18734.69124.21374.57461.78152.53093.10222.5363
H114.02542.62582.08622.60131.10472.18734.69124.57464.21371.78152.53092.53633.1022
H125.71064.49073.35084.77072.16251.10456.60235.85075.85072.53092.53091.79331.7933
H134.82413.92512.64364.56602.17081.10395.82134.99814.66823.10222.53631.79331.7911
H144.82413.92512.64364.56602.17081.10395.82134.66824.99812.53633.10221.79331.7911

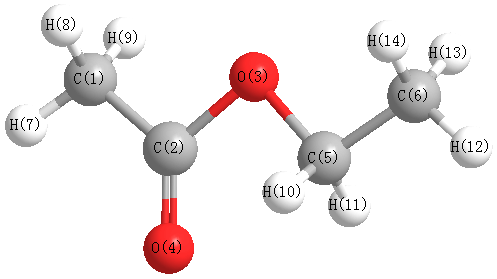
picture of Ethyl acetate state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O3 110.514 C1 C2 O4 125.795
C2 C1 H7 109.532 C2 C1 H8 109.633
C2 C1 H9 109.633 C2 O3 C5 114.619
O3 C2 O4 123.691 O3 C5 C6 107.058
O3 C5 H10 109.242 O3 C5 H11 109.242
C5 C6 H12 109.928 C5 C6 H13 110.619
C5 C6 H14 110.619 C6 C5 H10 111.890
C6 C5 H11 111.890 H7 C1 H8 110.212
H7 C1 H9 110.212 H8 C1 H9 107.592
H10 C5 H11 107.484 H12 C6 H13 108.591
H12 C6 H14 108.591 H13 C6 H14 108.432
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability