return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H8O2 (Ethyl acetate)

using model chemistry: CCD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CCD/cc-pVDZ
 hartrees
Energy at 0K-306.864745
Energy at 298.15K 
HF Energy-305.897328
Nuclear repulsion energy243.220859
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3215 3076 6.69      
2 A' 3169 3032 22.22      
3 A' 3093 2959 5.16      
4 A' 3090 2956 15.02      
5 A' 3078 2944 18.10      
6 A' 1897 1815 235.06      
7 A' 1530 1463 3.35      
8 A' 1502 1437 2.14      
9 A' 1475 1411 13.05      
10 A' 1455 1392 8.28      
11 A' 1421 1359 115.78      
12 A' 1397 1337 17.91      
13 A' 1316 1259 399.35      
14 A' 1160 1110 25.23      
15 A' 1125 1077 57.89      
16 A' 1025 980 10.00      
17 A' 976 934 6.40      
18 A' 889 850 10.28      
19 A' 655 627 10.24      
20 A' 430 412 0.64      
21 A' 380 363 9.49      
22 A' 200 191 4.14      
23 A" 3178 3040 6.81      
24 A" 3174 3037 32.07      
25 A" 3137 3002 14.30      
26 A" 1486 1422 5.02      
27 A" 1480 1416 6.99      
28 A" 1309 1252 1.44      
29 A" 1195 1143 4.47      
30 A" 1076 1029 5.69      
31 A" 817 782 0.16      
32 A" 613 586 5.93      
33 A" 274 263 1.04      
34 A" 153 146 4.00      
35 A" 89 85 0.12      
36 A" 75 72 0.66      

Unscaled Zero Point Vibrational Energy (zpe) 26265.8 cm-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 25128.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/cc-pVDZ
ABC
0.27965 0.06986 0.05774

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -2.314 0.031 0.000
C2 -0.904 -0.520 0.000
O3 0.000 0.485 0.000
O4 -0.605 -1.688 0.000
C5 1.374 0.061 0.000
C6 2.232 1.316 0.000
H7 -3.033 -0.800 0.000
H8 -2.464 0.665 0.890
H9 -2.464 0.665 -0.890
H10 1.564 -0.565 0.890
H11 1.564 -0.565 -0.890
H12 3.301 1.042 0.000
H13 2.025 1.928 -0.895
H14 2.025 1.928 0.895

Atom - Atom Distances (Å)
  C1 C2 O3 O4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14
C11.51332.35802.42373.68794.72371.09901.10291.10294.02284.02285.70504.81954.8195
C21.51331.35191.20612.35113.63392.14722.15122.15122.62412.62414.48583.92113.9211
O32.35801.35192.25621.43802.38143.29422.62572.62572.08332.08333.34752.64292.6429
O42.42371.20612.25622.64144.13242.58523.12793.12792.60012.60014.76584.56064.5606
C53.68792.35111.43802.64141.52064.49043.98563.98561.10441.10442.16232.17072.1707
C64.72373.63392.38144.13241.52065.67424.82314.82312.18562.18561.10391.10351.1035
H71.09902.14723.29422.58524.49045.67421.80621.80624.68834.68836.59645.81645.8164
H81.10292.15122.62573.12793.98564.82311.80621.77964.21114.57195.84514.99334.6634
H91.10292.15122.62573.12793.98564.82311.80621.77964.57194.21115.84514.66344.9933
H104.02282.62412.08332.60011.10442.18564.68834.21114.57191.78062.52803.10062.5349
H114.02282.62412.08332.60011.10442.18564.68834.57194.21111.78062.52802.53493.1006
H125.70504.48583.34754.76582.16231.10396.59645.84515.84512.52802.52801.79251.7925
H134.81953.92112.64294.56062.17071.10355.81644.99334.66343.10062.53491.79251.7903
H144.81953.92112.64294.56062.17071.10355.81644.66344.99332.53493.10061.79251.7903

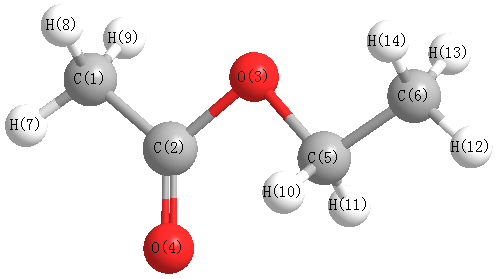
picture of Ethyl acetate state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O3 110.638 C1 C2 O4 125.691
C2 C1 H7 109.542 C2 C1 H8 109.628
C2 C1 H9 109.628 C2 O3 C5 114.816
O3 C2 O4 123.671 O3 C5 C6 107.170
O3 C5 H10 109.351 O3 C5 H11 109.351
C5 C6 H12 109.940 C5 C6 H13 110.621
C5 C6 H14 110.621 C6 C5 H10 111.755
C6 C5 H11 111.755 H7 C1 H8 110.225
H7 C1 H9 110.225 H8 C1 H9 107.567
H10 C5 H11 107.434 H12 C6 H13 108.589
H12 C6 H14 108.589 H13 C6 H14 108.421
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability