return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8 (Cyclopentene)

using model chemistry: CCD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CCD/cc-pVDZ
 hartrees
Energy at 0K-194.730270
Energy at 298.15K-194.739288
HF Energy-193.990423
Nuclear repulsion energy170.295682
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3242 3101 20.38      
2 A' 3144 3008 41.95      
3 A' 3115 2980 36.37      
4 A' 3082 2949 20.25      
5 A' 3052 2920 27.45      
6 A' 1707 1633 1.34      
7 A' 1509 1444 0.85      
8 A' 1481 1417 1.59      
9 A' 1351 1292 0.14      
10 A' 1242 1189 1.18      
11 A' 1128 1080 0.10      
12 A' 1078 1031 6.35      
13 A' 993 950 0.93      
14 A' 932 892 0.37      
15 A' 842 806 0.50      
16 A' 720 688 29.68      
17 A' 598 572 7.16      
18 A' 180 172 0.10      
19 A" 3215 3076 6.08      
20 A" 3113 2978 24.45      
21 A" 3054 2922 52.41      
22 A" 1491 1426 0.75      
23 A" 1396 1336 1.09      
24 A" 1345 1286 1.07      
25 A" 1311 1254 0.58      
26 A" 1238 1185 0.00      
27 A" 1153 1103 1.43      
28 A" 1058 1012 1.33      
29 A" 964 922 0.15      
30 A" 935 894 5.87      
31 A" 901 862 1.01      
32 A" 779 746 0.16      
33 A" 389 373 0.05      

Unscaled Zero Point Vibrational Energy (zpe) 25868.3 cm-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 24748.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/cc-pVDZ
ABC
0.24116 0.23894 0.13094

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.321 -1.189 0.000
C2 -0.083 -0.338 1.233
C3 -0.083 -0.338 -1.233
C4 -0.083 1.076 0.674
C5 -0.083 1.076 -0.674
H6 1.419 -1.319 0.000
H7 -0.135 -2.194 0.000
H8 -1.095 -0.604 1.600
H9 -1.095 -0.604 -1.600
H10 -0.128 1.972 1.302
H11 -0.128 1.972 -1.302
H12 0.611 -0.470 -2.083
H13 0.611 -0.470 2.083

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13
C11.55131.55132.39752.39751.10571.10342.21502.21503.44863.44862.22282.2228
C21.55132.46571.52102.37382.17622.22871.10913.02002.31213.43053.39031.1050
C31.55132.46572.37381.52102.17622.22873.02001.10913.43052.31211.10503.3903
C42.39751.52102.37381.34722.90613.33942.16913.00271.09572.17023.23602.2043
C52.39752.37381.52101.34722.90613.33943.00272.16912.17021.09572.20433.2360
H61.10572.17622.17622.90612.90611.78293.06423.06423.86313.86312.39022.3902
H71.10342.22872.22873.33943.33941.78292.45182.45184.36544.36542.80492.8049
H82.21501.10913.02002.16913.00273.06422.45183.20012.76763.99944.06111.7776
H92.21503.02001.10913.00272.16913.06422.45183.20013.99942.76761.77764.0611
H103.44862.31213.43051.09572.17023.86314.36542.76763.99942.60474.23962.6687
H113.44863.43052.31212.17021.09573.86314.36543.99942.76762.60472.66874.2396
H122.22283.39031.10503.23602.20432.39022.80494.06111.77764.23962.66874.1664
H132.22281.10503.39032.20433.23602.39022.80491.77764.06112.66874.23964.1664

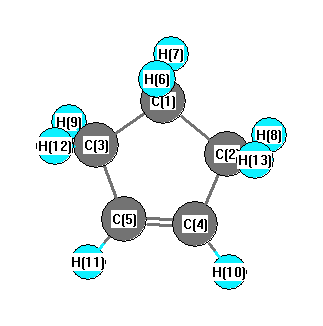
picture of Cyclopentene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 102.584 C1 C2 H8 111.652
C1 C2 H13 112.519 C1 C3 C5 102.584
C1 C3 H9 111.652 C1 C3 H12 112.519
C2 C1 C3 105.259 C2 C1 H6 108.820
C2 C1 H7 113.102 C2 C4 C5 111.575
C2 C4 H10 123.341 C3 C1 H6 108.820
C3 C1 H7 113.102 C3 C5 C4 111.575
C3 C5 H11 123.341 C4 C2 H8 110.141
C4 C2 H13 113.209 C4 C5 H11 125.017
C5 C3 H9 110.141 C5 C3 H12 113.209
C5 C4 H10 125.017 H6 C1 H7 107.619
H8 C2 H13 106.813 H9 C3 H12 106.813
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability