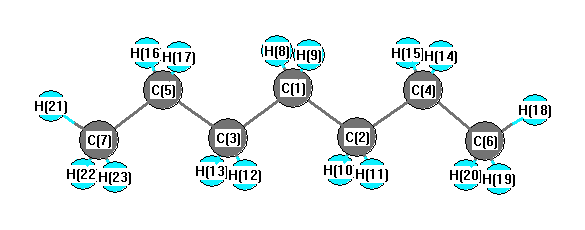
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3050 |
3032 |
37.87 |
|
|
|
| 2 |
A1 |
2964 |
2947 |
27.73 |
|
|
|
| 3 |
A1 |
2953 |
2936 |
135.38 |
|
|
|
| 4 |
A1 |
2942 |
2925 |
20.07 |
|
|
|
| 5 |
A1 |
2935 |
2918 |
0.45 |
|
|
|
| 6 |
A1 |
1438 |
1430 |
6.09 |
|
|
|
| 7 |
A1 |
1427 |
1419 |
0.74 |
|
|
|
| 8 |
A1 |
1413 |
1404 |
0.07 |
|
|
|
| 9 |
A1 |
1409 |
1401 |
0.02 |
|
|
|
| 10 |
A1 |
1349 |
1341 |
1.41 |
|
|
|
| 11 |
A1 |
1341 |
1334 |
0.10 |
|
|
|
| 12 |
A1 |
1258 |
1250 |
0.02 |
|
|
|
| 13 |
A1 |
1124 |
1118 |
0.82 |
|
|
|
| 14 |
A1 |
1063 |
1057 |
0.78 |
|
|
|
| 15 |
A1 |
993 |
987 |
0.01 |
|
|
|
| 16 |
A1 |
895 |
890 |
1.59 |
|
|
|
| 17 |
A1 |
404 |
402 |
0.07 |
|
|
|
| 18 |
A1 |
298 |
296 |
0.00 |
|
|
|
| 19 |
A1 |
94 |
94 |
0.01 |
|
|
|
| 20 |
A2 |
3041 |
3023 |
0.00 |
|
|
|
| 21 |
A2 |
2993 |
2976 |
0.00 |
|
|
|
| 22 |
A2 |
2968 |
2951 |
0.00 |
|
|
|
| 23 |
A2 |
1420 |
1412 |
0.00 |
|
|
|
| 24 |
A2 |
1279 |
1272 |
0.00 |
|
|
|
| 25 |
A2 |
1272 |
1265 |
0.00 |
|
|
|
| 26 |
A2 |
1188 |
1181 |
0.00 |
|
|
|
| 27 |
A2 |
988 |
982 |
0.00 |
|
|
|
| 28 |
A2 |
819 |
814 |
0.00 |
|
|
|
| 29 |
A2 |
724 |
719 |
0.00 |
|
|
|
| 30 |
A2 |
246 |
245 |
0.00 |
|
|
|
| 31 |
A2 |
147 |
146 |
0.00 |
|
|
|
| 32 |
A2 |
86 |
86 |
0.00 |
|
|
|
| 33 |
B1 |
3041 |
3023 |
93.43 |
|
|
|
| 34 |
B1 |
3002 |
2985 |
115.36 |
|
|
|
| 35 |
B1 |
2981 |
2964 |
0.00 |
|
|
|
| 36 |
B1 |
2961 |
2943 |
0.11 |
|
|
|
| 37 |
B1 |
1420 |
1412 |
12.67 |
|
|
|
| 38 |
B1 |
1286 |
1278 |
0.76 |
|
|
|
| 39 |
B1 |
1240 |
1233 |
0.04 |
|
|
|
| 40 |
B1 |
1153 |
1147 |
0.15 |
|
|
|
| 41 |
B1 |
905 |
900 |
1.01 |
|
|
|
| 42 |
B1 |
752 |
748 |
1.65 |
|
|
|
| 43 |
B1 |
722 |
718 |
6.23 |
|
|
|
| 44 |
B1 |
248 |
246 |
0.00 |
|
|
|
| 45 |
B1 |
154 |
153 |
0.00 |
|
|
|
| 46 |
B1 |
68 |
67 |
0.00 |
|
|
|
| 47 |
B2 |
3050 |
3032 |
29.05 |
|
|
|
| 48 |
B2 |
2964 |
2947 |
47.62 |
|
|
|
| 49 |
B2 |
2951 |
2934 |
0.03 |
|
|
|
| 50 |
B2 |
2936 |
2919 |
0.00 |
|
|
|
| 51 |
B2 |
1434 |
1425 |
0.72 |
|
|
|
| 52 |
B2 |
1418 |
1410 |
0.80 |
|
|
|
| 53 |
B2 |
1408 |
1400 |
0.38 |
|
|
|
| 54 |
B2 |
1353 |
1346 |
0.22 |
|
|
|
| 55 |
B2 |
1347 |
1339 |
1.42 |
|
|
|
| 56 |
B2 |
1309 |
1301 |
3.33 |
|
|
|
| 57 |
B2 |
1201 |
1194 |
6.46 |
|
|
|
| 58 |
B2 |
1068 |
1061 |
0.56 |
|
|
|
| 59 |
B2 |
1049 |
1043 |
1.64 |
|
|
|
| 60 |
B2 |
1038 |
1031 |
0.00 |
|
|
|
| 61 |
B2 |
868 |
863 |
2.40 |
|
|
|
| 62 |
B2 |
470 |
467 |
0.07 |
|
|
|
| 63 |
B2 |
235 |
234 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 46273.4 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 46005.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.035 |
|
|
|
| 2 |
C |
-0.038 |
|
|
|
| 3 |
C |
-0.038 |
|
|
|
| 4 |
C |
-0.069 |
|
|
|
| 5 |
C |
-0.069 |
|
|
|
| 6 |
C |
-0.045 |
|
|
|
| 7 |
C |
-0.045 |
|
|
|
| 8 |
H |
0.015 |
|
|
|
| 9 |
H |
0.015 |
|
|
|
| 10 |
H |
0.015 |
|
|
|
| 11 |
H |
0.015 |
|
|
|
| 12 |
H |
0.015 |
|
|
|
| 13 |
H |
0.015 |
|
|
|
| 14 |
H |
0.019 |
|
|
|
| 15 |
H |
0.019 |
|
|
|
| 16 |
H |
0.019 |
|
|
|
| 17 |
H |
0.019 |
|
|
|
| 18 |
H |
0.025 |
|
|
|
| 19 |
H |
0.030 |
|
|
|
| 20 |
H |
0.030 |
|
|
|
| 21 |
H |
0.025 |
|
|
|
| 22 |
H |
0.030 |
|
|
|
| 23 |
H |
0.030 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.064 |
0.064 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.853 |
0.000 |
0.000 |
| y |
0.000 |
-49.403 |
0.000 |
| z |
0.000 |
0.000 |
-48.999 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.348 |
0.000 |
0.000 |
| y |
0.000 |
-0.977 |
0.000 |
| z |
0.000 |
0.000 |
-0.371 |
|
| Polar |
|---|
| 3z2-r2 | -0.742 |
|---|
| x2-y2 | 1.550 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.800 |
0.000 |
0.000 |
| y |
0.000 |
15.628 |
0.000 |
| z |
0.000 |
0.000 |
11.391 |
<r2> (average value of r
2) Å
2
| <r2> |
480.736 |
| (<r2>)1/2 |
21.926 |