Jump to
S1C2
S1C3
Energy calculated at BLYP/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -255.126035 |
| Energy at 298.15K | -255.125325 |
| HF Energy | -255.126035 |
| Nuclear repulsion energy | 46.302953 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at BLYP/cc-pVDZ
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-0.680 |
| N2 |
0.000 |
0.000 |
-1.864 |
| Na3 |
0.000 |
0.000 |
1.557 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
Na3 |
| C1 | | 1.1839 | 2.2365 |
N2 | 1.1839 | | 3.4204 | Na3 | 2.2365 | 3.4204 | |
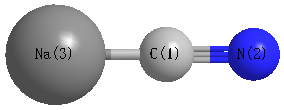 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
Na3 |
0.000 |
|
C1 |
Na3 |
N2 |
0.000 |
| N2 |
C1 |
Na3 |
180.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.362 |
|
|
|
| 2 |
N |
-0.179 |
|
|
|
| 3 |
Na |
0.541 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
9.750 |
9.750 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-17.578 |
0.000 |
0.000 |
| y |
0.000 |
-17.578 |
0.000 |
| z |
0.000 |
0.000 |
-12.666 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.456 |
0.000 |
0.000 |
| y |
0.000 |
-2.456 |
0.000 |
| z |
0.000 |
0.000 |
4.912 |
|
| Polar |
|---|
| 3z2-r2 | 9.824 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.416 |
0.000 |
0.000 |
| y |
0.000 |
3.416 |
0.000 |
| z |
0.000 |
0.000 |
7.261 |
<r2> (average value of r
2) Å
2
| <r2> |
63.698 |
| (<r2>)1/2 |
7.981 |
Jump to
S1C1
S1C3
Energy calculated at BLYP/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -255.132288 |
| Energy at 298.15K | -255.132012 |
| HF Energy | -255.132288 |
| Nuclear repulsion energy | 51.382470 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at BLYP/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.122 |
0.666 |
0.000 |
| N2 |
0.000 |
1.078 |
0.000 |
| Na3 |
-0.612 |
-1.049 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
Na3 |
| C1 | | 1.1955 | 2.4389 |
N2 | 1.1955 | | 2.2131 | Na3 | 2.4389 | 2.2131 | |
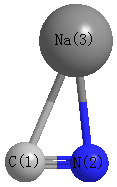 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
Na3 |
85.900 |
|
C1 |
Na3 |
N2 |
29.269 |
| N2 |
C1 |
Na3 |
64.831 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.210 |
|
|
|
| 2 |
N |
-0.190 |
|
|
|
| 3 |
Na |
0.400 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-4.533 |
-6.942 |
0.000 |
8.291 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-19.965 |
3.618 |
0.000 |
| y |
3.618 |
-14.143 |
0.000 |
| z |
0.000 |
0.000 |
-17.871 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.958 |
3.618 |
0.000 |
| y |
3.618 |
4.775 |
0.000 |
| z |
0.000 |
0.000 |
-0.817 |
|
| Polar |
|---|
| 3z2-r2 | -1.635 |
|---|
| x2-y2 | -5.822 |
|---|
| xy | 3.618 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.745 |
0.149 |
0.000 |
| y |
0.149 |
4.297 |
0.000 |
| z |
0.000 |
0.000 |
3.424 |
<r2> (average value of r
2) Å
2
| <r2> |
45.161 |
| (<r2>)1/2 |
6.720 |
Jump to
S1C1
S1C2
Energy calculated at BLYP/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -255.127819 |
| Energy at 298.15K | |
| HF Energy | -255.127819 |
| Nuclear repulsion energy | 48.378437 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at BLYP/cc-pVDZ
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.867 |
| N2 |
0.000 |
0.000 |
-0.671 |
| Na3 |
0.000 |
0.000 |
1.446 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
Na3 |
| C1 | | 1.1958 | 3.3126 |
N2 | 1.1958 | | 2.1168 | Na3 | 3.3126 | 2.1168 | |
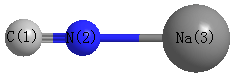 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
Na3 |
180.000 |
|
C1 |
Na3 |
N2 |
0.000 |
| N2 |
C1 |
Na3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.267 |
|
|
|
| 2 |
N |
-0.244 |
|
|
|
| 3 |
Na |
0.511 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
10.164 |
10.164 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-17.601 |
0.000 |
0.000 |
| y |
0.000 |
-17.601 |
0.000 |
| z |
0.000 |
0.000 |
-15.587 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.007 |
0.000 |
0.000 |
| y |
0.000 |
-1.007 |
0.000 |
| z |
0.000 |
0.000 |
2.014 |
|
| Polar |
|---|
| 3z2-r2 | 4.028 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.382 |
0.000 |
0.000 |
| y |
0.000 |
3.382 |
0.000 |
| z |
0.000 |
0.000 |
8.504 |
<r2> (average value of r
2) Å
2
| <r2> |
57.629 |
| (<r2>)1/2 |
7.591 |
