Jump to
S1C2
S1C3
Energy calculated at B3PW91/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -255.071206 |
| Energy at 298.15K | -255.070529 |
| HF Energy | -255.071206 |
| Nuclear repulsion energy | 46.470336 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at B3PW91/cc-pVDZ
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-0.685 |
| N2 |
0.000 |
0.000 |
-1.858 |
| Na3 |
0.000 |
0.000 |
1.556 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
Na3 |
| C1 | | 1.1728 | 2.2410 |
N2 | 1.1728 | | 3.4139 | Na3 | 2.2410 | 3.4139 | |
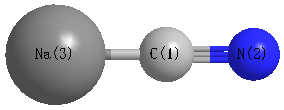 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
Na3 |
0.000 |
|
C1 |
Na3 |
N2 |
0.000 |
| N2 |
C1 |
Na3 |
180.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.417 |
|
|
|
| 2 |
N |
-0.220 |
|
|
|
| 3 |
Na |
0.637 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
10.408 |
10.408 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-17.110 |
0.000 |
0.000 |
| y |
0.000 |
-17.110 |
0.000 |
| z |
0.000 |
0.000 |
-12.338 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.386 |
0.000 |
0.000 |
| y |
0.000 |
-2.386 |
0.000 |
| z |
0.000 |
0.000 |
4.773 |
|
| Polar |
|---|
| 3z2-r2 | 9.545 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.860 |
0.000 |
0.000 |
| y |
0.000 |
2.860 |
0.000 |
| z |
0.000 |
0.000 |
5.309 |
<r2> (average value of r
2) Å
2
| <r2> |
63.303 |
| (<r2>)1/2 |
7.956 |
Jump to
S1C1
S1C3
Energy calculated at B3PW91/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -255.077519 |
| Energy at 298.15K | -255.077251 |
| HF Energy | -255.077519 |
| Nuclear repulsion energy | 51.633035 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at B3PW91/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.116 |
0.675 |
0.000 |
| N2 |
0.000 |
1.074 |
0.000 |
| Na3 |
-0.609 |
-1.052 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
Na3 |
| C1 | | 1.1850 | 2.4397 |
N2 | 1.1850 | | 2.2111 | Na3 | 2.4397 | 2.2111 | |
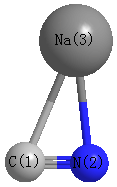 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
Na3 |
86.267 |
|
C1 |
Na3 |
N2 |
28.992 |
| N2 |
C1 |
Na3 |
64.741 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.253 |
|
|
|
| 2 |
N |
-0.255 |
|
|
|
| 3 |
Na |
0.508 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-4.684 |
-7.320 |
0.000 |
8.691 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-19.602 |
3.534 |
0.000 |
| y |
3.534 |
-13.803 |
0.000 |
| z |
0.000 |
0.000 |
-17.361 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.021 |
3.534 |
0.000 |
| y |
3.534 |
4.679 |
0.000 |
| z |
0.000 |
0.000 |
-0.658 |
|
| Polar |
|---|
| 3z2-r2 | -1.317 |
|---|
| x2-y2 | -5.800 |
|---|
| xy | 3.534 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.064 |
-0.132 |
0.000 |
| y |
-0.132 |
3.426 |
0.000 |
| z |
0.000 |
0.000 |
2.910 |
<r2> (average value of r
2) Å
2
| <r2> |
44.731 |
| (<r2>)1/2 |
6.688 |
Jump to
S1C1
S1C2
Energy calculated at B3PW91/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -255.073295 |
| Energy at 298.15K | |
| HF Energy | -255.073295 |
| Nuclear repulsion energy | 48.661229 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at B3PW91/cc-pVDZ
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.856 |
| N2 |
0.000 |
0.000 |
-0.671 |
| Na3 |
0.000 |
0.000 |
1.440 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
Na3 |
| C1 | | 1.1847 | 3.2956 |
N2 | 1.1847 | | 2.1109 | Na3 | 3.2956 | 2.1109 | |
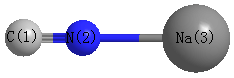 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
Na3 |
180.000 |
|
C1 |
Na3 |
N2 |
0.000 |
| N2 |
C1 |
Na3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.276 |
|
|
|
| 2 |
N |
-0.340 |
|
|
|
| 3 |
Na |
0.616 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
10.712 |
10.712 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-17.054 |
0.000 |
0.000 |
| y |
0.000 |
-17.054 |
0.000 |
| z |
0.000 |
0.000 |
-15.337 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.858 |
0.000 |
0.000 |
| y |
0.000 |
-0.858 |
0.000 |
| z |
0.000 |
0.000 |
1.717 |
|
| Polar |
|---|
| 3z2-r2 | 3.434 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.703 |
0.000 |
0.000 |
| y |
0.000 |
2.703 |
0.000 |
| z |
0.000 |
0.000 |
5.353 |
<r2> (average value of r
2) Å
2
| <r2> |
56.914 |
| (<r2>)1/2 |
7.544 |
