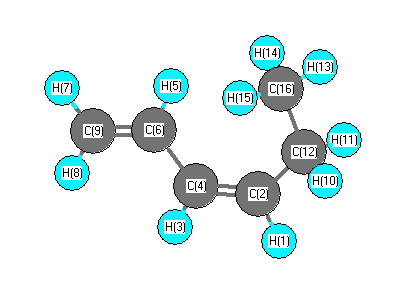
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3175 |
3156 |
13.88 |
|
|
|
| 2 |
A' |
3133 |
3115 |
3.98 |
|
|
|
| 3 |
A' |
3073 |
3055 |
29.66 |
|
|
|
| 4 |
A' |
3070 |
3053 |
7.40 |
|
|
|
| 5 |
A' |
3052 |
3034 |
32.83 |
|
|
|
| 6 |
A' |
3048 |
3030 |
3.65 |
|
|
|
| 7 |
A' |
2976 |
2959 |
18.97 |
|
|
|
| 8 |
A' |
2926 |
2909 |
29.06 |
|
|
|
| 9 |
A' |
1670 |
1661 |
15.43 |
|
|
|
| 10 |
A' |
1612 |
1603 |
2.23 |
|
|
|
| 11 |
A' |
1436 |
1427 |
7.69 |
|
|
|
| 12 |
A' |
1411 |
1402 |
8.77 |
|
|
|
| 13 |
A' |
1385 |
1377 |
14.57 |
|
|
|
| 14 |
A' |
1354 |
1346 |
2.51 |
|
|
|
| 15 |
A' |
1340 |
1333 |
1.06 |
|
|
|
| 16 |
A' |
1329 |
1321 |
6.03 |
|
|
|
| 17 |
A' |
1263 |
1255 |
0.70 |
|
|
|
| 18 |
A' |
1254 |
1247 |
12.31 |
|
|
|
| 19 |
A' |
1146 |
1139 |
0.84 |
|
|
|
| 20 |
A' |
1049 |
1043 |
5.93 |
|
|
|
| 21 |
A' |
1020 |
1014 |
1.98 |
|
|
|
| 22 |
A' |
914 |
908 |
3.97 |
|
|
|
| 23 |
A' |
803 |
798 |
7.27 |
|
|
|
| 24 |
A' |
700 |
696 |
1.40 |
|
|
|
| 25 |
A' |
413 |
410 |
0.35 |
|
|
|
| 26 |
A' |
274 |
272 |
2.42 |
|
|
|
| 27 |
A' |
180 |
179 |
0.36 |
|
|
|
| 28 |
A" |
3063 |
3045 |
20.04 |
|
|
|
| 29 |
A" |
2945 |
2928 |
15.62 |
|
|
|
| 30 |
A" |
1425 |
1417 |
5.63 |
|
|
|
| 31 |
A" |
1248 |
1240 |
0.21 |
|
|
|
| 32 |
A" |
1050 |
1044 |
3.94 |
|
|
|
| 33 |
A" |
1006 |
1001 |
9.27 |
|
|
|
| 34 |
A" |
951 |
946 |
1.97 |
|
|
|
| 35 |
A" |
875 |
870 |
32.19 |
|
|
|
| 36 |
A" |
807 |
802 |
1.56 |
|
|
|
| 37 |
A" |
737 |
733 |
1.73 |
|
|
|
| 38 |
A" |
611 |
607 |
21.99 |
|
|
|
| 39 |
A" |
376 |
374 |
0.00 |
|
|
|
| 40 |
A" |
291 |
290 |
0.00 |
|
|
|
| 41 |
A" |
131 |
130 |
0.22 |
|
|
|
| 42 |
A" |
81 |
80 |
0.47 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30299.4 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 30123.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
-0.003 |
|
|
|
| 2 |
C |
-0.051 |
|
|
|
| 3 |
H |
-0.021 |
|
|
|
| 4 |
C |
0.005 |
|
|
|
| 5 |
H |
-0.026 |
|
|
|
| 6 |
C |
-0.041 |
|
|
|
| 7 |
H |
0.021 |
|
|
|
| 8 |
H |
0.017 |
|
|
|
| 9 |
C |
-0.003 |
|
|
|
| 10 |
H |
0.042 |
|
|
|
| 11 |
H |
0.042 |
|
|
|
| 12 |
C |
-0.053 |
|
|
|
| 13 |
H |
0.026 |
|
|
|
| 14 |
H |
0.050 |
|
|
|
| 15 |
H |
0.050 |
|
|
|
| 16 |
C |
-0.055 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.646 |
-0.573 |
0.000 |
0.863 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.142 |
0.302 |
0.000 |
| y |
0.302 |
-36.675 |
0.000 |
| z |
0.000 |
0.000 |
-41.026 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.708 |
0.302 |
0.000 |
| y |
0.302 |
1.909 |
0.000 |
| z |
0.000 |
0.000 |
-4.617 |
|
| Polar |
|---|
| 3z2-r2 | -9.235 |
|---|
| x2-y2 | 0.533 |
|---|
| xy | 0.302 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
16.146 |
2.789 |
0.000 |
| y |
2.789 |
11.182 |
0.000 |
| z |
0.000 |
0.000 |
5.866 |
<r2> (average value of r
2) Å
2
| <r2> |
221.920 |
| (<r2>)1/2 |
14.897 |