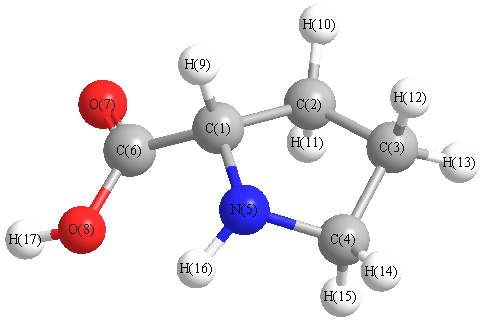
Vibrational Frequencies calculated at B3PW91/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3733 |
3603 |
69.20 |
|
|
|
| 2 |
A |
3551 |
3426 |
6.80 |
|
|
|
| 3 |
A |
3148 |
3038 |
16.79 |
|
|
|
| 4 |
A |
3124 |
3015 |
12.10 |
|
|
|
| 5 |
A |
3097 |
2988 |
23.58 |
|
|
|
| 6 |
A |
3090 |
2982 |
50.45 |
|
|
|
| 7 |
A |
3072 |
2964 |
15.83 |
|
|
|
| 8 |
A |
3032 |
2926 |
21.07 |
|
|
|
| 9 |
A |
2939 |
2836 |
79.50 |
|
|
|
| 10 |
A |
1846 |
1782 |
246.22 |
|
|
|
| 11 |
A |
1498 |
1445 |
0.77 |
|
|
|
| 12 |
A |
1479 |
1428 |
5.90 |
|
|
|
| 13 |
A |
1459 |
1408 |
1.54 |
|
|
|
| 14 |
A |
1444 |
1394 |
20.63 |
|
|
|
| 15 |
A |
1371 |
1323 |
31.95 |
|
|
|
| 16 |
A |
1359 |
1311 |
6.68 |
|
|
|
| 17 |
A |
1324 |
1278 |
11.71 |
|
|
|
| 18 |
A |
1314 |
1268 |
9.75 |
|
|
|
| 19 |
A |
1301 |
1255 |
1.88 |
|
|
|
| 20 |
A |
1249 |
1205 |
7.49 |
|
|
|
| 21 |
A |
1235 |
1192 |
2.67 |
|
|
|
| 22 |
A |
1212 |
1170 |
16.38 |
|
|
|
| 23 |
A |
1193 |
1152 |
14.72 |
|
|
|
| 24 |
A |
1172 |
1131 |
199.74 |
|
|
|
| 25 |
A |
1156 |
1116 |
33.98 |
|
|
|
| 26 |
A |
1100 |
1062 |
1.56 |
|
|
|
| 27 |
A |
1082 |
1044 |
4.10 |
|
|
|
| 28 |
A |
1008 |
972 |
4.12 |
|
|
|
| 29 |
A |
972 |
938 |
6.98 |
|
|
|
| 30 |
A |
939 |
906 |
3.91 |
|
|
|
| 31 |
A |
923 |
891 |
2.49 |
|
|
|
| 32 |
A |
895 |
863 |
55.77 |
|
|
|
| 33 |
A |
845 |
815 |
26.32 |
|
|
|
| 34 |
A |
786 |
759 |
3.18 |
|
|
|
| 35 |
A |
748 |
722 |
37.32 |
|
|
|
| 36 |
A |
676 |
652 |
91.96 |
|
|
|
| 37 |
A |
624 |
602 |
47.58 |
|
|
|
| 38 |
A |
590 |
569 |
13.41 |
|
|
|
| 39 |
A |
510 |
492 |
31.99 |
|
|
|
| 40 |
A |
495 |
478 |
14.13 |
|
|
|
| 41 |
A |
350 |
338 |
2.13 |
|
|
|
| 42 |
A |
260 |
251 |
3.12 |
|
|
|
| 43 |
A |
187 |
181 |
0.18 |
|
|
|
| 44 |
A |
62 |
60 |
0.29 |
|
|
|
| 45 |
A |
44 |
42 |
1.70 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31747.1 cm
-1
Scaled (by 0.965) Zero Point Vibrational Energy (zpe) 30636.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.073 |
|
|
|
| 2 |
C |
-0.012 |
|
|
|
| 3 |
C |
-0.052 |
|
|
|
| 4 |
C |
0.026 |
|
|
|
| 5 |
N |
-0.206 |
|
|
|
| 6 |
C |
0.216 |
|
|
|
| 7 |
O |
-0.250 |
|
|
|
| 8 |
O |
-0.188 |
|
|
|
| 9 |
H |
0.056 |
|
|
|
| 10 |
H |
0.041 |
|
|
|
| 11 |
H |
0.052 |
|
|
|
| 12 |
H |
0.044 |
|
|
|
| 13 |
H |
0.033 |
|
|
|
| 14 |
H |
0.035 |
|
|
|
| 15 |
H |
0.020 |
|
|
|
| 16 |
H |
0.095 |
|
|
|
| 17 |
H |
0.162 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.026 |
-1.358 |
-0.351 |
1.403 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-45.578 |
5.661 |
-0.920 |
| y |
5.661 |
-48.955 |
2.467 |
| z |
-0.920 |
2.467 |
-46.915 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.357 |
5.661 |
-0.920 |
| y |
5.661 |
-2.709 |
2.467 |
| z |
-0.920 |
2.467 |
0.352 |
|
| Polar |
|---|
| 3z2-r2 | 0.704 |
|---|
| x2-y2 | 3.378 |
|---|
| xy | 5.661 |
|---|
| xz | -0.920 |
|---|
| yz | 2.467 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.257 |
0.068 |
0.057 |
| y |
0.068 |
9.634 |
0.216 |
| z |
0.057 |
0.216 |
8.017 |
<r2> (average value of r
2) Å
2
| <r2> |
258.724 |
| (<r2>)1/2 |
16.085 |