return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H9NO2 (Proline)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-400.079256
Energy at 298.15K-400.091810
HF Energy-398.795523
Nuclear repulsion energy397.188446
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3789 3635 68.62      
2 A 3564 3419 4.75      
3 A 3167 3038 18.81      
4 A 3140 3013 16.10      
5 A 3120 2993 33.66      
6 A 3110 2984 36.55      
7 A 3088 2963 22.52      
8 A 3074 2949 18.76      
9 A 2980 2859 68.51      
10 A 1856 1781 221.61      
11 A 1527 1465 0.86      
12 A 1508 1447 4.17      
13 A 1484 1424 0.74      
14 A 1467 1407 21.29      
15 A 1403 1346 25.94      
16 A 1386 1329 5.72      
17 A 1344 1290 10.99      
18 A 1337 1283 5.64      
19 A 1324 1270 1.94      
20 A 1276 1224 12.64      
21 A 1258 1207 0.67      
22 A 1233 1183 12.42      
23 A 1215 1166 45.13      
24 A 1195 1146 195.72      
25 A 1164 1117 10.60      
26 A 1115 1070 2.56      
27 A 1092 1048 4.25      
28 A 1015 974 4.16      
29 A 982 942 11.21      
30 A 946 908 7.51      
31 A 935 897 30.96      
32 A 921 884 35.69      
33 A 858 823 14.09      
34 A 790 758 1.91      
35 A 764 733 34.24      
36 A 681 653 88.61      
37 A 626 600 49.18      
38 A 597 572 13.82      
39 A 512 491 20.75      
40 A 509 488 22.28      
41 A 367 352 1.72      
42 A 270 259 2.70      
43 A 189 181 0.36      
44 A 57 55 0.09      
45 A 41 39 1.40      

Unscaled Zero Point Vibrational Energy (zpe) 32136.0 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 30831.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.11697 0.05795 0.04682

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.051 0.149 0.809
C2 0.921 1.259 0.334
C3 2.008 0.499 -0.483
C4 1.470 -0.943 -0.574
N5 0.732 -1.077 0.686
C6 -1.361 0.175 0.001
O7 -1.903 1.172 -0.421
O8 -1.862 -1.073 -0.160
H9 -0.359 0.301 1.862
H10 1.363 1.776 1.201
H11 0.382 2.009 -0.266
H12 2.971 0.507 0.055
H13 2.171 0.944 -1.480
H14 2.273 -1.699 -0.636
H15 0.824 -1.057 -1.476
H16 0.140 -1.907 0.706
H17 -2.695 -0.959 -0.643

Atom - Atom Distances (Å)
  C1 C2 C3 C4 N5 C6 O7 O8 H9 H10 H11 H12 H13 H14 H15 H16 H17
C11.54962.45542.32801.46021.53982.44772.39021.10752.19052.19133.13523.28723.30202.72802.06713.2141
C21.54961.55722.44422.36992.54802.92473.66382.21171.10261.10172.20142.22483.39312.94053.28144.3531
C32.45541.55721.54192.34073.41843.96904.18903.33772.20982.22981.10381.10362.21872.19273.26954.9263
C42.32802.44421.54191.46663.09763.98483.36023.29073.24923.16172.18012.20741.10401.11532.08244.1659
N51.46022.36992.34071.46662.53363.63742.72872.11492.96703.24892.81473.29342.12362.16431.01933.6781
C61.53982.54803.41843.09762.53361.21071.35412.11743.37982.54374.34523.90574.13782.91082.66151.8657
O72.44772.92473.96903.98483.63741.21072.26012.89033.69662.43824.94284.21575.07233.67713.86302.2842
O82.39023.66384.18903.36022.72871.35412.26012.86944.51303.81335.08954.69794.20922.99112.33530.9698
H91.10752.21173.33773.29072.11492.11742.89032.86942.36172.82743.79474.24044.14343.79292.54123.6496
H102.19051.10262.20983.24922.96703.37983.69664.51302.36171.78072.34752.92134.03443.93503.91175.2297
H112.19131.10172.22983.16173.24892.54372.43823.81332.82741.78073.01112.41034.17873.32584.04184.2915
H123.13522.20141.10382.18012.81474.34524.94285.08953.79472.34753.01111.78542.41453.06623.77725.8946
H133.28722.22481.10362.20743.29343.90574.21574.69794.24042.92132.41031.78542.77552.41184.12655.2913
H143.30203.39312.21871.10402.12364.13785.07234.20924.14344.03444.17872.41452.77551.79332.52915.0230
H152.72802.94052.19271.11532.16432.91083.67712.99113.79293.93503.32583.06622.41181.79332.43973.6179
H162.06713.28143.26952.08241.01932.66153.86302.33532.54123.91174.04183.77724.12652.52912.43973.2797
H173.21414.35314.92634.16593.67811.86572.28420.96983.64965.22974.29155.89465.29135.02303.61793.2797

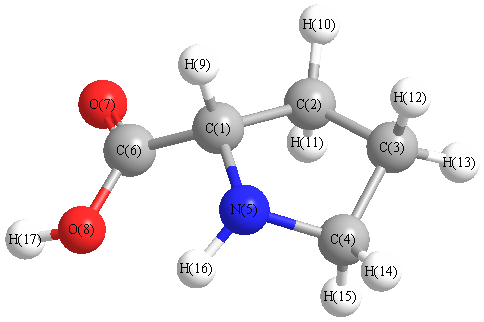
picture of Proline state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 104.433 C1 C2 H10 110.222
C1 C2 H11 110.338 C1 N5 C4 105.388
C1 N5 H16 111.730 C1 C6 O7 125.303
C1 C6 O8 111.204 C2 C1 N5 103.850
C2 C1 C6 111.124 C2 C1 H9 111.610
C2 C3 C4 104.122 C2 C3 H12 110.483
C2 C3 H13 112.348 C3 C2 H10 111.219
C3 C2 H11 112.864 C3 C4 N5 102.135
C3 C4 H14 112.931 C3 C4 H15 110.182
C4 C3 H12 109.871 C4 C3 H13 112.042
C4 N5 H16 112.554 N5 C1 C6 115.221
N5 C1 H9 110.149 N5 C4 H14 110.613
N5 C4 H15 113.216 C6 C1 H9 105.057
C6 O8 H17 105.608 O7 C6 O8 123.476
H10 C2 H11 107.767 H12 C3 H13 107.963
H14 C4 H15 107.806
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability