return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H9NO2 (Proline)

using model chemistry: CCD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at CCD/cc-pVDZ
 hartrees
Energy at 0K-400.066097
Energy at 298.15K-400.078678
HF Energy-398.796507
Nuclear repulsion energy397.554466
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3823 3657 74.09      
2 A 3579 3424 5.51      
3 A 3178 3040 17.50      
4 A 3152 3016 15.00      
5 A 3130 2995 33.34      
6 A 3120 2985 33.57      
7 A 3098 2964 21.22      
8 A 3082 2948 18.69      
9 A 2994 2865 64.97      
10 A 1906 1824 236.16      
11 A 1531 1464 0.83      
12 A 1511 1445 3.99      
13 A 1488 1423 1.38      
14 A 1473 1409 22.93      
15 A 1415 1353 31.70      
16 A 1392 1331 5.88      
17 A 1349 1291 12.81      
18 A 1343 1285 3.81      
19 A 1328 1271 2.15      
20 A 1282 1226 16.74      
21 A 1262 1207 0.75      
22 A 1238 1185 16.01      
23 A 1223 1170 110.03      
24 A 1206 1154 134.89      
25 A 1170 1120 9.65      
26 A 1118 1070 2.15      
27 A 1097 1049 3.61      
28 A 1020 975 3.60      
29 A 988 946 10.59      
30 A 951 909 8.35      
31 A 939 898 30.70      
32 A 926 886 34.38      
33 A 862 825 13.99      
34 A 793 759 1.94      
35 A 771 737 35.71      
36 A 682 652 85.65      
37 A 629 602 54.85      
38 A 600 574 13.38      
39 A 519 496 16.26      
40 A 511 489 27.00      
41 A 369 353 1.73      
42 A 272 260 2.72      
43 A 188 180 0.36      
44 A 55 53 0.09      
45 A 41 39 1.39      

Unscaled Zero Point Vibrational Energy (zpe) 32299.3 cm-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 30900.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/cc-pVDZ
ABC
0.11734 0.05797 0.04687

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/cc-pVDZ

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.051 0.150 0.808
C2 0.922 1.258 0.333
C3 2.008 0.496 -0.483
C4 1.468 -0.945 -0.573
N5 0.730 -1.077 0.686
C6 -1.360 0.176 0.001
O7 -1.898 1.170 -0.421
O8 -1.864 -1.068 -0.159
H9 -0.359 0.303 1.861
H10 1.365 1.774 1.201
H11 0.385 2.009 -0.266
H12 2.971 0.503 0.056
H13 2.172 0.940 -1.479
H14 2.270 -1.701 -0.635
H15 0.822 -1.058 -1.474
H16 0.136 -1.905 0.706
H17 -2.696 -0.955 -0.642

Atom - Atom Distances (Å)
  C1 C2 C3 C4 N5 C6 O7 O8 H9 H10 H11 H12 H13 H14 H15 H16 H17
C11.54942.45462.32661.45931.53842.44222.38861.10722.19032.19133.13403.28653.30042.72612.06583.2123
C21.54941.55682.44352.36932.54762.92123.66252.21101.10231.10142.20062.22443.39202.93983.28024.3519
C32.45461.55681.54172.34013.41723.96414.18793.33662.20912.22921.10351.10322.21802.19243.26834.9248
C42.32662.44351.54171.46583.09543.97813.35933.28913.24803.16122.17952.20731.10371.11472.08104.1641
N51.45932.36932.34011.46582.53123.63062.72792.11402.96603.24832.81363.29272.12282.16271.01873.6761
C61.53842.54763.41723.09542.53121.20631.35142.11583.37932.54474.34343.90534.13522.90842.65841.8645
O72.44222.92123.96413.97813.63061.20632.25302.88553.69402.43744.93764.21215.06533.67053.85542.2801
O82.38863.66254.18793.35932.72791.35142.25302.86734.51123.81245.08784.69734.20842.99012.33460.9685
H91.10722.21103.33663.28912.11402.11582.88552.86732.36112.82663.79324.23934.14183.79072.54023.6473
H102.19031.10232.20913.24802.96603.37933.69404.51122.36111.78002.34622.92044.03273.93373.91025.2282
H112.19131.10142.22923.16123.24832.54472.43743.81242.82661.78003.01002.41014.17763.32574.04074.2914
H123.13402.20061.10352.17952.81364.34344.93765.08783.79322.34623.01001.78482.41363.06553.77555.8925
H133.28652.22441.10322.20733.29273.90534.21214.69734.23932.92042.41011.78482.77472.41244.12525.2907
H143.30043.39202.21801.10372.12284.13525.06534.20844.14184.03274.17762.41362.77471.79262.52795.0211
H152.72612.93982.19241.11472.16272.90843.67052.99013.79073.93373.32573.06552.41241.79262.43723.6161
H162.06583.28023.26832.08101.01872.65843.85542.33462.54023.91024.04073.77554.12522.52792.43723.2774
H173.21234.35194.92484.16413.67611.86452.28010.96853.64735.22824.29145.89255.29075.02113.61613.2774

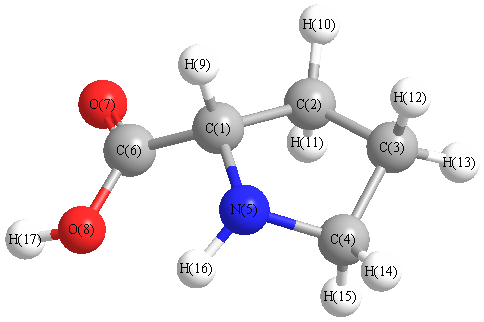
picture of Proline state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 104.413 C1 C2 H10 110.237
C1 C2 H11 110.367 C1 N5 C4 105.387
C1 N5 H16 111.721 C1 C6 O7 125.265
C1 C6 O8 111.328 C2 C1 N5 103.859
C2 C1 C6 111.187 C2 C1 H9 111.577
C2 C3 C4 104.109 C2 C3 H12 110.462
C2 C3 H13 112.369 C3 C2 H10 111.203
C3 C2 H11 112.867 C3 C4 N5 102.147
C3 C4 H14 112.906 C3 C4 H15 110.211
C4 C3 H12 109.860 C4 C3 H13 112.074
C4 N5 H16 112.526 N5 C1 C6 115.192
N5 C1 H9 110.154 N5 C4 H14 110.626
N5 C4 H15 113.184 C6 C1 H9 105.040
C6 O8 H17 105.784 O7 C6 O8 123.391
H10 C2 H11 107.757 H12 C3 H13 107.953
H14 C4 H15 107.808
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability