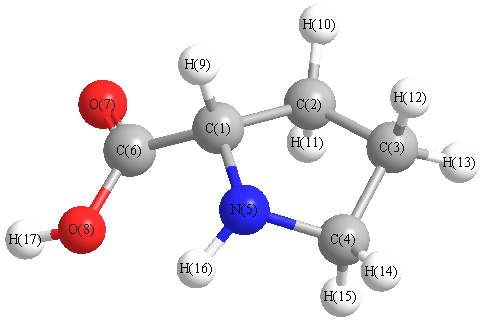
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3576 |
3555 |
41.65 |
|
|
|
| 2 |
A |
3432 |
3412 |
3.69 |
|
|
|
| 3 |
A |
3071 |
3053 |
18.32 |
|
|
|
| 4 |
A |
3049 |
3032 |
9.76 |
|
|
|
| 5 |
A |
3021 |
3003 |
23.63 |
|
|
|
| 6 |
A |
3013 |
2995 |
50.44 |
|
|
|
| 7 |
A |
2998 |
2981 |
13.43 |
|
|
|
| 8 |
A |
2953 |
2936 |
20.72 |
|
|
|
| 9 |
A |
2850 |
2834 |
82.74 |
|
|
|
| 10 |
A |
1774 |
1764 |
210.98 |
|
|
|
| 11 |
A |
1445 |
1437 |
0.83 |
|
|
|
| 12 |
A |
1429 |
1420 |
5.96 |
|
|
|
| 13 |
A |
1409 |
1401 |
0.82 |
|
|
|
| 14 |
A |
1392 |
1384 |
15.60 |
|
|
|
| 15 |
A |
1315 |
1308 |
16.36 |
|
|
|
| 16 |
A |
1309 |
1301 |
7.50 |
|
|
|
| 17 |
A |
1278 |
1271 |
8.18 |
|
|
|
| 18 |
A |
1265 |
1258 |
8.37 |
|
|
|
| 19 |
A |
1254 |
1247 |
1.87 |
|
|
|
| 20 |
A |
1201 |
1194 |
2.25 |
|
|
|
| 21 |
A |
1190 |
1183 |
2.48 |
|
|
|
| 22 |
A |
1164 |
1158 |
16.95 |
|
|
|
| 23 |
A |
1152 |
1146 |
4.66 |
|
|
|
| 24 |
A |
1118 |
1112 |
83.67 |
|
|
|
| 25 |
A |
1109 |
1103 |
144.38 |
|
|
|
| 26 |
A |
1064 |
1058 |
3.54 |
|
|
|
| 27 |
A |
1051 |
1044 |
5.36 |
|
|
|
| 28 |
A |
980 |
974 |
4.24 |
|
|
|
| 29 |
A |
941 |
935 |
6.24 |
|
|
|
| 30 |
A |
912 |
907 |
3.53 |
|
|
|
| 31 |
A |
896 |
891 |
4.63 |
|
|
|
| 32 |
A |
869 |
864 |
57.73 |
|
|
|
| 33 |
A |
817 |
812 |
20.63 |
|
|
|
| 34 |
A |
757 |
753 |
1.88 |
|
|
|
| 35 |
A |
722 |
718 |
34.55 |
|
|
|
| 36 |
A |
663 |
659 |
87.42 |
|
|
|
| 37 |
A |
603 |
600 |
36.85 |
|
|
|
| 38 |
A |
573 |
570 |
12.69 |
|
|
|
| 39 |
A |
496 |
493 |
32.07 |
|
|
|
| 40 |
A |
474 |
471 |
12.08 |
|
|
|
| 41 |
A |
342 |
340 |
2.12 |
|
|
|
| 42 |
A |
252 |
251 |
2.78 |
|
|
|
| 43 |
A |
186 |
185 |
0.18 |
|
|
|
| 44 |
A |
67 |
66 |
0.23 |
|
|
|
| 45 |
A |
46 |
45 |
1.56 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30738.8 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 30560.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.068 |
|
|
|
| 2 |
C |
0.016 |
|
|
|
| 3 |
C |
-0.024 |
|
|
|
| 4 |
C |
0.032 |
|
|
|
| 5 |
N |
-0.157 |
|
|
|
| 6 |
C |
0.159 |
|
|
|
| 7 |
O |
-0.212 |
|
|
|
| 8 |
O |
-0.152 |
|
|
|
| 9 |
H |
0.038 |
|
|
|
| 10 |
H |
0.027 |
|
|
|
| 11 |
H |
0.034 |
|
|
|
| 12 |
H |
0.031 |
|
|
|
| 13 |
H |
0.019 |
|
|
|
| 14 |
H |
0.020 |
|
|
|
| 15 |
H |
0.007 |
|
|
|
| 16 |
H |
0.081 |
|
|
|
| 17 |
H |
0.148 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.032 |
-1.321 |
-0.334 |
1.363 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.106 |
5.501 |
-0.817 |
| y |
5.501 |
-49.067 |
2.274 |
| z |
-0.817 |
2.274 |
-47.316 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.085 |
5.501 |
-0.817 |
| y |
5.501 |
-2.356 |
2.274 |
| z |
-0.817 |
2.274 |
0.271 |
|
| Polar |
|---|
| 3z2-r2 | 0.542 |
|---|
| x2-y2 | 2.961 |
|---|
| xy | 5.501 |
|---|
| xz | -0.817 |
|---|
| yz | 2.274 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.824 |
0.129 |
0.105 |
| y |
0.129 |
10.072 |
0.272 |
| z |
0.105 |
0.272 |
8.372 |
<r2> (average value of r
2) Å
2
| <r2> |
260.504 |
| (<r2>)1/2 |
16.140 |