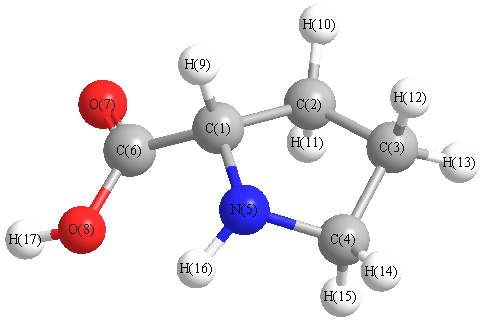
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3573 |
3553 |
41.66 |
|
|
|
| 2 |
A |
3434 |
3414 |
3.74 |
|
|
|
| 3 |
A |
3071 |
3053 |
18.48 |
|
|
|
| 4 |
A |
3049 |
3031 |
9.98 |
|
|
|
| 5 |
A |
3021 |
3003 |
22.93 |
|
|
|
| 6 |
A |
3013 |
2996 |
50.00 |
|
|
|
| 7 |
A |
2997 |
2980 |
14.23 |
|
|
|
| 8 |
A |
2953 |
2936 |
20.80 |
|
|
|
| 9 |
A |
2851 |
2835 |
83.01 |
|
|
|
| 10 |
A |
1774 |
1764 |
210.88 |
|
|
|
| 11 |
A |
1444 |
1436 |
0.94 |
|
|
|
| 12 |
A |
1429 |
1420 |
5.75 |
|
|
|
| 13 |
A |
1408 |
1399 |
1.01 |
|
|
|
| 14 |
A |
1392 |
1384 |
15.41 |
|
|
|
| 15 |
A |
1315 |
1308 |
15.60 |
|
|
|
| 16 |
A |
1309 |
1301 |
8.08 |
|
|
|
| 17 |
A |
1277 |
1270 |
8.42 |
|
|
|
| 18 |
A |
1265 |
1258 |
8.01 |
|
|
|
| 19 |
A |
1255 |
1248 |
1.92 |
|
|
|
| 20 |
A |
1202 |
1195 |
2.13 |
|
|
|
| 21 |
A |
1191 |
1184 |
2.56 |
|
|
|
| 22 |
A |
1165 |
1158 |
16.72 |
|
|
|
| 23 |
A |
1152 |
1146 |
4.87 |
|
|
|
| 24 |
A |
1119 |
1112 |
79.53 |
|
|
|
| 25 |
A |
1110 |
1103 |
149.68 |
|
|
|
| 26 |
A |
1064 |
1057 |
3.50 |
|
|
|
| 27 |
A |
1050 |
1044 |
5.33 |
|
|
|
| 28 |
A |
980 |
974 |
4.06 |
|
|
|
| 29 |
A |
941 |
935 |
6.26 |
|
|
|
| 30 |
A |
912 |
907 |
3.49 |
|
|
|
| 31 |
A |
896 |
891 |
4.10 |
|
|
|
| 32 |
A |
869 |
863 |
57.94 |
|
|
|
| 33 |
A |
816 |
811 |
20.39 |
|
|
|
| 34 |
A |
758 |
753 |
1.94 |
|
|
|
| 35 |
A |
722 |
718 |
34.99 |
|
|
|
| 36 |
A |
663 |
659 |
87.75 |
|
|
|
| 37 |
A |
603 |
600 |
36.80 |
|
|
|
| 38 |
A |
572 |
569 |
13.09 |
|
|
|
| 39 |
A |
496 |
494 |
31.29 |
|
|
|
| 40 |
A |
474 |
471 |
12.04 |
|
|
|
| 41 |
A |
342 |
340 |
2.14 |
|
|
|
| 42 |
A |
252 |
251 |
2.82 |
|
|
|
| 43 |
A |
185 |
184 |
0.17 |
|
|
|
| 44 |
A |
65 |
65 |
0.24 |
|
|
|
| 45 |
A |
44 |
44 |
1.55 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30735.2 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 30556.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.067 |
|
|
|
| 2 |
C |
0.016 |
|
|
|
| 3 |
C |
-0.024 |
|
|
|
| 4 |
C |
0.032 |
|
|
|
| 5 |
N |
-0.157 |
|
|
|
| 6 |
C |
0.160 |
|
|
|
| 7 |
O |
-0.212 |
|
|
|
| 8 |
O |
-0.152 |
|
|
|
| 9 |
H |
0.038 |
|
|
|
| 10 |
H |
0.027 |
|
|
|
| 11 |
H |
0.034 |
|
|
|
| 12 |
H |
0.031 |
|
|
|
| 13 |
H |
0.019 |
|
|
|
| 14 |
H |
0.020 |
|
|
|
| 15 |
H |
0.007 |
|
|
|
| 16 |
H |
0.081 |
|
|
|
| 17 |
H |
0.149 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.035 |
-1.319 |
-0.334 |
1.361 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.074 |
5.496 |
-0.818 |
| y |
5.496 |
-49.084 |
2.292 |
| z |
-0.818 |
2.292 |
-47.323 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.129 |
5.496 |
-0.818 |
| y |
5.496 |
-2.385 |
2.292 |
| z |
-0.818 |
2.292 |
0.256 |
|
| Polar |
|---|
| 3z2-r2 | 0.512 |
|---|
| x2-y2 | 3.010 |
|---|
| xy | 5.496 |
|---|
| xz | -0.818 |
|---|
| yz | 2.292 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.829 |
0.132 |
0.105 |
| y |
0.132 |
10.069 |
0.269 |
| z |
0.105 |
0.269 |
8.371 |
<r2> (average value of r
2) Å
2
| <r2> |
260.636 |
| (<r2>)1/2 |
16.144 |