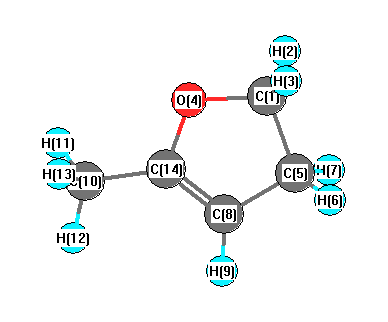
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3179 |
3160 |
3.85 |
|
|
|
| 2 |
A |
3090 |
3072 |
9.48 |
|
|
|
| 3 |
A |
3040 |
3023 |
32.87 |
|
|
|
| 4 |
A |
3034 |
3016 |
10.36 |
|
|
|
| 5 |
A |
2999 |
2982 |
33.37 |
|
|
|
| 6 |
A |
2971 |
2954 |
25.47 |
|
|
|
| 7 |
A |
2948 |
2930 |
55.34 |
|
|
|
| 8 |
A |
2926 |
2909 |
55.90 |
|
|
|
| 9 |
A |
1688 |
1678 |
51.86 |
|
|
|
| 10 |
A |
1436 |
1428 |
1.42 |
|
|
|
| 11 |
A |
1415 |
1406 |
0.68 |
|
|
|
| 12 |
A |
1410 |
1401 |
2.97 |
|
|
|
| 13 |
A |
1388 |
1380 |
7.93 |
|
|
|
| 14 |
A |
1359 |
1351 |
31.61 |
|
|
|
| 15 |
A |
1336 |
1329 |
2.07 |
|
|
|
| 16 |
A |
1271 |
1264 |
11.29 |
|
|
|
| 17 |
A |
1213 |
1206 |
55.75 |
|
|
|
| 18 |
A |
1206 |
1199 |
1.82 |
|
|
|
| 19 |
A |
1165 |
1159 |
16.30 |
|
|
|
| 20 |
A |
1153 |
1146 |
2.54 |
|
|
|
| 21 |
A |
1032 |
1026 |
0.51 |
|
|
|
| 22 |
A |
1008 |
1002 |
5.19 |
|
|
|
| 23 |
A |
999 |
993 |
25.96 |
|
|
|
| 24 |
A |
987 |
981 |
20.06 |
|
|
|
| 25 |
A |
950 |
944 |
24.44 |
|
|
|
| 26 |
A |
923 |
918 |
7.40 |
|
|
|
| 27 |
A |
889 |
884 |
21.83 |
|
|
|
| 28 |
A |
827 |
823 |
4.57 |
|
|
|
| 29 |
A |
689 |
685 |
20.15 |
|
|
|
| 30 |
A |
665 |
661 |
3.03 |
|
|
|
| 31 |
A |
622 |
618 |
0.91 |
|
|
|
| 32 |
A |
538 |
534 |
1.17 |
|
|
|
| 33 |
A |
326 |
324 |
2.07 |
|
|
|
| 34 |
A |
218 |
217 |
6.35 |
|
|
|
| 35 |
A |
162 |
161 |
2.37 |
|
|
|
| 36 |
A |
132 |
132 |
0.89 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25595.4 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 25446.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.106 |
|
|
|
| 2 |
H |
0.032 |
|
|
|
| 3 |
H |
0.034 |
|
|
|
| 4 |
O |
-0.258 |
|
|
|
| 5 |
C |
-0.011 |
|
|
|
| 6 |
H |
0.031 |
|
|
|
| 7 |
H |
0.037 |
|
|
|
| 8 |
C |
-0.145 |
|
|
|
| 9 |
H |
-0.031 |
|
|
|
| 10 |
C |
0.008 |
|
|
|
| 11 |
H |
0.053 |
|
|
|
| 12 |
H |
0.040 |
|
|
|
| 13 |
H |
0.052 |
|
|
|
| 14 |
C |
0.052 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.296 |
0.456 |
0.110 |
0.555 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.518 |
1.091 |
-0.041 |
| y |
1.091 |
-37.489 |
-0.159 |
| z |
-0.041 |
-0.159 |
-37.388 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.920 |
1.091 |
-0.041 |
| y |
1.091 |
-2.536 |
-0.159 |
| z |
-0.041 |
-0.159 |
-2.384 |
|
| Polar |
|---|
| 3z2-r2 | -4.768 |
|---|
| x2-y2 | 4.971 |
|---|
| xy | 1.091 |
|---|
| xz | -0.041 |
|---|
| yz | -0.159 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.665 |
-0.520 |
0.029 |
| y |
-0.520 |
8.833 |
-0.022 |
| z |
0.029 |
-0.022 |
6.194 |
<r2> (average value of r
2) Å
2
| <r2> |
157.579 |
| (<r2>)1/2 |
12.553 |