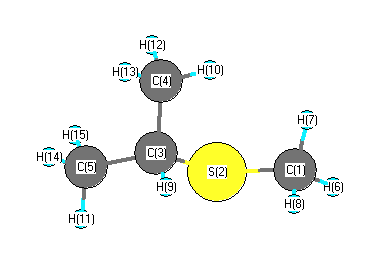
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3080 |
3062 |
5.05 |
|
|
|
| 2 |
A |
3065 |
3048 |
16.32 |
|
|
|
| 3 |
A |
3059 |
3041 |
19.52 |
|
|
|
| 4 |
A |
3054 |
3037 |
13.12 |
|
|
|
| 5 |
A |
3047 |
3029 |
25.26 |
|
|
|
| 6 |
A |
3038 |
3021 |
16.81 |
|
|
|
| 7 |
A |
2969 |
2952 |
19.71 |
|
|
|
| 8 |
A |
2965 |
2948 |
32.47 |
|
|
|
| 9 |
A |
2961 |
2944 |
23.33 |
|
|
|
| 10 |
A |
2957 |
2940 |
26.13 |
|
|
|
| 11 |
A |
1429 |
1420 |
5.36 |
|
|
|
| 12 |
A |
1417 |
1409 |
6.42 |
|
|
|
| 13 |
A |
1407 |
1399 |
7.22 |
|
|
|
| 14 |
A |
1405 |
1396 |
3.93 |
|
|
|
| 15 |
A |
1399 |
1391 |
4.86 |
|
|
|
| 16 |
A |
1384 |
1376 |
7.83 |
|
|
|
| 17 |
A |
1345 |
1337 |
3.57 |
|
|
|
| 18 |
A |
1331 |
1323 |
7.15 |
|
|
|
| 19 |
A |
1287 |
1279 |
2.58 |
|
|
|
| 20 |
A |
1272 |
1265 |
1.54 |
|
|
|
| 21 |
A |
1207 |
1200 |
29.85 |
|
|
|
| 22 |
A |
1132 |
1126 |
14.10 |
|
|
|
| 23 |
A |
1109 |
1103 |
1.15 |
|
|
|
| 24 |
A |
1028 |
1022 |
4.06 |
|
|
|
| 25 |
A |
932 |
927 |
4.39 |
|
|
|
| 26 |
A |
920 |
915 |
3.33 |
|
|
|
| 27 |
A |
916 |
911 |
4.05 |
|
|
|
| 28 |
A |
893 |
888 |
1.25 |
|
|
|
| 29 |
A |
875 |
870 |
0.84 |
|
|
|
| 30 |
A |
709 |
705 |
1.04 |
|
|
|
| 31 |
A |
616 |
612 |
2.25 |
|
|
|
| 32 |
A |
409 |
406 |
0.82 |
|
|
|
| 33 |
A |
333 |
331 |
1.02 |
|
|
|
| 34 |
A |
315 |
313 |
0.34 |
|
|
|
| 35 |
A |
248 |
247 |
0.10 |
|
|
|
| 36 |
A |
221 |
220 |
0.00 |
|
|
|
| 37 |
A |
195 |
194 |
0.46 |
|
|
|
| 38 |
A |
154 |
154 |
0.49 |
|
|
|
| 39 |
A |
65 |
65 |
1.33 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28073.9 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 27911.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.159 |
|
|
|
| 2 |
S |
-0.056 |
|
|
|
| 3 |
C |
-0.241 |
|
|
|
| 4 |
C |
0.003 |
|
|
|
| 5 |
C |
-0.009 |
|
|
|
| 6 |
H |
0.068 |
|
|
|
| 7 |
H |
0.068 |
|
|
|
| 8 |
H |
0.064 |
|
|
|
| 9 |
H |
0.043 |
|
|
|
| 10 |
H |
0.029 |
|
|
|
| 11 |
H |
0.041 |
|
|
|
| 12 |
H |
0.039 |
|
|
|
| 13 |
H |
0.032 |
|
|
|
| 14 |
H |
0.034 |
|
|
|
| 15 |
H |
0.043 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.231 |
1.188 |
0.863 |
1.486 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.274 |
1.222 |
0.411 |
| y |
1.222 |
-41.849 |
0.081 |
| z |
0.411 |
0.081 |
-41.362 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.331 |
1.222 |
0.411 |
| y |
1.222 |
-2.531 |
0.081 |
| z |
0.411 |
0.081 |
-1.800 |
|
| Polar |
|---|
| 3z2-r2 | -3.601 |
|---|
| x2-y2 | 4.574 |
|---|
| xy | 1.222 |
|---|
| xz | 0.411 |
|---|
| yz | 0.081 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.965 |
0.133 |
0.014 |
| y |
0.133 |
9.257 |
0.362 |
| z |
0.014 |
0.362 |
7.827 |
<r2> (average value of r
2) Å
2
| <r2> |
185.489 |
| (<r2>)1/2 |
13.619 |