return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8 (1,3-Pentadiene, (Z)-)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-194.712590
Energy at 298.15K-194.719967
HF Energy-193.971398
Nuclear repulsion energy154.137956
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3263 3131 15.32      
2 A' 3203 3073 15.47      
3 A' 3186 3057 27.34      
4 A' 3168 3040 10.03      
5 A' 3166 3037 7.01      
6 A' 3157 3028 1.32      
7 A' 3056 2932 29.78      
8 A' 1752 1681 5.00      
9 A' 1676 1608 1.75      
10 A' 1492 1432 4.08      
11 A' 1475 1415 10.25      
12 A' 1424 1366 1.50      
13 A' 1391 1335 2.07      
14 A' 1318 1264 1.80      
15 A' 1284 1232 1.13      
16 A' 1191 1143 0.55      
17 A' 1062 1019 0.08      
18 A' 971 932 7.43      
19 A' 901 864 1.14      
20 A' 611 586 4.03      
21 A' 385 369 3.00      
22 A' 220 211 0.22      
23 A" 3118 2992 20.67      
24 A" 1481 1421 5.18      
25 A" 1059 1016 1.59      
26 A" 1025 983 12.82      
27 A" 979 939 3.20      
28 A" 921 883 31.37      
29 A" 794 762 8.45      
30 A" 633 608 21.24      
31 A" 351 337 0.14      
32 A" 123 118 0.00      
33 A" 109 105 0.47      

Unscaled Zero Point Vibrational Energy (zpe) 24971.7 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 23957.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.52034 0.08659 0.07530

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 2.413 0.150 0.000
C2 1.112 -0.221 0.000
C3 0.000 0.747 0.000
C4 -1.325 0.459 0.000
C5 -1.967 -0.909 0.000
H6 2.704 1.208 0.000
H7 3.220 -0.592 0.000
H8 0.867 -1.291 0.000
H9 0.296 1.806 0.000
H10 -2.024 1.307 0.000
H11 -1.231 -1.729 0.000
H12 -2.614 -1.031 0.889
H13 -2.614 -1.031 -0.889

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13
C11.35292.48573.75034.50621.09781.09572.11332.68754.58544.09935.24015.2401
C21.35291.47442.52963.15492.13982.14011.09722.18503.48862.78573.91563.9156
C32.48571.47441.35572.57132.74313.48692.21441.09942.10022.76473.28443.2844
C43.75032.52961.35571.51084.09794.66412.80392.10771.09972.18922.16162.1616
C54.50623.15492.57131.51085.12865.19652.85943.53442.21681.10191.10651.1065
H61.09782.13982.74314.09795.12861.87223.10182.48094.72924.91005.83885.8388
H71.09572.14013.48694.66415.19651.87222.45473.78075.57694.59345.91785.9178
H82.11331.09722.21442.80392.85943.10182.45473.14863.88662.14263.60223.6022
H92.68752.18501.09942.10773.53442.48093.78073.14862.37333.85014.16044.1604
H104.58543.48862.10021.09972.21684.72925.57693.88662.37333.13772.57012.5701
H114.09932.78572.76472.18921.10194.91004.59342.14263.85013.13771.78661.7866
H125.24013.91563.28442.16161.10655.83885.91783.60224.16042.57011.78661.7782
H135.24013.91563.28442.16161.10655.83885.91783.60224.16042.57011.78661.7782

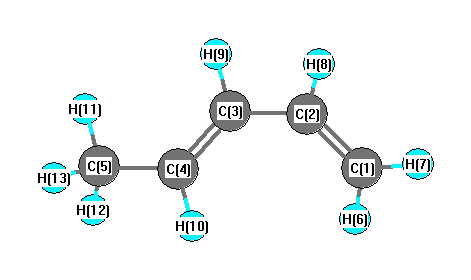
picture of 1,3-Pentadiene, (Z)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 123.029 C1 C2 H8 118.834
C2 C1 H6 121.299 C2 C1 H7 121.503
C2 C3 C4 126.659 C2 C3 H9 115.421
C3 C2 H8 118.137 C3 C4 C5 127.456
C3 C4 H10 117.207 C4 C3 H9 117.920
C4 C5 H11 112.899 C4 C5 H12 110.401
C4 C5 H13 110.401 C5 C4 H10 115.337
H6 C1 H7 117.198 H11 C5 H12 107.993
H11 C5 H13 107.993 H12 C5 H13 106.931
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability