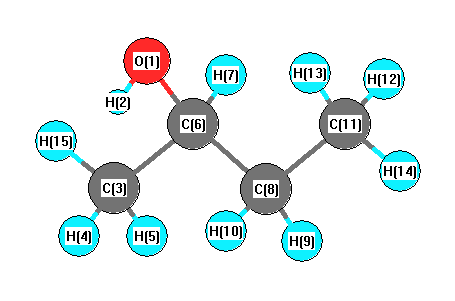
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3625 |
3604 |
6.47 |
|
|
|
| 2 |
A |
3068 |
3050 |
24.71 |
|
|
|
| 3 |
A |
3065 |
3047 |
12.85 |
|
|
|
| 4 |
A |
3049 |
3032 |
35.73 |
|
|
|
| 5 |
A |
3037 |
3019 |
31.68 |
|
|
|
| 6 |
A |
2989 |
2972 |
26.14 |
|
|
|
| 7 |
A |
2972 |
2955 |
29.94 |
|
|
|
| 8 |
A |
2962 |
2944 |
45.15 |
|
|
|
| 9 |
A |
2955 |
2938 |
7.29 |
|
|
|
| 10 |
A |
2930 |
2913 |
19.74 |
|
|
|
| 11 |
A |
1433 |
1425 |
5.63 |
|
|
|
| 12 |
A |
1417 |
1409 |
6.63 |
|
|
|
| 13 |
A |
1417 |
1409 |
14.67 |
|
|
|
| 14 |
A |
1413 |
1405 |
0.43 |
|
|
|
| 15 |
A |
1399 |
1391 |
0.77 |
|
|
|
| 16 |
A |
1369 |
1361 |
38.18 |
|
|
|
| 17 |
A |
1340 |
1332 |
2.75 |
|
|
|
| 18 |
A |
1334 |
1326 |
11.81 |
|
|
|
| 19 |
A |
1326 |
1318 |
1.35 |
|
|
|
| 20 |
A |
1287 |
1280 |
1.49 |
|
|
|
| 21 |
A |
1260 |
1253 |
5.96 |
|
|
|
| 22 |
A |
1210 |
1203 |
4.71 |
|
|
|
| 23 |
A |
1140 |
1134 |
19.41 |
|
|
|
| 24 |
A |
1101 |
1095 |
18.05 |
|
|
|
| 25 |
A |
1055 |
1049 |
58.86 |
|
|
|
| 26 |
A |
1032 |
1026 |
1.00 |
|
|
|
| 27 |
A |
968 |
962 |
21.65 |
|
|
|
| 28 |
A |
938 |
933 |
9.59 |
|
|
|
| 29 |
A |
895 |
890 |
9.23 |
|
|
|
| 30 |
A |
795 |
791 |
4.62 |
|
|
|
| 31 |
A |
753 |
749 |
0.81 |
|
|
|
| 32 |
A |
483 |
480 |
14.30 |
|
|
|
| 33 |
A |
450 |
447 |
6.33 |
|
|
|
| 34 |
A |
365 |
363 |
18.97 |
|
|
|
| 35 |
A |
313 |
311 |
80.91 |
|
|
|
| 36 |
A |
249 |
248 |
0.32 |
|
|
|
| 37 |
A |
231 |
230 |
3.01 |
|
|
|
| 38 |
A |
222 |
221 |
0.09 |
|
|
|
| 39 |
A |
123 |
122 |
0.14 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28985.5 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 28817.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.238 |
|
|
|
| 2 |
H |
0.132 |
|
|
|
| 3 |
C |
-0.014 |
|
|
|
| 4 |
H |
0.018 |
|
|
|
| 5 |
H |
0.028 |
|
|
|
| 6 |
C |
0.008 |
|
|
|
| 7 |
H |
0.017 |
|
|
|
| 8 |
C |
-0.052 |
|
|
|
| 9 |
H |
0.016 |
|
|
|
| 10 |
H |
0.006 |
|
|
|
| 11 |
C |
-0.039 |
|
|
|
| 12 |
H |
0.030 |
|
|
|
| 13 |
H |
0.031 |
|
|
|
| 14 |
H |
0.024 |
|
|
|
| 15 |
H |
0.033 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.197 |
-1.033 |
0.882 |
1.372 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.722 |
0.118 |
-0.612 |
| y |
0.118 |
-35.369 |
2.186 |
| z |
-0.612 |
2.186 |
-31.128 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.474 |
0.118 |
-0.612 |
| y |
0.118 |
-2.944 |
2.186 |
| z |
-0.612 |
2.186 |
3.418 |
|
| Polar |
|---|
| 3z2-r2 | 6.836 |
|---|
| x2-y2 | 1.647 |
|---|
| xy | 0.118 |
|---|
| xz | -0.612 |
|---|
| yz | 2.186 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.676 |
0.175 |
-0.062 |
| y |
0.175 |
7.369 |
0.105 |
| z |
-0.062 |
0.105 |
7.250 |
<r2> (average value of r
2) Å
2
| <r2> |
147.422 |
| (<r2>)1/2 |
12.142 |