return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10 (Cyclopropane, 1,1-dimethyl-)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-195.924926
Energy at 298.15K-195.935736
HF Energy-195.145193
Nuclear repulsion energy183.218342
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3155 3027 11.26      
2 A 3134 3007 37.07      
3 A 3044 2921 36.92      
4 A 1536 1474 0.35      
5 A 1501 1440 5.08      
6 A 1433 1375 0.11      
7 A 1372 1317 1.14      
8 A 1098 1053 0.23      
9 A 1054 1011 5.21      
10 A 967 928 10.96      
11 A 700 671 1.66      
12 A 350 335 0.05      
13 A 3234 3103 0.00      
14 A 3116 2990 0.00      
15 A 1480 1420 0.00      
16 A 1188 1140 0.00      
17 A 1066 1022 0.00      
18 A 896 860 0.00      
19 A 332 319 0.00      
20 A 222 213 0.00      
21 A 3147 3019 18.97      
22 A 3119 2992 60.28      
23 A 1498 1437 5.27      
24 A 1464 1404 2.79      
25 A 1168 1121 3.87      
26 A 1079 1035 0.40      
27 A 878 843 6.44      
28 A 393 377 0.75      
29 A 251 241 0.00      
30 A 3247 3115 23.56      
31 A 3132 3005 16.38      
32 A 3040 2917 33.53      
33 A 1492 1432 3.24      
34 A 1428 1370 4.66      
35 A 1340 1285 0.54      
36 A 977 937 0.56      
37 A 962 923 0.02      
38 A 788 756 0.08      
39 A 357 343 0.94      

Unscaled Zero Point Vibrational Energy (zpe) 30319.1 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 29088.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.20272 0.17131 0.12565

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.153
C2 0.762 0.000 -1.159
C3 -0.762 0.000 -1.159
C4 0.000 1.280 0.974
C5 0.000 -1.280 0.974
H6 1.273 0.922 -1.460
H7 1.273 -0.922 -1.460
H8 -1.273 -0.922 -1.460
H9 -1.273 0.922 -1.460
H10 -0.895 1.333 1.623
H11 -0.895 -1.333 1.623
H12 0.895 -1.333 1.623
H13 0.895 1.333 1.623
H14 0.000 2.173 0.322
H15 0.000 -2.173 0.322

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15
C11.51731.51731.52121.52122.25232.25232.25232.25232.17702.17702.17702.17702.17952.1795
C21.51731.52432.60212.60211.09641.09642.25422.25423.50203.50203.08783.08782.73802.7380
C31.51731.52432.60212.60212.25422.25421.09641.09643.08783.08783.50203.50202.73802.7380
C41.52122.60212.60212.56092.77033.52123.52122.77031.10702.83752.83751.10701.10523.5143
C51.52122.60212.60212.56093.52122.77032.77033.52122.83751.10701.10702.83753.51431.1052
H62.25231.09642.25422.77033.52121.84473.14332.54513.79164.39243.83883.13362.52213.7918
H72.25231.09642.25423.52122.77031.84472.54513.14334.39243.79163.13363.83883.79182.5221
H82.25232.25421.09643.52122.77033.14332.54511.84473.83883.13363.79164.39243.79182.5221
H92.25232.25421.09642.77033.52122.54513.14331.84473.13363.83884.39243.79162.52213.7918
H102.17703.50203.08781.10702.83753.79164.39243.83883.13362.66553.21101.79051.78883.8449
H112.17703.50203.08782.83751.10704.39243.79163.13363.83882.66551.79053.21103.84491.7888
H122.17703.08783.50202.83751.10703.83883.13363.79164.39243.21101.79052.66553.84491.7888
H132.17703.08783.50201.10702.83753.13363.83884.39243.79161.79053.21102.66551.78883.8449
H142.17952.73802.73801.10523.51432.52213.79183.79182.52211.78883.84493.84491.78884.3458
H152.17952.73802.73803.51431.10523.79182.52212.52213.79183.84491.78881.78883.84494.3458

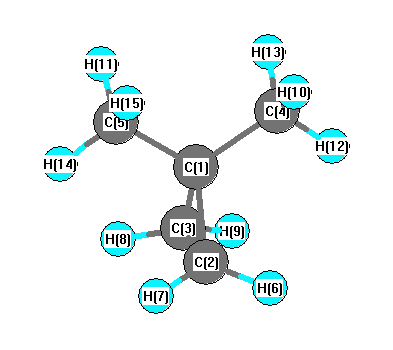
picture of Cyclopropane, 1,1-dimethyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 59.846 C1 C2 H6 118.134
C1 C2 H7 118.134 C1 C3 C2 59.846
C1 C3 H8 118.134 C1 C3 H9 118.134
C1 C4 H10 110.871 C1 C4 H13 110.871
C1 C4 H14 111.177 C1 C5 H11 110.871
C1 C5 H12 110.871 C1 C5 H15 111.177
C2 C1 C3 60.307 C2 C1 C4 117.827
C2 C1 C5 117.827 C2 C3 H8 117.744
C2 C3 H9 117.744 C3 C1 C4 117.827
C3 C1 C5 117.827 C3 C2 H6 117.744
C3 C2 H7 117.744 C4 C1 C5 114.651
H6 C2 H7 114.547 H8 C3 H9 114.547
H10 C4 H13 107.943 H10 C4 H14 107.921
H11 C5 H12 107.943 H11 C5 H15 107.921
H12 C5 H15 107.921 H13 C4 H14 107.921
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability