return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10 (Cyclopropane, 1,1-dimethyl-)

using model chemistry: CCD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at CCD/cc-pVDZ
 hartrees
Energy at 0K-195.920833
Energy at 298.15K-195.931658
HF Energy-195.145331
Nuclear repulsion energy183.293980
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3164 3027 10.93      
2 A 3146 3010 35.01      
3 A 3054 2922 34.83      
4 A 1542 1475 0.39      
5 A 1504 1439 5.31      
6 A 1436 1374 0.07      
7 A 1377 1317 1.00      
8 A 1103 1055 0.32      
9 A 1058 1012 5.25      
10 A 972 930 11.46      
11 A 703 673 1.68      
12 A 350 335 0.04      
13 A 3245 3104 0.00      
14 A 3129 2994 0.00      
15 A 1482 1418 0.00      
16 A 1191 1140 0.00      
17 A 1068 1021 0.00      
18 A 899 860 0.00      
19 A 333 319 0.00      
20 A 222 212 0.00      
21 A 3156 3020 17.28      
22 A 3132 2996 57.13      
23 A 1500 1435 5.40      
24 A 1467 1404 3.11      
25 A 1172 1122 4.01      
26 A 1084 1037 0.45      
27 A 884 846 6.22      
28 A 394 377 0.78      
29 A 251 240 0.00      
30 A 3258 3116 21.94      
31 A 3144 3008 15.28      
32 A 3050 2918 32.11      
33 A 1495 1430 3.26      
34 A 1431 1369 5.07      
35 A 1343 1285 0.48      
36 A 979 937 0.61      
37 A 964 922 0.02      
38 A 791 757 0.07      
39 A 359 343 0.91      

Unscaled Zero Point Vibrational Energy (zpe) 30415.5 cm-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 29098.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/cc-pVDZ
ABC
0.20285 0.17148 0.12573

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/cc-pVDZ

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.153
C2 0.762 0.000 -1.158
C3 -0.762 0.000 -1.158
C4 0.000 1.280 0.974
C5 0.000 -1.280 0.974
H6 1.272 0.922 -1.460
H7 1.272 -0.922 -1.460
H8 -1.272 -0.922 -1.460
H9 -1.272 0.922 -1.460
H10 -0.895 1.333 1.623
H11 -0.895 -1.333 1.623
H12 0.895 -1.333 1.623
H13 0.895 1.333 1.623
H14 0.000 2.172 0.322
H15 0.000 -2.172 0.322

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15
C11.51621.51621.52111.52112.25102.25102.25102.25102.17652.17652.17652.17652.17902.1790
C21.51621.52302.60112.60111.09591.09592.25272.25273.50053.50053.08673.08672.73722.7372
C31.51621.52302.60112.60112.25272.25271.09591.09593.08673.08673.50053.50052.73722.7372
C41.52112.60112.60112.56062.76913.51983.51982.76911.10652.83702.83701.10651.10473.5137
C51.52112.60112.60112.56063.51982.76912.76913.51982.83701.10651.10652.83703.51371.1047
H62.25101.09592.25272.76913.51981.84393.14152.54353.78984.39053.83733.13222.52123.7905
H72.25101.09592.25273.51982.76911.84392.54353.14154.39053.78983.13223.83733.79052.5212
H82.25102.25271.09593.51982.76913.14152.54351.84393.83733.13223.78984.39053.79052.5212
H92.25102.25271.09592.76913.51982.54353.14151.84393.13223.83734.39053.78982.52123.7905
H102.17653.50053.08671.10652.83703.78984.39053.83733.13222.66513.21031.78971.78793.8440
H112.17653.50053.08672.83701.10654.39053.78983.13223.83732.66511.78973.21033.84401.7879
H122.17653.08673.50052.83701.10653.83733.13223.78984.39053.21031.78972.66513.84401.7879
H132.17653.08673.50051.10652.83703.13223.83734.39053.78981.78973.21032.66511.78793.8440
H142.17902.73722.73721.10473.51372.52123.79053.79052.52121.78793.84403.84401.78794.3448
H152.17902.73722.73723.51371.10473.79052.52122.52123.79053.84401.78791.78793.84404.3448

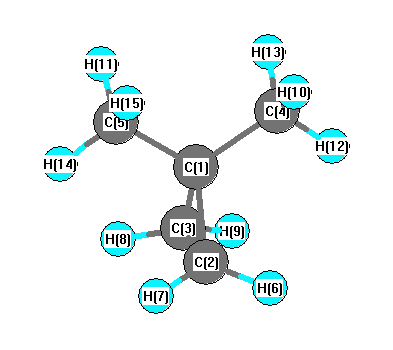
picture of Cyclopropane, 1,1-dimethyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 59.851 C1 C2 H6 118.129
C1 C2 H7 118.129 C1 C3 C2 59.851
C1 C3 H8 118.129 C1 C3 H9 118.129
C1 C4 H10 110.871 C1 C4 H13 110.871
C1 C4 H14 111.181 C1 C5 H11 110.871
C1 C5 H12 110.871 C1 C5 H15 111.181
C2 C1 C3 60.297 C2 C1 C4 117.831
C2 C1 C5 117.831 C2 C3 H8 117.746
C2 C3 H9 117.746 C3 C1 C4 117.831
C3 C1 C5 117.831 C3 C2 H6 117.746
C3 C2 H7 117.746 C4 C1 C5 114.647
H6 C2 H7 114.549 H8 C3 H9 114.549
H10 C4 H13 107.946 H10 C4 H14 107.917
H11 C5 H12 107.946 H11 C5 H15 107.917
H12 C5 H15 107.917 H13 C4 H14 107.917
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability